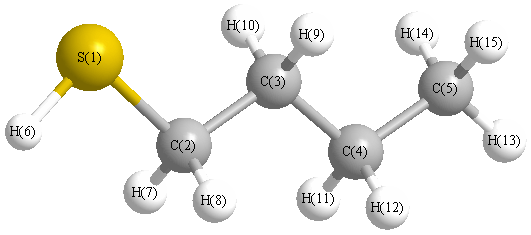
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3069 |
2990 |
64.87 |
|
|
|
| 2 |
A' |
3027 |
2950 |
47.24 |
|
|
|
| 3 |
A' |
2993 |
2917 |
37.81 |
|
|
|
| 4 |
A' |
2981 |
2905 |
27.12 |
|
|
|
| 5 |
A' |
2973 |
2897 |
43.39 |
|
|
|
| 6 |
A' |
2471 |
2408 |
43.83 |
|
|
|
| 7 |
A' |
1535 |
1496 |
9.21 |
|
|
|
| 8 |
A' |
1521 |
1482 |
1.09 |
|
|
|
| 9 |
A' |
1515 |
1476 |
1.48 |
|
|
|
| 10 |
A' |
1497 |
1459 |
1.95 |
|
|
|
| 11 |
A' |
1430 |
1394 |
3.53 |
|
|
|
| 12 |
A' |
1394 |
1358 |
5.59 |
|
|
|
| 13 |
A' |
1347 |
1313 |
11.82 |
|
|
|
| 14 |
A' |
1267 |
1235 |
33.93 |
|
|
|
| 15 |
A' |
1129 |
1100 |
3.46 |
|
|
|
| 16 |
A' |
1072 |
1045 |
1.29 |
|
|
|
| 17 |
A' |
1027 |
1001 |
0.81 |
|
|
|
| 18 |
A' |
913 |
890 |
2.80 |
|
|
|
| 19 |
A' |
809 |
788 |
2.02 |
|
|
|
| 20 |
A' |
691 |
673 |
9.70 |
|
|
|
| 21 |
A' |
383 |
373 |
1.11 |
|
|
|
| 22 |
A' |
307 |
300 |
2.00 |
|
|
|
| 23 |
A' |
148 |
145 |
2.00 |
|
|
|
| 24 |
A" |
3102 |
3023 |
53.02 |
|
|
|
| 25 |
A" |
3068 |
2990 |
86.99 |
|
|
|
| 26 |
A" |
3043 |
2966 |
11.47 |
|
|
|
| 27 |
A" |
3010 |
2933 |
7.23 |
|
|
|
| 28 |
A" |
1521 |
1482 |
8.75 |
|
|
|
| 29 |
A" |
1332 |
1298 |
0.29 |
|
|
|
| 30 |
A" |
1309 |
1275 |
0.68 |
|
|
|
| 31 |
A" |
1234 |
1202 |
0.48 |
|
|
|
| 32 |
A" |
1075 |
1048 |
0.98 |
|
|
|
| 33 |
A" |
934 |
910 |
2.01 |
|
|
|
| 34 |
A" |
801 |
781 |
0.12 |
|
|
|
| 35 |
A" |
750 |
731 |
5.73 |
|
|
|
| 36 |
A" |
229 |
223 |
0.00 |
|
|
|
| 37 |
A" |
165 |
160 |
21.74 |
|
|
|
| 38 |
A" |
106 |
103 |
1.25 |
|
|
|
| 39 |
A" |
92 |
90 |
4.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28634.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 27904.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.050 |
|
|
|
| 2 |
C |
-0.554 |
|
|
|
| 3 |
C |
-0.192 |
|
|
|
| 4 |
C |
-0.200 |
|
|
|
| 5 |
C |
-0.472 |
|
|
|
| 6 |
H |
0.041 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
H |
0.143 |
|
|
|
| 10 |
H |
0.143 |
|
|
|
| 11 |
H |
0.126 |
|
|
|
| 12 |
H |
0.126 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
| 14 |
H |
0.144 |
|
|
|
| 15 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.064 |
0.278 |
0.000 |
2.082 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.033 |
2.607 |
0.000 |
| y |
2.607 |
-37.785 |
0.000 |
| z |
0.000 |
0.000 |
-41.706 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.288 |
2.607 |
0.000 |
| y |
2.607 |
5.085 |
0.000 |
| z |
0.000 |
0.000 |
-0.796 |
|
| Polar |
|---|
| 3z2-r2 | -1.593 |
|---|
| x2-y2 | -6.249 |
|---|
| xy | 2.607 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.016 |
-1.685 |
0.000 |
| y |
-1.685 |
11.828 |
0.000 |
| z |
0.000 |
0.000 |
7.798 |
<r2> (average value of r
2) Å
2
| <r2> |
188.175 |
| (<r2>)1/2 |
13.718 |