Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -42.065680 |
| Energy at 298.15K | -42.071388 |
| Nuclear repulsion energy | 84.652125 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3251 |
3168 |
20.35 |
|
|
|
| 2 |
A1 |
3145 |
3065 |
8.55 |
|
|
|
| 3 |
A1 |
3120 |
3040 |
36.52 |
|
|
|
| 4 |
A1 |
1698 |
1655 |
4.08 |
|
|
|
| 5 |
A1 |
1436 |
1399 |
0.68 |
|
|
|
| 6 |
A1 |
1349 |
1314 |
5.93 |
|
|
|
| 7 |
A1 |
1165 |
1136 |
16.05 |
|
|
|
| 8 |
A1 |
826 |
805 |
1.98 |
|
|
|
| 9 |
A1 |
499 |
486 |
0.10 |
|
|
|
| 10 |
A1 |
216 |
211 |
0.12 |
|
|
|
| 11 |
A2 |
958 |
933 |
0.00 |
|
|
|
| 12 |
A2 |
885 |
862 |
0.00 |
|
|
|
| 13 |
A2 |
704 |
686 |
0.00 |
|
|
|
| 14 |
A2 |
99 |
97 |
0.00 |
|
|
|
| 15 |
B1 |
974 |
949 |
80.53 |
|
|
|
| 16 |
B1 |
890 |
867 |
144.37 |
|
|
|
| 17 |
B1 |
674 |
657 |
8.62 |
|
|
|
| 18 |
B1 |
89 |
87 |
3.47 |
|
|
|
| 19 |
B2 |
3251 |
3168 |
7.25 |
|
|
|
| 20 |
B2 |
3143 |
3063 |
0.85 |
|
|
|
| 21 |
B2 |
3113 |
3034 |
6.63 |
|
|
|
| 22 |
B2 |
1646 |
1604 |
422.37 |
|
|
|
| 23 |
B2 |
1422 |
1386 |
47.69 |
|
|
|
| 24 |
B2 |
1323 |
1289 |
9.37 |
|
|
|
| 25 |
B2 |
1172 |
1142 |
446.45 |
|
|
|
| 26 |
B2 |
1023 |
997 |
26.90 |
|
|
|
| 27 |
B2 |
475 |
463 |
6.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19271.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 18780.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.334 |
| C2 |
0.000 |
1.211 |
-0.386 |
| C3 |
0.000 |
-1.211 |
-0.386 |
| C4 |
0.000 |
2.400 |
0.251 |
| C5 |
0.000 |
-2.400 |
0.251 |
| H6 |
0.000 |
1.102 |
-1.472 |
| H7 |
0.000 |
-1.102 |
-1.472 |
| H8 |
0.000 |
3.319 |
-0.326 |
| H9 |
0.000 |
2.460 |
1.335 |
| H10 |
0.000 |
-3.319 |
-0.326 |
| H11 |
0.000 |
-2.460 |
1.335 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.4092 | 1.4092 | 2.4012 | 2.4012 | 2.1162 | 2.1162 | 3.3844 | 2.6555 | 3.3844 | 2.6555 |
C2 | 1.4092 | | 2.4227 | 1.3480 | 3.6667 | 1.0920 | 2.5560 | 2.1089 | 2.1259 | 4.5311 | 4.0544 | C3 | 1.4092 | 2.4227 | | 3.6667 | 1.3480 | 2.5560 | 1.0920 | 4.5311 | 4.0544 | 2.1089 | 2.1259 | C4 | 2.4012 | 1.3480 | 3.6667 | | 4.7994 | 2.1568 | 3.9027 | 1.0854 | 1.0861 | 5.7481 | 4.9790 | C5 | 2.4012 | 3.6667 | 1.3480 | 4.7994 | | 3.9027 | 2.1568 | 5.7481 | 4.9790 | 1.0854 | 1.0861 | H6 | 2.1162 | 1.0920 | 2.5560 | 2.1568 | 3.9027 | | 2.2043 | 2.4961 | 3.1183 | 4.5678 | 4.5352 | H7 | 2.1162 | 2.5560 | 1.0920 | 3.9027 | 2.1568 | 2.2043 | | 4.5678 | 4.5352 | 2.4961 | 3.1183 | H8 | 3.3844 | 2.1089 | 4.5311 | 1.0854 | 5.7481 | 2.4961 | 4.5678 | | 1.8700 | 6.6388 | 6.0131 | H9 | 2.6555 | 2.1259 | 4.0544 | 1.0861 | 4.9790 | 3.1183 | 4.5352 | 1.8700 | | 6.0131 | 4.9196 | H10 | 3.3844 | 4.5311 | 2.1089 | 5.7481 | 1.0854 | 4.5678 | 2.4961 | 6.6388 | 6.0131 | | 1.8700 | H11 | 2.6555 | 4.0544 | 2.1259 | 4.9790 | 1.0861 | 4.5352 | 3.1183 | 6.0131 | 4.9196 | 1.8700 | |
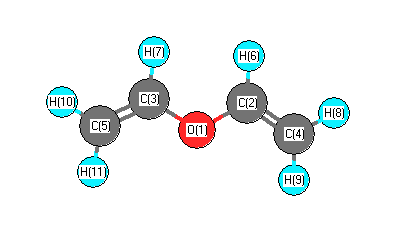 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
121.109 |
|
O1 |
C2 |
H6 |
114.990 |
| O1 |
C3 |
C5 |
121.109 |
|
O1 |
C3 |
H7 |
114.990 |
| C2 |
O1 |
C3 |
118.547 |
|
C2 |
C4 |
H8 |
119.764 |
| C2 |
C4 |
H9 |
121.336 |
|
C3 |
C5 |
H10 |
119.764 |
| C3 |
C5 |
H11 |
121.336 |
|
C4 |
C2 |
H6 |
123.901 |
| C5 |
C3 |
H7 |
123.901 |
|
H8 |
C4 |
H9 |
118.900 |
| H10 |
C5 |
H11 |
118.900 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.196 |
|
|
|
| 2 |
C |
0.094 |
|
|
|
| 3 |
C |
0.094 |
|
|
|
| 4 |
C |
-0.388 |
|
|
|
| 5 |
C |
-0.388 |
|
|
|
| 6 |
H |
0.114 |
|
|
|
| 7 |
H |
0.114 |
|
|
|
| 8 |
H |
0.134 |
|
|
|
| 9 |
H |
0.144 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
| 11 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.717 |
1.717 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.660 |
0.000 |
0.000 |
| y |
0.000 |
-26.770 |
0.000 |
| z |
0.000 |
0.000 |
-27.871 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.340 |
0.000 |
0.000 |
| y |
0.000 |
4.496 |
0.000 |
| z |
0.000 |
0.000 |
2.844 |
|
| Polar |
|---|
| 3z2-r2 | 5.687 |
|---|
| x2-y2 | -7.891 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.809 |
0.000 |
0.000 |
| y |
0.000 |
13.588 |
0.000 |
| z |
0.000 |
0.000 |
6.696 |
<r2> (average value of r
2) Å
2
| <r2> |
123.441 |
| (<r2>)1/2 |
11.110 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -42.065871 |
| Energy at 298.15K | -42.071831 |
| HF Energy | -42.065871 |
| Nuclear repulsion energy | 85.885203 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3247 |
3164 |
15.50 |
|
|
|
| 2 |
A |
3242 |
3159 |
22.34 |
|
|
|
| 3 |
A |
3196 |
3115 |
9.91 |
|
|
|
| 4 |
A |
3149 |
3069 |
10.88 |
|
|
|
| 5 |
A |
3136 |
3056 |
1.36 |
|
|
|
| 6 |
A |
3126 |
3046 |
5.64 |
|
|
|
| 7 |
A |
1668 |
1625 |
108.36 |
|
|
|
| 8 |
A |
1640 |
1598 |
162.91 |
|
|
|
| 9 |
A |
1437 |
1400 |
3.31 |
|
|
|
| 10 |
A |
1422 |
1386 |
15.94 |
|
|
|
| 11 |
A |
1354 |
1319 |
41.24 |
|
|
|
| 12 |
A |
1320 |
1287 |
5.74 |
|
|
|
| 13 |
A |
1206 |
1175 |
294.25 |
|
|
|
| 14 |
A |
1115 |
1086 |
13.09 |
|
|
|
| 15 |
A |
996 |
971 |
62.79 |
|
|
|
| 16 |
A |
986 |
961 |
6.08 |
|
|
|
| 17 |
A |
980 |
955 |
57.10 |
|
|
|
| 18 |
A |
904 |
881 |
70.29 |
|
|
|
| 19 |
A |
887 |
864 |
78.38 |
|
|
|
| 20 |
A |
827 |
806 |
11.28 |
|
|
|
| 21 |
A |
716 |
698 |
7.07 |
|
|
|
| 22 |
A |
700 |
682 |
2.65 |
|
|
|
| 23 |
A |
577 |
563 |
2.42 |
|
|
|
| 24 |
A |
436 |
425 |
3.86 |
|
|
|
| 25 |
A |
259 |
253 |
1.85 |
|
|
|
| 26 |
A |
195 |
190 |
10.79 |
|
|
|
| 27 |
A |
86 |
84 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19403.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 18908.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.050 |
-0.789 |
0.052 |
| C2 |
1.330 |
-0.508 |
0.014 |
| C3 |
-0.968 |
0.259 |
0.294 |
| C4 |
1.905 |
0.705 |
-0.152 |
| C5 |
-2.223 |
0.201 |
-0.196 |
| H6 |
1.886 |
-1.434 |
0.119 |
| H7 |
-0.594 |
1.051 |
0.943 |
| H8 |
2.989 |
0.775 |
-0.158 |
| H9 |
1.340 |
1.620 |
-0.301 |
| H10 |
-2.941 |
0.973 |
0.063 |
| H11 |
-2.540 |
-0.615 |
-0.840 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.4088 | 1.4137 | 2.4691 | 2.4000 | 2.0424 | 2.1155 | 3.4241 | 2.8038 | 3.3854 | 2.6499 |
C2 | 1.4088 | | 2.4385 | 1.3524 | 3.6284 | 1.0858 | 2.6447 | 2.1041 | 2.1513 | 4.5205 | 3.9638 | C3 | 1.4137 | 2.4385 | | 2.9418 | 1.3480 | 3.3233 | 1.0904 | 4.0159 | 2.7451 | 2.1110 | 2.1255 | C4 | 2.4691 | 1.3524 | 2.9418 | | 4.1589 | 2.1560 | 2.7504 | 1.0857 | 1.0863 | 4.8586 | 4.6875 | C5 | 2.4000 | 3.6284 | 1.3480 | 4.1589 | | 4.4335 | 2.1615 | 5.2430 | 3.8366 | 1.0859 | 1.0864 | H6 | 2.0424 | 1.0858 | 3.3233 | 2.1560 | 4.4335 | | 3.6066 | 2.4843 | 3.1312 | 5.3946 | 4.6022 | H7 | 2.1155 | 2.6447 | 1.0904 | 2.7504 | 2.1615 | 3.6066 | | 3.7581 | 2.3688 | 2.5080 | 3.1206 | H8 | 3.4241 | 2.1041 | 4.0159 | 1.0857 | 5.2430 | 2.4843 | 3.7581 | | 1.8582 | 5.9373 | 5.7410 | H9 | 2.8038 | 2.1513 | 2.7451 | 1.0863 | 3.8366 | 3.1312 | 2.3688 | 1.8582 | | 4.3452 | 4.5098 | H10 | 3.3854 | 4.5205 | 2.1110 | 4.8586 | 1.0859 | 5.3946 | 2.5080 | 5.9373 | 4.3452 | | 1.8697 | H11 | 2.6499 | 3.9638 | 2.1255 | 4.6875 | 1.0864 | 4.6022 | 3.1206 | 5.7410 | 4.5098 | 1.8697 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
126.801 |
|
O1 |
C2 |
H6 |
109.231 |
| O1 |
C3 |
C5 |
120.675 |
|
O1 |
C3 |
H7 |
114.689 |
| C2 |
O1 |
C3 |
119.526 |
|
C2 |
C4 |
H8 |
118.908 |
| C2 |
C4 |
H9 |
123.437 |
|
C3 |
C5 |
H10 |
119.920 |
| C3 |
C5 |
H11 |
121.274 |
|
C4 |
C2 |
H6 |
123.957 |
| C5 |
C3 |
H7 |
124.525 |
|
H8 |
C4 |
H9 |
117.642 |
| H10 |
C5 |
H11 |
118.802 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.203 |
|
|
|
| 2 |
C |
0.064 |
|
|
|
| 3 |
C |
-0.003 |
|
|
|
| 4 |
C |
-0.390 |
|
|
|
| 5 |
C |
-0.334 |
|
|
|
| 6 |
H |
0.148 |
|
|
|
| 7 |
H |
0.142 |
|
|
|
| 8 |
H |
0.139 |
|
|
|
| 9 |
H |
0.158 |
|
|
|
| 10 |
H |
0.135 |
|
|
|
| 11 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.429 |
1.072 |
0.297 |
1.192 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.516 |
-1.946 |
0.543 |
| y |
-1.946 |
-30.103 |
1.053 |
| z |
0.543 |
1.053 |
-33.207 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.139 |
-1.946 |
0.543 |
| y |
-1.946 |
-0.242 |
1.053 |
| z |
0.543 |
1.053 |
-4.898 |
|
| Polar |
|---|
| 3z2-r2 | -9.796 |
|---|
| x2-y2 | 3.587 |
|---|
| xy | -1.946 |
|---|
| xz | 0.543 |
|---|
| yz | 1.053 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.480 |
0.591 |
0.592 |
| y |
0.591 |
6.911 |
0.169 |
| z |
0.592 |
0.169 |
4.490 |
<r2> (average value of r
2) Å
2
| <r2> |
110.788 |
| (<r2>)1/2 |
10.526 |
