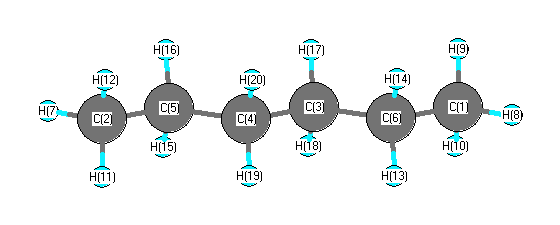
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3062 |
2984 |
0.00 |
|
|
|
| 2 |
Ag |
2977 |
2901 |
0.00 |
|
|
|
| 3 |
Ag |
2969 |
2893 |
0.00 |
|
|
|
| 4 |
Ag |
2956 |
2881 |
0.00 |
|
|
|
| 5 |
Ag |
1530 |
1491 |
0.00 |
|
|
|
| 6 |
Ag |
1516 |
1477 |
0.00 |
|
|
|
| 7 |
Ag |
1510 |
1471 |
0.00 |
|
|
|
| 8 |
Ag |
1428 |
1392 |
0.00 |
|
|
|
| 9 |
Ag |
1392 |
1356 |
0.00 |
|
|
|
| 10 |
Ag |
1343 |
1309 |
0.00 |
|
|
|
| 11 |
Ag |
1157 |
1127 |
0.00 |
|
|
|
| 12 |
Ag |
1077 |
1050 |
0.00 |
|
|
|
| 13 |
Ag |
1008 |
982 |
0.00 |
|
|
|
| 14 |
Ag |
903 |
880 |
0.00 |
|
|
|
| 15 |
Ag |
358 |
349 |
0.00 |
|
|
|
| 16 |
Ag |
290 |
283 |
0.00 |
|
|
|
| 17 |
Au |
3063 |
2985 |
252.08 |
|
|
|
| 18 |
Au |
3031 |
2954 |
70.69 |
|
|
|
| 19 |
Au |
2996 |
2920 |
0.04 |
|
|
|
| 20 |
Au |
1521 |
1482 |
16.38 |
|
|
|
| 21 |
Au |
1329 |
1295 |
0.75 |
|
|
|
| 22 |
Au |
1255 |
1223 |
0.01 |
|
|
|
| 23 |
Au |
1017 |
991 |
0.47 |
|
|
|
| 24 |
Au |
814 |
793 |
1.57 |
|
|
|
| 25 |
Au |
735 |
716 |
5.43 |
|
|
|
| 26 |
Au |
240 |
234 |
0.00 |
|
|
|
| 27 |
Au |
93 |
91 |
0.00 |
|
|
|
| 28 |
Au |
66 |
65 |
0.01 |
|
|
|
| 29 |
Bg |
3060 |
2982 |
0.00 |
|
|
|
| 30 |
Bg |
3014 |
2937 |
0.00 |
|
|
|
| 31 |
Bg |
2985 |
2908 |
0.00 |
|
|
|
| 32 |
Bg |
1521 |
1483 |
0.00 |
|
|
|
| 33 |
Bg |
1325 |
1291 |
0.00 |
|
|
|
| 34 |
Bg |
1304 |
1271 |
0.00 |
|
|
|
| 35 |
Bg |
1213 |
1182 |
0.00 |
|
|
|
| 36 |
Bg |
913 |
890 |
0.00 |
|
|
|
| 37 |
Bg |
752 |
733 |
0.00 |
|
|
|
| 38 |
Bg |
231 |
225 |
0.00 |
|
|
|
| 39 |
Bg |
136 |
132 |
0.00 |
|
|
|
| 40 |
Bu |
3062 |
2984 |
142.53 |
|
|
|
| 41 |
Bu |
2975 |
2900 |
52.60 |
|
|
|
| 42 |
Bu |
2974 |
2898 |
227.96 |
|
|
|
| 43 |
Bu |
2957 |
2882 |
12.19 |
|
|
|
| 44 |
Bu |
1537 |
1498 |
13.42 |
|
|
|
| 45 |
Bu |
1522 |
1483 |
3.51 |
|
|
|
| 46 |
Bu |
1510 |
1472 |
0.55 |
|
|
|
| 47 |
Bu |
1427 |
1390 |
9.08 |
|
|
|
| 48 |
Bu |
1384 |
1349 |
2.74 |
|
|
|
| 49 |
Bu |
1283 |
1250 |
1.04 |
|
|
|
| 50 |
Bu |
1080 |
1053 |
7.72 |
|
|
|
| 51 |
Bu |
1052 |
1026 |
0.55 |
|
|
|
| 52 |
Bu |
882 |
860 |
3.87 |
|
|
|
| 53 |
Bu |
457 |
446 |
0.10 |
|
|
|
| 54 |
Bu |
130 |
127 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41161.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 40112.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.472 |
|
|
|
| 2 |
C |
-0.472 |
|
|
|
| 3 |
C |
-0.227 |
|
|
|
| 4 |
C |
-0.227 |
|
|
|
| 5 |
C |
-0.193 |
|
|
|
| 6 |
C |
-0.193 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
| 9 |
H |
0.139 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.139 |
|
|
|
| 12 |
H |
0.139 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
| 14 |
H |
0.122 |
|
|
|
| 15 |
H |
0.122 |
|
|
|
| 16 |
H |
0.122 |
|
|
|
| 17 |
H |
0.113 |
|
|
|
| 18 |
H |
0.113 |
|
|
|
| 19 |
H |
0.113 |
|
|
|
| 20 |
H |
0.113 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.507 |
0.007 |
0.000 |
| y |
0.007 |
-43.497 |
0.000 |
| z |
0.000 |
0.000 |
-41.464 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.026 |
0.007 |
0.000 |
| y |
0.007 |
-1.512 |
0.000 |
| z |
0.000 |
0.000 |
1.538 |
|
| Polar |
|---|
| 3z2-r2 | 3.077 |
|---|
| x2-y2 | 0.990 |
|---|
| xy | 0.007 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.658 |
1.173 |
0.000 |
| y |
1.173 |
12.140 |
0.000 |
| z |
0.000 |
0.000 |
9.684 |
<r2> (average value of r
2) Å
2
| <r2> |
266.389 |
| (<r2>)1/2 |
16.321 |