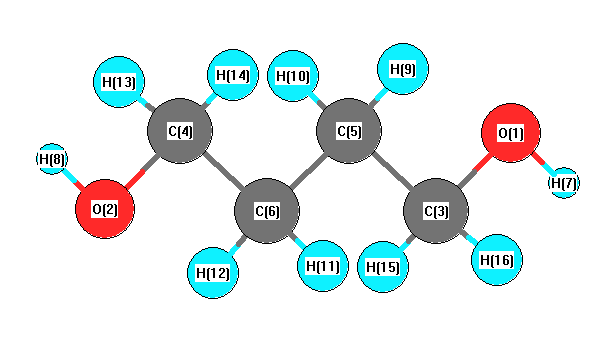
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3660 |
3566 |
0.00 |
|
|
|
| 2 |
Ag |
2996 |
2920 |
0.00 |
|
|
|
| 3 |
Ag |
2952 |
2877 |
0.00 |
|
|
|
| 4 |
Ag |
1536 |
1497 |
0.00 |
|
|
|
| 5 |
Ag |
1517 |
1478 |
0.00 |
|
|
|
| 6 |
Ag |
1442 |
1405 |
0.00 |
|
|
|
| 7 |
Ag |
1383 |
1348 |
0.00 |
|
|
|
| 8 |
Ag |
1238 |
1206 |
0.00 |
|
|
|
| 9 |
Ag |
1071 |
1043 |
0.00 |
|
|
|
| 10 |
Ag |
1034 |
1008 |
0.00 |
|
|
|
| 11 |
Ag |
992 |
967 |
0.00 |
|
|
|
| 12 |
Ag |
356 |
347 |
0.00 |
|
|
|
| 13 |
Ag |
322 |
314 |
0.00 |
|
|
|
| 14 |
Au |
3071 |
2992 |
145.70 |
|
|
|
| 15 |
Au |
2999 |
2922 |
87.94 |
|
|
|
| 16 |
Au |
1321 |
1287 |
2.99 |
|
|
|
| 17 |
Au |
1196 |
1165 |
2.30 |
|
|
|
| 18 |
Au |
947 |
923 |
1.92 |
|
|
|
| 19 |
Au |
771 |
751 |
1.41 |
|
|
|
| 20 |
Au |
273 |
266 |
330.80 |
|
|
|
| 21 |
Au |
94 |
92 |
23.23 |
|
|
|
| 22 |
Au |
71 |
70 |
6.14 |
|
|
|
| 23 |
Bg |
3040 |
2963 |
0.00 |
|
|
|
| 24 |
Bg |
3001 |
2925 |
0.00 |
|
|
|
| 25 |
Bg |
1317 |
1284 |
0.00 |
|
|
|
| 26 |
Bg |
1275 |
1243 |
0.00 |
|
|
|
| 27 |
Bg |
1164 |
1134 |
0.00 |
|
|
|
| 28 |
Bg |
826 |
805 |
0.00 |
|
|
|
| 29 |
Bg |
267 |
261 |
0.00 |
|
|
|
| 30 |
Bg |
138 |
134 |
0.00 |
|
|
|
| 31 |
Bu |
3660 |
3566 |
7.51 |
|
|
|
| 32 |
Bu |
3003 |
2926 |
96.86 |
|
|
|
| 33 |
Bu |
2952 |
2877 |
122.28 |
|
|
|
| 34 |
Bu |
1545 |
1506 |
6.86 |
|
|
|
| 35 |
Bu |
1522 |
1483 |
5.37 |
|
|
|
| 36 |
Bu |
1441 |
1405 |
17.52 |
|
|
|
| 37 |
Bu |
1310 |
1277 |
28.64 |
|
|
|
| 38 |
Bu |
1194 |
1164 |
68.04 |
|
|
|
| 39 |
Bu |
1029 |
1003 |
196.21 |
|
|
|
| 40 |
Bu |
963 |
939 |
25.82 |
|
|
|
| 41 |
Bu |
502 |
489 |
49.97 |
|
|
|
| 42 |
Bu |
134 |
130 |
9.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30761.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29977.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.471 |
|
|
|
| 2 |
O |
-0.471 |
|
|
|
| 3 |
C |
-0.177 |
|
|
|
| 4 |
C |
-0.177 |
|
|
|
| 5 |
C |
-0.204 |
|
|
|
| 6 |
C |
-0.204 |
|
|
|
| 7 |
H |
0.334 |
|
|
|
| 8 |
H |
0.334 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
H |
0.136 |
|
|
|
| 11 |
H |
0.136 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.123 |
|
|
|
| 14 |
H |
0.123 |
|
|
|
| 15 |
H |
0.123 |
|
|
|
| 16 |
H |
0.123 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.643 |
9.421 |
0.000 |
| y |
9.421 |
-46.350 |
0.000 |
| z |
0.000 |
0.000 |
-38.747 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
15.905 |
9.421 |
0.000 |
| y |
9.421 |
-13.655 |
0.000 |
| z |
0.000 |
0.000 |
-2.250 |
|
| Polar |
|---|
| 3z2-r2 | -4.500 |
|---|
| x2-y2 | 19.707 |
|---|
| xy | 9.421 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.504 |
1.167 |
0.000 |
| y |
1.167 |
9.444 |
0.000 |
| z |
0.000 |
0.000 |
6.922 |
<r2> (average value of r
2) Å
2
| <r2> |
238.072 |
| (<r2>)1/2 |
15.430 |