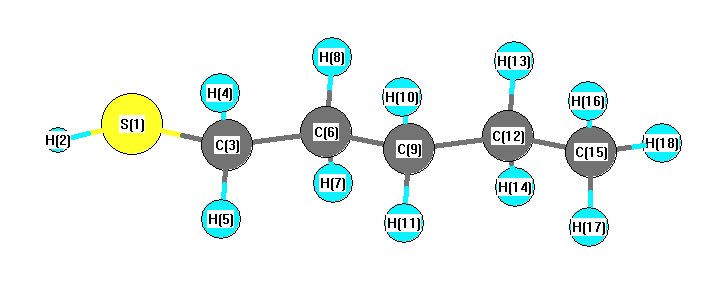
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3066 |
2988 |
67.59 |
|
|
|
| 2 |
A' |
3026 |
2949 |
49.96 |
|
|
|
| 3 |
A' |
2993 |
2917 |
40.88 |
|
|
|
| 4 |
A' |
2978 |
2902 |
30.56 |
|
|
|
| 5 |
A' |
2974 |
2898 |
71.39 |
|
|
|
| 6 |
A' |
2963 |
2887 |
10.36 |
|
|
|
| 7 |
A' |
2470 |
2407 |
44.15 |
|
|
|
| 8 |
A' |
1537 |
1498 |
10.78 |
|
|
|
| 9 |
A' |
1526 |
1487 |
0.77 |
|
|
|
| 10 |
A' |
1516 |
1477 |
0.94 |
|
|
|
| 11 |
A' |
1512 |
1474 |
0.91 |
|
|
|
| 12 |
A' |
1497 |
1459 |
1.97 |
|
|
|
| 13 |
A' |
1429 |
1392 |
4.60 |
|
|
|
| 14 |
A' |
1394 |
1358 |
0.06 |
|
|
|
| 15 |
A' |
1375 |
1340 |
12.60 |
|
|
|
| 16 |
A' |
1316 |
1282 |
14.96 |
|
|
|
| 17 |
A' |
1258 |
1226 |
24.10 |
|
|
|
| 18 |
A' |
1132 |
1103 |
5.76 |
|
|
|
| 19 |
A' |
1070 |
1043 |
0.08 |
|
|
|
| 20 |
A' |
1059 |
1032 |
2.61 |
|
|
|
| 21 |
A' |
1014 |
988 |
0.35 |
|
|
|
| 22 |
A' |
900 |
877 |
0.75 |
|
|
|
| 23 |
A' |
819 |
798 |
2.29 |
|
|
|
| 24 |
A' |
688 |
670 |
9.42 |
|
|
|
| 25 |
A' |
427 |
416 |
2.59 |
|
|
|
| 26 |
A' |
336 |
328 |
0.27 |
|
|
|
| 27 |
A' |
237 |
231 |
2.07 |
|
|
|
| 28 |
A' |
112 |
109 |
1.42 |
|
|
|
| 29 |
A" |
3101 |
3022 |
51.05 |
|
|
|
| 30 |
A" |
3064 |
2986 |
120.42 |
|
|
|
| 31 |
A" |
3046 |
2968 |
22.96 |
|
|
|
| 32 |
A" |
3020 |
2943 |
13.49 |
|
|
|
| 33 |
A" |
2997 |
2920 |
1.40 |
|
|
|
| 34 |
A" |
1521 |
1482 |
8.59 |
|
|
|
| 35 |
A" |
1329 |
1295 |
0.02 |
|
|
|
| 36 |
A" |
1327 |
1294 |
0.76 |
|
|
|
| 37 |
A" |
1286 |
1253 |
0.13 |
|
|
|
| 38 |
A" |
1227 |
1195 |
0.17 |
|
|
|
| 39 |
A" |
1079 |
1052 |
1.93 |
|
|
|
| 40 |
A" |
969 |
944 |
0.07 |
|
|
|
| 41 |
A" |
849 |
827 |
2.06 |
|
|
|
| 42 |
A" |
770 |
750 |
0.49 |
|
|
|
| 43 |
A" |
742 |
723 |
5.72 |
|
|
|
| 44 |
A" |
230 |
224 |
0.02 |
|
|
|
| 45 |
A" |
158 |
154 |
19.81 |
|
|
|
| 46 |
A" |
131 |
128 |
2.39 |
|
|
|
| 47 |
A" |
89 |
87 |
4.13 |
|
|
|
| 48 |
A" |
58 |
57 |
1.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34807.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 33920.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.053 |
|
|
|
| 2 |
H |
0.041 |
|
|
|
| 3 |
C |
-0.563 |
|
|
|
| 4 |
H |
0.178 |
|
|
|
| 5 |
H |
0.178 |
|
|
|
| 6 |
C |
-0.183 |
|
|
|
| 7 |
H |
0.142 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
| 9 |
C |
-0.239 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.118 |
|
|
|
| 12 |
C |
-0.194 |
|
|
|
| 13 |
H |
0.127 |
|
|
|
| 14 |
H |
0.127 |
|
|
|
| 15 |
C |
-0.473 |
|
|
|
| 16 |
H |
0.140 |
|
|
|
| 17 |
H |
0.140 |
|
|
|
| 18 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.762 |
-1.903 |
0.000 |
2.050 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.870 |
-1.755 |
0.000 |
| y |
-1.755 |
-55.720 |
0.000 |
| z |
0.000 |
0.000 |
-48.308 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.144 |
-1.755 |
0.000 |
| y |
-1.755 |
-10.131 |
0.000 |
| z |
0.000 |
0.000 |
0.987 |
|
| Polar |
|---|
| 3z2-r2 | 1.975 |
|---|
| x2-y2 | 12.850 |
|---|
| xy | -1.755 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.448 |
-1.927 |
0.000 |
| y |
-1.927 |
14.372 |
0.000 |
| z |
0.000 |
0.000 |
9.254 |
<r2> (average value of r
2) Å
2
| <r2> |
279.306 |
| (<r2>)1/2 |
16.712 |