Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -44.713215 |
| Energy at 298.15K | -44.727468 |
| Nuclear repulsion energy | 143.426796 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3499 |
3410 |
5.40 |
|
|
|
| 2 |
A' |
3052 |
2974 |
150.53 |
|
|
|
| 3 |
A' |
3039 |
2961 |
8.09 |
|
|
|
| 4 |
A' |
3026 |
2948 |
88.37 |
|
|
|
| 5 |
A' |
2988 |
2912 |
18.81 |
|
|
|
| 6 |
A' |
2966 |
2890 |
41.89 |
|
|
|
| 7 |
A' |
2856 |
2783 |
213.69 |
|
|
|
| 8 |
A' |
1525 |
1486 |
2.62 |
|
|
|
| 9 |
A' |
1512 |
1473 |
7.19 |
|
|
|
| 10 |
A' |
1506 |
1468 |
8.40 |
|
|
|
| 11 |
A' |
1415 |
1379 |
3.52 |
|
|
|
| 12 |
A' |
1380 |
1345 |
0.09 |
|
|
|
| 13 |
A' |
1307 |
1274 |
0.49 |
|
|
|
| 14 |
A' |
1285 |
1253 |
2.72 |
|
|
|
| 15 |
A' |
1166 |
1137 |
3.10 |
|
|
|
| 16 |
A' |
1059 |
1032 |
1.03 |
|
|
|
| 17 |
A' |
1048 |
1021 |
8.43 |
|
|
|
| 18 |
A' |
894 |
871 |
5.41 |
|
|
|
| 19 |
A' |
864 |
842 |
4.69 |
|
|
|
| 20 |
A' |
806 |
786 |
0.59 |
|
|
|
| 21 |
A' |
673 |
656 |
98.49 |
|
|
|
| 22 |
A' |
530 |
516 |
51.48 |
|
|
|
| 23 |
A' |
424 |
413 |
18.10 |
|
|
|
| 24 |
A' |
373 |
364 |
3.40 |
|
|
|
| 25 |
A' |
233 |
227 |
1.66 |
|
|
|
| 26 |
A" |
3043 |
2965 |
1.66 |
|
|
|
| 27 |
A" |
3035 |
2958 |
135.56 |
|
|
|
| 28 |
A" |
2987 |
2911 |
57.96 |
|
|
|
| 29 |
A" |
2847 |
2774 |
46.57 |
|
|
|
| 30 |
A" |
1511 |
1473 |
0.00 |
|
|
|
| 31 |
A" |
1499 |
1460 |
1.47 |
|
|
|
| 32 |
A" |
1484 |
1446 |
4.02 |
|
|
|
| 33 |
A" |
1375 |
1340 |
1.68 |
|
|
|
| 34 |
A" |
1370 |
1335 |
1.75 |
|
|
|
| 35 |
A" |
1352 |
1317 |
34.22 |
|
|
|
| 36 |
A" |
1290 |
1257 |
2.21 |
|
|
|
| 37 |
A" |
1192 |
1162 |
8.32 |
|
|
|
| 38 |
A" |
1173 |
1143 |
9.09 |
|
|
|
| 39 |
A" |
1135 |
1106 |
15.48 |
|
|
|
| 40 |
A" |
1075 |
1047 |
3.39 |
|
|
|
| 41 |
A" |
982 |
957 |
0.91 |
|
|
|
| 42 |
A" |
875 |
852 |
0.53 |
|
|
|
| 43 |
A" |
824 |
803 |
0.77 |
|
|
|
| 44 |
A" |
445 |
433 |
1.11 |
|
|
|
| 45 |
A" |
235 |
229 |
0.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34576.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 33694.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.640 |
1.346 |
0.000 |
| H2 |
0.571 |
2.444 |
0.000 |
| H3 |
1.714 |
1.095 |
0.000 |
| C4 |
-0.022 |
0.764 |
1.281 |
| C5 |
-0.022 |
0.764 |
-1.281 |
| C6 |
-0.022 |
-0.786 |
-1.244 |
| C7 |
-0.022 |
-0.786 |
1.244 |
| N8 |
-0.683 |
-1.254 |
0.000 |
| H9 |
-0.872 |
-2.256 |
0.000 |
| H10 |
0.507 |
1.114 |
2.180 |
| H11 |
0.507 |
1.114 |
-2.180 |
| H12 |
-1.063 |
1.112 |
1.351 |
| H13 |
-1.063 |
1.112 |
-1.351 |
| H14 |
1.028 |
-1.147 |
-1.321 |
| H15 |
1.028 |
-1.147 |
1.321 |
| H16 |
-0.573 |
-1.188 |
-2.106 |
| H17 |
-0.573 |
-1.188 |
2.106 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0999 | 1.1033 | 1.5553 | 1.5553 | 2.5557 | 2.5557 | 2.9170 | 3.9059 | 2.1968 | 2.1968 | 2.1860 | 2.1860 | 2.8477 | 2.8477 | 3.5104 | 3.5104 |
H2 | 1.0999 | | 1.7676 | 2.1947 | 2.1947 | 3.5117 | 3.5117 | 3.9046 | 4.9160 | 2.5550 | 2.5550 | 2.5039 | 2.5039 | 3.8529 | 3.8529 | 4.3509 | 4.3509 | H3 | 1.1033 | 1.7676 | | 2.1831 | 2.1831 | 2.8463 | 2.8463 | 3.3562 | 4.2325 | 2.4922 | 2.4922 | 3.0882 | 3.0882 | 2.6908 | 2.6908 | 3.8567 | 3.8567 | C4 | 1.5553 | 2.1947 | 2.1831 | | 2.5627 | 2.9633 | 1.5502 | 2.4799 | 3.3883 | 1.1004 | 3.5195 | 1.0997 | 2.8519 | 3.3951 | 2.1805 | 3.9476 | 2.1888 | C5 | 1.5553 | 2.1947 | 2.1831 | 2.5627 | | 1.5502 | 2.9633 | 2.4799 | 3.3883 | 3.5195 | 1.1004 | 2.8519 | 1.0997 | 2.1805 | 3.3951 | 2.1888 | 3.9476 | C6 | 2.5557 | 3.5117 | 2.8463 | 2.9633 | 1.5502 | | 2.4886 | 1.4846 | 2.1048 | 3.9519 | 2.1829 | 3.3794 | 2.1672 | 1.1135 | 2.7958 | 1.0983 | 3.4186 | C7 | 2.5557 | 3.5117 | 2.8463 | 1.5502 | 2.9633 | 2.4886 | | 1.4846 | 2.1048 | 2.1829 | 3.9519 | 2.1672 | 3.3794 | 2.7958 | 1.1135 | 3.4186 | 1.0983 | N8 | 2.9170 | 3.9046 | 3.3562 | 2.4799 | 2.4799 | 1.4846 | 1.4846 | | 1.0194 | 3.4316 | 3.4316 | 2.7508 | 2.7508 | 2.1649 | 2.1649 | 2.1096 | 2.1096 | H9 | 3.9059 | 4.9160 | 4.2325 | 3.3883 | 3.3883 | 2.1048 | 2.1048 | 1.0194 | | 4.2435 | 4.2435 | 3.6335 | 3.6335 | 2.5666 | 2.5666 | 2.3799 | 2.3799 | H10 | 2.1968 | 2.5550 | 2.4922 | 1.1004 | 3.5195 | 3.9519 | 2.1829 | 3.4316 | 4.2435 | | 4.3609 | 1.7759 | 3.8647 | 4.2002 | 2.4734 | 4.9832 | 2.5431 | H11 | 2.1968 | 2.5550 | 2.4922 | 3.5195 | 1.1004 | 2.1829 | 3.9519 | 3.4316 | 4.2435 | 4.3609 | | 3.8647 | 1.7759 | 2.4734 | 4.2002 | 2.5431 | 4.9832 | H12 | 2.1860 | 2.5039 | 3.0882 | 1.0997 | 2.8519 | 3.3794 | 2.1672 | 2.7508 | 3.6335 | 1.7759 | 3.8647 | | 2.7018 | 4.0762 | 3.0783 | 4.1805 | 2.4696 | H13 | 2.1860 | 2.5039 | 3.0882 | 2.8519 | 1.0997 | 2.1672 | 3.3794 | 2.7508 | 3.6335 | 3.8647 | 1.7759 | 2.7018 | | 3.0783 | 4.0762 | 2.4696 | 4.1805 | H14 | 2.8477 | 3.8529 | 2.6908 | 3.3951 | 2.1805 | 1.1135 | 2.7958 | 2.1649 | 2.5666 | 4.2002 | 2.4734 | 4.0762 | 3.0783 | | 2.6427 | 1.7834 | 3.7828 | H15 | 2.8477 | 3.8529 | 2.6908 | 2.1805 | 3.3951 | 2.7958 | 1.1135 | 2.1649 | 2.5666 | 2.4734 | 4.2002 | 3.0783 | 4.0762 | 2.6427 | | 3.7828 | 1.7834 | H16 | 3.5104 | 4.3509 | 3.8567 | 3.9476 | 2.1888 | 1.0983 | 3.4186 | 2.1096 | 2.3799 | 4.9832 | 2.5431 | 4.1805 | 2.4696 | 1.7834 | 3.7828 | | 4.2113 | H17 | 3.5104 | 4.3509 | 3.8567 | 2.1888 | 3.9476 | 3.4186 | 1.0983 | 2.1096 | 2.3799 | 2.5431 | 4.9832 | 2.4696 | 4.1805 | 3.7828 | 1.7834 | 4.2113 | |
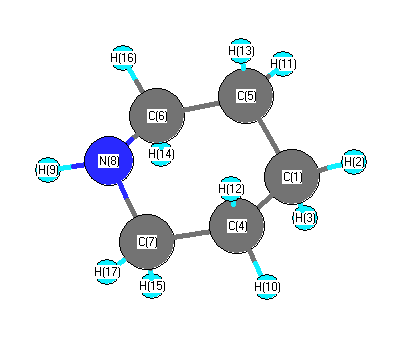 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.766 |
|
C1 |
C4 |
H10 |
110.454 |
| C1 |
C4 |
H12 |
109.651 |
|
C1 |
C5 |
C6 |
110.766 |
| C1 |
C5 |
H11 |
110.454 |
|
C1 |
C5 |
H13 |
109.651 |
| H2 |
C1 |
H3 |
106.702 |
|
H2 |
C1 |
C4 |
110.316 |
| H2 |
C1 |
C5 |
110.316 |
|
H3 |
C1 |
C4 |
109.225 |
| H3 |
C1 |
C5 |
109.225 |
|
C4 |
C1 |
C5 |
110.952 |
| C4 |
C7 |
N8 |
109.581 |
|
C4 |
C7 |
H15 |
108.782 |
| C4 |
C7 |
H17 |
110.300 |
|
C5 |
C6 |
N8 |
109.581 |
| C5 |
C6 |
H14 |
108.782 |
|
C5 |
C6 |
H16 |
110.300 |
| C6 |
C5 |
H11 |
109.713 |
|
C6 |
C5 |
H13 |
108.543 |
| C6 |
N8 |
C7 |
113.894 |
|
C6 |
N8 |
H9 |
113.088 |
| C7 |
C4 |
H10 |
109.713 |
|
C7 |
C4 |
H12 |
108.543 |
| C7 |
N8 |
H9 |
113.088 |
|
N8 |
C6 |
H14 |
112.084 |
| N8 |
C6 |
H16 |
108.601 |
|
N8 |
C7 |
H15 |
112.084 |
| N8 |
C7 |
H17 |
108.601 |
|
H10 |
C4 |
H12 |
107.638 |
| H11 |
C5 |
H13 |
107.638 |
|
H14 |
C6 |
H16 |
107.471 |
| H15 |
C7 |
H17 |
107.471 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.254 |
|
|
|
| 2 |
H |
0.137 |
|
|
|
| 3 |
H |
0.115 |
|
|
|
| 4 |
C |
-0.209 |
|
|
|
| 5 |
C |
-0.209 |
|
|
|
| 6 |
C |
-0.244 |
|
|
|
| 7 |
C |
-0.244 |
|
|
|
| 8 |
N |
-0.313 |
|
|
|
| 9 |
H |
0.255 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
H |
0.126 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
H |
0.099 |
|
|
|
| 15 |
H |
0.099 |
|
|
|
| 16 |
H |
0.137 |
|
|
|
| 17 |
H |
0.137 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.687 |
-0.593 |
0.000 |
0.907 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.958 |
0.225 |
0.000 |
| y |
0.225 |
-36.640 |
0.000 |
| z |
0.000 |
0.000 |
-38.410 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.433 |
0.225 |
0.000 |
| y |
0.225 |
3.543 |
0.000 |
| z |
0.000 |
0.000 |
0.890 |
|
| Polar |
|---|
| 3z2-r2 | 1.779 |
|---|
| x2-y2 | -5.318 |
|---|
| xy | 0.225 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.883 |
0.300 |
0.000 |
| y |
0.300 |
9.657 |
0.000 |
| z |
0.000 |
0.000 |
10.264 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -44.711768 |
| Energy at 298.15K | -44.725965 |
| Nuclear repulsion energy | 143.238124 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3481 |
3393 |
5.89 |
|
|
|
| 2 |
A' |
3050 |
2973 |
106.13 |
|
|
|
| 3 |
A' |
3031 |
2953 |
98.86 |
|
|
|
| 4 |
A' |
3021 |
2944 |
76.77 |
|
|
|
| 5 |
A' |
2977 |
2901 |
50.82 |
|
|
|
| 6 |
A' |
2967 |
2891 |
38.41 |
|
|
|
| 7 |
A' |
2963 |
2888 |
41.23 |
|
|
|
| 8 |
A' |
1521 |
1482 |
4.53 |
|
|
|
| 9 |
A' |
1508 |
1470 |
9.49 |
|
|
|
| 10 |
A' |
1503 |
1465 |
4.66 |
|
|
|
| 11 |
A' |
1394 |
1359 |
3.44 |
|
|
|
| 12 |
A' |
1374 |
1339 |
0.81 |
|
|
|
| 13 |
A' |
1337 |
1303 |
0.08 |
|
|
|
| 14 |
A' |
1274 |
1242 |
6.63 |
|
|
|
| 15 |
A' |
1187 |
1157 |
1.65 |
|
|
|
| 16 |
A' |
1044 |
1018 |
3.03 |
|
|
|
| 17 |
A' |
1011 |
985 |
6.09 |
|
|
|
| 18 |
A' |
894 |
871 |
2.51 |
|
|
|
| 19 |
A' |
855 |
833 |
18.06 |
|
|
|
| 20 |
A' |
801 |
781 |
4.18 |
|
|
|
| 21 |
A' |
626 |
610 |
181.86 |
|
|
|
| 22 |
A' |
550 |
536 |
12.39 |
|
|
|
| 23 |
A' |
429 |
418 |
3.67 |
|
|
|
| 24 |
A' |
368 |
358 |
12.09 |
|
|
|
| 25 |
A' |
228 |
222 |
8.48 |
|
|
|
| 26 |
A" |
3047 |
2969 |
49.72 |
|
|
|
| 27 |
A" |
3019 |
2942 |
107.15 |
|
|
|
| 28 |
A" |
2971 |
2896 |
23.64 |
|
|
|
| 29 |
A" |
2963 |
2888 |
54.17 |
|
|
|
| 30 |
A" |
1505 |
1467 |
0.06 |
|
|
|
| 31 |
A" |
1500 |
1462 |
7.65 |
|
|
|
| 32 |
A" |
1480 |
1442 |
9.79 |
|
|
|
| 33 |
A" |
1380 |
1345 |
3.82 |
|
|
|
| 34 |
A" |
1364 |
1330 |
0.87 |
|
|
|
| 35 |
A" |
1363 |
1328 |
2.18 |
|
|
|
| 36 |
A" |
1297 |
1264 |
0.61 |
|
|
|
| 37 |
A" |
1213 |
1182 |
18.42 |
|
|
|
| 38 |
A" |
1130 |
1102 |
3.42 |
|
|
|
| 39 |
A" |
1120 |
1091 |
16.68 |
|
|
|
| 40 |
A" |
1074 |
1046 |
4.83 |
|
|
|
| 41 |
A" |
955 |
931 |
4.76 |
|
|
|
| 42 |
A" |
864 |
842 |
0.35 |
|
|
|
| 43 |
A" |
810 |
790 |
0.16 |
|
|
|
| 44 |
A" |
449 |
438 |
0.85 |
|
|
|
| 45 |
A" |
224 |
218 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34561.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 33680.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.651 |
1.339 |
0.000 |
| H2 |
0.593 |
2.437 |
0.000 |
| H3 |
1.724 |
1.080 |
0.000 |
| C4 |
-0.011 |
0.760 |
1.284 |
| C5 |
-0.011 |
0.760 |
-1.284 |
| C6 |
-0.011 |
-0.798 |
-1.245 |
| C7 |
-0.011 |
-0.798 |
1.245 |
| N8 |
-0.600 |
-1.359 |
0.000 |
| H9 |
-1.621 |
-1.322 |
0.000 |
| H10 |
0.517 |
1.115 |
2.183 |
| H11 |
0.517 |
1.115 |
-2.183 |
| H12 |
-1.051 |
1.121 |
1.357 |
| H13 |
-1.051 |
1.121 |
-1.357 |
| H14 |
1.027 |
-1.160 |
-1.317 |
| H15 |
1.027 |
-1.160 |
1.317 |
| H16 |
-0.556 |
-1.211 |
-2.105 |
| H17 |
-0.556 |
-1.211 |
2.105 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0999 | 1.1034 | 1.5563 | 1.5563 | 2.5604 | 2.5604 | 2.9746 | 3.4990 | 2.1981 | 2.1981 | 2.1876 | 2.1876 | 2.8499 | 2.8499 | 3.5198 | 3.5198 |
H2 | 1.0999 | | 1.7667 | 2.1967 | 2.1967 | 3.5187 | 3.5187 | 3.9798 | 4.3626 | 2.5528 | 2.5528 | 2.5048 | 2.5048 | 3.8556 | 3.8556 | 4.3655 | 4.3655 | H3 | 1.1034 | 1.7667 | | 2.1822 | 2.1822 | 2.8441 | 2.8441 | 3.3696 | 4.1179 | 2.4945 | 2.4945 | 3.0891 | 3.0891 | 2.6908 | 2.6908 | 3.8572 | 3.8572 | C4 | 1.5563 | 2.1967 | 2.1822 | | 2.5674 | 2.9703 | 1.5585 | 2.5471 | 2.9280 | 1.1014 | 3.5242 | 1.1029 | 2.8604 | 3.3955 | 2.1835 | 3.9575 | 2.2035 | C5 | 1.5563 | 2.1967 | 2.1822 | 2.5674 | | 1.5585 | 2.9703 | 2.5471 | 2.9280 | 3.5242 | 1.1014 | 2.8604 | 1.1029 | 2.1835 | 3.3955 | 2.2035 | 3.9575 | C6 | 2.5604 | 3.5187 | 2.8441 | 2.9703 | 1.5585 | | 2.4904 | 1.4875 | 2.1011 | 3.9609 | 2.1949 | 3.3960 | 2.1854 | 1.1020 | 2.7882 | 1.0983 | 3.4189 | C7 | 2.5604 | 3.5187 | 2.8441 | 1.5585 | 2.9703 | 2.4904 | | 1.4875 | 2.1011 | 2.1949 | 3.9609 | 2.1854 | 3.3960 | 2.7882 | 1.1020 | 3.4189 | 1.0983 | N8 | 2.9746 | 3.9798 | 3.3696 | 2.5471 | 2.5471 | 1.4875 | 1.4875 | | 1.0210 | 3.4837 | 3.4837 | 2.8632 | 2.8632 | 2.1027 | 2.1027 | 2.1104 | 2.1104 | H9 | 3.4990 | 4.3626 | 4.1179 | 2.9280 | 2.9280 | 2.1011 | 2.1011 | 1.0210 | | 3.9080 | 3.9080 | 2.8522 | 2.8522 | 2.9612 | 2.9612 | 2.3610 | 2.3610 | H10 | 2.1981 | 2.5528 | 2.4945 | 1.1014 | 3.5242 | 3.9609 | 2.1949 | 3.4837 | 3.9080 | | 4.3651 | 1.7720 | 3.8708 | 4.2054 | 2.4876 | 4.9942 | 2.5628 | H11 | 2.1981 | 2.5528 | 2.4945 | 3.5242 | 1.1014 | 2.1949 | 3.9609 | 3.4837 | 3.9080 | 4.3651 | | 3.8708 | 1.7720 | 2.4876 | 4.2054 | 2.5628 | 4.9942 | H12 | 2.1876 | 2.5048 | 3.0891 | 1.1029 | 2.8604 | 3.3960 | 2.1854 | 2.8632 | 2.8522 | 1.7720 | 3.8708 | | 2.7131 | 4.0830 | 3.0862 | 4.2027 | 2.4985 | H13 | 2.1876 | 2.5048 | 3.0891 | 2.8604 | 1.1029 | 2.1854 | 3.3960 | 2.8632 | 2.8522 | 3.8708 | 1.7720 | 2.7131 | | 3.0862 | 4.0830 | 2.4985 | 4.2027 | H14 | 2.8499 | 3.8556 | 2.6908 | 3.3955 | 2.1835 | 1.1020 | 2.7882 | 2.1027 | 2.9612 | 4.2054 | 2.4876 | 4.0830 | 3.0862 | | 2.6339 | 1.7690 | 3.7705 | H15 | 2.8499 | 3.8556 | 2.6908 | 2.1835 | 3.3955 | 2.7882 | 1.1020 | 2.1027 | 2.9612 | 2.4876 | 4.2054 | 3.0862 | 4.0830 | 2.6339 | | 3.7705 | 1.7690 | H16 | 3.5198 | 4.3655 | 3.8572 | 3.9575 | 2.2035 | 1.0983 | 3.4189 | 2.1104 | 2.3610 | 4.9942 | 2.5628 | 4.2027 | 2.4985 | 1.7690 | 3.7705 | | 4.2093 | H17 | 3.5198 | 4.3655 | 3.8572 | 2.2035 | 3.9575 | 3.4189 | 1.0983 | 2.1104 | 2.3610 | 2.5628 | 4.9942 | 2.4985 | 4.2027 | 3.7705 | 1.7690 | 4.2093 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.571 |
|
C1 |
C4 |
H10 |
110.427 |
| C1 |
C4 |
H12 |
109.522 |
|
C1 |
C5 |
C6 |
110.571 |
| C1 |
C5 |
H11 |
110.427 |
|
C1 |
C5 |
H13 |
109.522 |
| H2 |
C1 |
H3 |
106.615 |
|
H2 |
C1 |
C4 |
110.405 |
| H2 |
C1 |
C5 |
110.405 |
|
H3 |
C1 |
C4 |
109.076 |
| H3 |
C1 |
C5 |
109.076 |
|
C4 |
C1 |
C5 |
111.139 |
| C4 |
C7 |
N8 |
113.464 |
|
C4 |
C7 |
H15 |
109.105 |
| C4 |
C7 |
H17 |
110.886 |
|
C5 |
C6 |
N8 |
113.464 |
| C5 |
C6 |
H14 |
109.105 |
|
C5 |
C6 |
H16 |
110.886 |
| C6 |
C5 |
H11 |
110.029 |
|
C6 |
C5 |
H13 |
109.206 |
| C6 |
N8 |
C7 |
113.669 |
|
C6 |
N8 |
H9 |
112.442 |
| C7 |
C4 |
H10 |
110.029 |
|
C7 |
C4 |
H12 |
109.206 |
| C7 |
N8 |
H9 |
112.442 |
|
N8 |
C6 |
H14 |
107.650 |
| N8 |
C6 |
H16 |
108.462 |
|
N8 |
C7 |
H15 |
107.650 |
| N8 |
C7 |
H17 |
108.462 |
|
H10 |
C4 |
H12 |
107.007 |
| H11 |
C5 |
H13 |
107.007 |
|
H14 |
C6 |
H16 |
107.028 |
| H15 |
C7 |
H17 |
107.028 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.240 |
|
|
|
| 2 |
H |
0.137 |
|
|
|
| 3 |
H |
0.121 |
|
|
|
| 4 |
C |
-0.230 |
|
|
|
| 5 |
C |
-0.230 |
|
|
|
| 6 |
C |
-0.280 |
|
|
|
| 7 |
C |
-0.280 |
|
|
|
| 8 |
N |
-0.266 |
|
|
|
| 9 |
H |
0.242 |
|
|
|
| 10 |
H |
0.125 |
|
|
|
| 11 |
H |
0.125 |
|
|
|
| 12 |
H |
0.110 |
|
|
|
| 13 |
H |
0.110 |
|
|
|
| 14 |
H |
0.134 |
|
|
|
| 15 |
H |
0.134 |
|
|
|
| 16 |
H |
0.143 |
|
|
|
| 17 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.433 |
1.196 |
0.000 |
1.272 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.194 |
0.292 |
0.000 |
| y |
0.292 |
-44.527 |
0.000 |
| z |
0.000 |
0.000 |
-38.306 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.223 |
0.292 |
0.000 |
| y |
0.292 |
-6.777 |
0.000 |
| z |
0.000 |
0.000 |
2.554 |
|
| Polar |
|---|
| 3z2-r2 | 5.109 |
|---|
| x2-y2 | 7.333 |
|---|
| xy | 0.292 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.033 |
0.291 |
0.000 |
| y |
0.291 |
9.266 |
0.000 |
| z |
0.000 |
0.000 |
10.252 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
