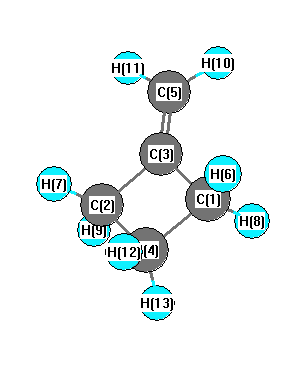
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3098 |
3019 |
84.69 |
|
|
|
| 2 |
A' |
3094 |
3015 |
14.79 |
|
|
|
| 3 |
A' |
3057 |
2979 |
28.91 |
|
|
|
| 4 |
A' |
3030 |
2953 |
62.89 |
|
|
|
| 5 |
A' |
3004 |
2927 |
10.49 |
|
|
|
| 6 |
A' |
1714 |
1670 |
26.65 |
|
|
|
| 7 |
A' |
1514 |
1476 |
0.26 |
|
|
|
| 8 |
A' |
1493 |
1455 |
1.35 |
|
|
|
| 9 |
A' |
1450 |
1413 |
1.23 |
|
|
|
| 10 |
A' |
1269 |
1237 |
0.20 |
|
|
|
| 11 |
A' |
1219 |
1188 |
0.10 |
|
|
|
| 12 |
A' |
1102 |
1074 |
0.02 |
|
|
|
| 13 |
A' |
956 |
932 |
1.26 |
|
|
|
| 14 |
A' |
927 |
904 |
62.70 |
|
|
|
| 15 |
A' |
870 |
848 |
0.04 |
|
|
|
| 16 |
A' |
747 |
728 |
2.20 |
|
|
|
| 17 |
A' |
665 |
648 |
1.92 |
|
|
|
| 18 |
A' |
377 |
368 |
7.92 |
|
|
|
| 19 |
A' |
76 |
74 |
0.01 |
|
|
|
| 20 |
A" |
3189 |
3108 |
28.68 |
|
|
|
| 21 |
A" |
3062 |
2984 |
12.40 |
|
|
|
| 22 |
A" |
3000 |
2924 |
83.67 |
|
|
|
| 23 |
A" |
1483 |
1445 |
6.28 |
|
|
|
| 24 |
A" |
1287 |
1254 |
0.00 |
|
|
|
| 25 |
A" |
1251 |
1219 |
1.08 |
|
|
|
| 26 |
A" |
1228 |
1196 |
1.64 |
|
|
|
| 27 |
A" |
1196 |
1166 |
4.22 |
|
|
|
| 28 |
A" |
1049 |
1022 |
0.08 |
|
|
|
| 29 |
A" |
931 |
907 |
1.51 |
|
|
|
| 30 |
A" |
902 |
879 |
2.63 |
|
|
|
| 31 |
A" |
817 |
796 |
0.03 |
|
|
|
| 32 |
A" |
644 |
627 |
0.07 |
|
|
|
| 33 |
A" |
351 |
342 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25024.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 24386.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.410 |
|
|
|
| 2 |
C |
-0.410 |
|
|
|
| 3 |
C |
0.507 |
|
|
|
| 4 |
C |
-0.241 |
|
|
|
| 5 |
C |
-0.546 |
|
|
|
| 6 |
H |
0.147 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.127 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.064 |
-0.606 |
0.000 |
0.610 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.686 |
-0.439 |
0.000 |
| y |
-0.439 |
-32.325 |
0.000 |
| z |
0.000 |
0.000 |
-30.958 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.044 |
-0.439 |
0.000 |
| y |
-0.439 |
-0.503 |
0.000 |
| z |
0.000 |
0.000 |
1.548 |
|
| Polar |
|---|
| 3z2-r2 | 3.095 |
|---|
| x2-y2 | -0.361 |
|---|
| xy | -0.439 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.819 |
-0.001 |
-0.336 |
| y |
-0.001 |
8.078 |
0.004 |
| z |
-0.336 |
0.004 |
6.147 |
<r2> (average value of r
2) Å
2
| <r2> |
100.393 |
| (<r2>)1/2 |
10.020 |