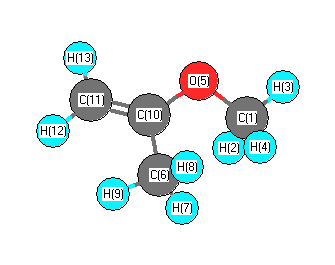
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3242 |
3159 |
17.87 |
|
|
|
| 2 |
A |
3134 |
3054 |
2.87 |
|
|
|
| 3 |
A |
3126 |
3046 |
29.46 |
|
|
|
| 4 |
A |
3106 |
3026 |
29.37 |
|
|
|
| 5 |
A |
3058 |
2980 |
76.68 |
|
|
|
| 6 |
A |
3056 |
2978 |
20.22 |
|
|
|
| 7 |
A |
2989 |
2913 |
42.91 |
|
|
|
| 8 |
A |
2972 |
2896 |
58.12 |
|
|
|
| 9 |
A |
1687 |
1644 |
107.41 |
|
|
|
| 10 |
A |
1519 |
1480 |
0.34 |
|
|
|
| 11 |
A |
1512 |
1474 |
21.24 |
|
|
|
| 12 |
A |
1503 |
1464 |
18.65 |
|
|
|
| 13 |
A |
1500 |
1462 |
8.71 |
|
|
|
| 14 |
A |
1478 |
1440 |
0.17 |
|
|
|
| 15 |
A |
1443 |
1406 |
10.42 |
|
|
|
| 16 |
A |
1424 |
1387 |
9.01 |
|
|
|
| 17 |
A |
1274 |
1241 |
166.65 |
|
|
|
| 18 |
A |
1185 |
1155 |
4.20 |
|
|
|
| 19 |
A |
1140 |
1111 |
4.09 |
|
|
|
| 20 |
A |
1080 |
1053 |
1.27 |
|
|
|
| 21 |
A |
1068 |
1041 |
109.60 |
|
|
|
| 22 |
A |
1022 |
996 |
3.05 |
|
|
|
| 23 |
A |
917 |
894 |
5.66 |
|
|
|
| 24 |
A |
890 |
867 |
78.71 |
|
|
|
| 25 |
A |
764 |
744 |
10.04 |
|
|
|
| 26 |
A |
730 |
712 |
5.46 |
|
|
|
| 27 |
A |
521 |
508 |
7.87 |
|
|
|
| 28 |
A |
459 |
448 |
4.15 |
|
|
|
| 29 |
A |
410 |
400 |
1.09 |
|
|
|
| 30 |
A |
286 |
279 |
2.35 |
|
|
|
| 31 |
A |
224 |
218 |
1.99 |
|
|
|
| 32 |
A |
182 |
178 |
1.48 |
|
|
|
| 33 |
A |
77 |
75 |
10.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24488.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 23863.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.311 |
|
|
|
| 2 |
H |
0.151 |
|
|
|
| 3 |
H |
0.176 |
|
|
|
| 4 |
H |
0.149 |
|
|
|
| 5 |
O |
-0.246 |
|
|
|
| 6 |
C |
-0.670 |
|
|
|
| 7 |
H |
0.167 |
|
|
|
| 8 |
H |
0.174 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
C |
0.435 |
|
|
|
| 11 |
C |
-0.471 |
|
|
|
| 12 |
H |
0.145 |
|
|
|
| 13 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.846 |
1.977 |
0.784 |
2.288 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.771 |
0.113 |
0.612 |
| y |
0.113 |
-32.151 |
-0.797 |
| z |
0.612 |
-0.797 |
-34.048 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.328 |
0.113 |
0.612 |
| y |
0.113 |
-1.242 |
-0.797 |
| z |
0.612 |
-0.797 |
-4.086 |
|
| Polar |
|---|
| 3z2-r2 | -8.173 |
|---|
| x2-y2 | 4.380 |
|---|
| xy | 0.113 |
|---|
| xz | 0.612 |
|---|
| yz | -0.797 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.932 |
1.216 |
-0.244 |
| y |
1.216 |
7.887 |
-0.140 |
| z |
-0.244 |
-0.140 |
5.408 |
<r2> (average value of r
2) Å
2
| <r2> |
111.895 |
| (<r2>)1/2 |
10.578 |