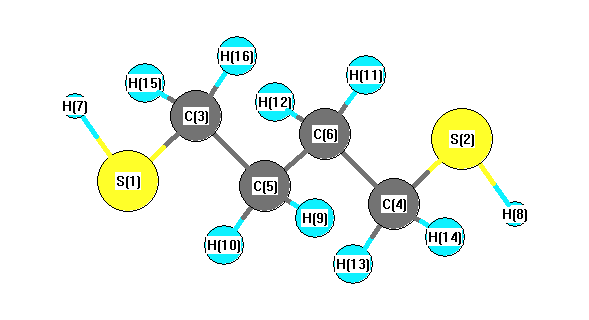
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3029 |
2951 |
0.00 |
|
|
|
| 2 |
Ag |
2992 |
2915 |
0.00 |
|
|
|
| 3 |
Ag |
2476 |
2413 |
0.00 |
|
|
|
| 4 |
Ag |
1519 |
1481 |
0.00 |
|
|
|
| 5 |
Ag |
1500 |
1461 |
0.00 |
|
|
|
| 6 |
Ag |
1394 |
1359 |
0.00 |
|
|
|
| 7 |
Ag |
1283 |
1250 |
0.00 |
|
|
|
| 8 |
Ag |
1096 |
1068 |
0.00 |
|
|
|
| 9 |
Ag |
1046 |
1019 |
0.00 |
|
|
|
| 10 |
Ag |
808 |
787 |
0.00 |
|
|
|
| 11 |
Ag |
709 |
691 |
0.00 |
|
|
|
| 12 |
Ag |
324 |
316 |
0.00 |
|
|
|
| 13 |
Ag |
195 |
190 |
0.00 |
|
|
|
| 14 |
Au |
3107 |
3028 |
77.10 |
|
|
|
| 15 |
Au |
3054 |
2976 |
25.79 |
|
|
|
| 16 |
Au |
1320 |
1286 |
0.80 |
|
|
|
| 17 |
Au |
1101 |
1073 |
3.56 |
|
|
|
| 18 |
Au |
900 |
877 |
2.64 |
|
|
|
| 19 |
Au |
757 |
738 |
7.26 |
|
|
|
| 20 |
Au |
150 |
146 |
36.78 |
|
|
|
| 21 |
Au |
85 |
83 |
18.36 |
|
|
|
| 22 |
Au |
49 |
48 |
8.81 |
|
|
|
| 23 |
Bg |
3102 |
3023 |
0.00 |
|
|
|
| 24 |
Bg |
3031 |
2954 |
0.00 |
|
|
|
| 25 |
Bg |
1335 |
1301 |
0.00 |
|
|
|
| 26 |
Bg |
1256 |
1224 |
0.00 |
|
|
|
| 27 |
Bg |
1040 |
1014 |
0.00 |
|
|
|
| 28 |
Bg |
791 |
771 |
0.00 |
|
|
|
| 29 |
Bg |
147 |
143 |
0.00 |
|
|
|
| 30 |
Bg |
124 |
121 |
0.00 |
|
|
|
| 31 |
Bu |
3030 |
2953 |
87.94 |
|
|
|
| 32 |
Bu |
2997 |
2921 |
30.63 |
|
|
|
| 33 |
Bu |
2476 |
2413 |
80.98 |
|
|
|
| 34 |
Bu |
1536 |
1497 |
7.52 |
|
|
|
| 35 |
Bu |
1497 |
1459 |
3.03 |
|
|
|
| 36 |
Bu |
1353 |
1319 |
54.29 |
|
|
|
| 37 |
Bu |
1248 |
1216 |
43.97 |
|
|
|
| 38 |
Bu |
1044 |
1018 |
2.91 |
|
|
|
| 39 |
Bu |
837 |
816 |
5.14 |
|
|
|
| 40 |
Bu |
673 |
656 |
11.55 |
|
|
|
| 41 |
Bu |
371 |
362 |
10.38 |
|
|
|
| 42 |
Bu |
96 |
94 |
4.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28439.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 27714.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.057 |
|
|
|
| 2 |
S |
0.057 |
|
|
|
| 3 |
C |
-0.564 |
|
|
|
| 4 |
C |
-0.564 |
|
|
|
| 5 |
C |
-0.194 |
|
|
|
| 6 |
C |
-0.194 |
|
|
|
| 7 |
H |
0.045 |
|
|
|
| 8 |
H |
0.045 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.147 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
| 13 |
H |
0.182 |
|
|
|
| 14 |
H |
0.182 |
|
|
|
| 15 |
H |
0.182 |
|
|
|
| 16 |
H |
0.182 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.295 |
4.226 |
0.000 |
| y |
4.226 |
-67.675 |
0.000 |
| z |
0.000 |
0.000 |
-55.173 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
18.129 |
4.226 |
0.000 |
| y |
4.226 |
-18.441 |
0.000 |
| z |
0.000 |
0.000 |
0.312 |
|
| Polar |
|---|
| 3z2-r2 | 0.624 |
|---|
| x2-y2 | 24.380 |
|---|
| xy | 4.226 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.042 |
2.523 |
0.000 |
| y |
2.523 |
16.805 |
0.000 |
| z |
0.000 |
0.000 |
8.835 |
<r2> (average value of r
2) Å
2
| <r2> |
292.236 |
| (<r2>)1/2 |
17.095 |