Jump to
S2C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -22.234219 |
| Energy at 298.15K | |
| HF Energy | -22.234219 |
| Nuclear repulsion energy | 27.584950 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
2053 |
2001 |
0.00 |
92.12 |
0.57 |
0.72 |
| 2 |
Σg |
882 |
859 |
0.00 |
112.64 |
0.41 |
0.58 |
| 3 |
Σu |
1466 |
1429 |
304.27 |
0.00 |
0.00 |
0.00 |
| 4 |
Πg |
569 |
555 |
0.00 |
4.74 |
0.75 |
0.86 |
| 4 |
Πg |
569 |
555 |
0.00 |
4.74 |
0.75 |
0.86 |
| 5 |
Πu |
227 |
221 |
19.44 |
0.00 |
0.00 |
0.00 |
| 5 |
Πu |
227 |
221 |
19.44 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 2996.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 2919.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.653 |
| C2 |
0.000 |
0.000 |
-0.653 |
| C3 |
0.000 |
0.000 |
1.997 |
| C4 |
0.000 |
0.000 |
-1.997 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
| C1 | | 1.3068 | 1.3441 | 2.6509 |
C2 | 1.3068 | | 2.6509 | 1.3441 | C3 | 1.3441 | 2.6509 | | 3.9950 | C4 | 2.6509 | 1.3441 | 3.9950 | |
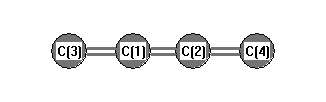 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
180.000 |
|
C2 |
C1 |
C3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.374 |
|
|
|
| 2 |
C |
0.374 |
|
|
|
| 3 |
C |
-0.374 |
|
|
|
| 4 |
C |
-0.374 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.983 |
0.000 |
0.000 |
| y |
0.000 |
-21.983 |
0.000 |
| z |
0.000 |
0.000 |
-34.138 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.077 |
0.000 |
0.000 |
| y |
0.000 |
6.077 |
0.000 |
| z |
0.000 |
0.000 |
-12.154 |
|
| Polar |
|---|
| 3z2-r2 | -24.309 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.576 |
0.000 |
0.000 |
| y |
0.000 |
3.576 |
0.000 |
| z |
0.000 |
0.000 |
14.158 |
<r2> (average value of r
2) Å
2
| <r2> |
51.596 |
| (<r2>)1/2 |
7.183 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -22.207222 |
| Energy at 298.15K | |
| HF Energy | -22.207222 |
| Nuclear repulsion energy | 27.552408 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
2060 |
2008 |
0.00 |
90.58 |
0.39 |
0.56 |
| 2 |
Σg |
881 |
859 |
0.00 |
93.42 |
0.32 |
0.48 |
| 3 |
Σu |
1486 |
1448 |
394.54 |
0.00 |
0.00 |
0.00 |
| 4 |
Πg |
627 |
611 |
0.00 |
11.36 |
0.75 |
0.86 |
| 4 |
Πg |
566 |
552 |
0.00 |
3.60 |
0.75 |
0.86 |
| 5 |
Πu |
238 |
232 |
29.42 |
0.00 |
0.00 |
0.00 |
| 5 |
Πu |
236 |
230 |
3.86 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 3047.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 2969.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.656 |
| C2 |
0.000 |
0.000 |
-0.656 |
| C3 |
0.000 |
0.000 |
2.000 |
| C4 |
0.000 |
0.000 |
-2.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
| C1 | | 1.3112 | 1.3439 | 2.6551 |
C2 | 1.3112 | | 2.6551 | 1.3439 | C3 | 1.3439 | 2.6551 | | 3.9990 | C4 | 2.6551 | 1.3439 | 3.9990 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.380 |
|
|
|
| 2 |
C |
0.380 |
|
|
|
| 3 |
C |
-0.380 |
|
|
|
| 4 |
C |
-0.380 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.116 |
0.000 |
0.000 |
| y |
0.000 |
-24.021 |
0.000 |
| z |
0.000 |
0.000 |
-33.825 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.807 |
0.000 |
0.000 |
| y |
0.000 |
2.950 |
0.000 |
| z |
0.000 |
0.000 |
-11.757 |
|
| Polar |
|---|
| 3z2-r2 | -23.514 |
|---|
| x2-y2 | 3.905 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.303 |
0.000 |
0.000 |
| y |
0.000 |
3.203 |
0.000 |
| z |
0.000 |
0.000 |
13.485 |
<r2> (average value of r
2) Å
2
| <r2> |
51.655 |
| (<r2>)1/2 |
7.187 |
