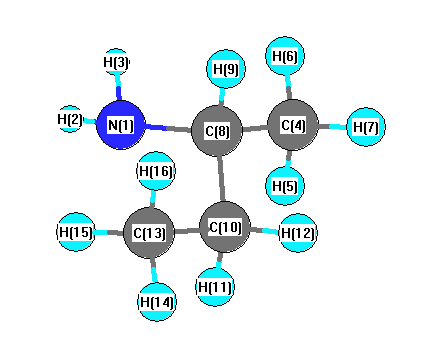
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3606 |
3514 |
1.12 |
|
|
|
| 2 |
A |
3482 |
3394 |
4.49 |
|
|
|
| 3 |
A |
3082 |
3004 |
72.71 |
|
|
|
| 4 |
A |
3066 |
2988 |
76.52 |
|
|
|
| 5 |
A |
3062 |
2984 |
81.28 |
|
|
|
| 6 |
A |
3060 |
2982 |
34.90 |
|
|
|
| 7 |
A |
3031 |
2954 |
5.86 |
|
|
|
| 8 |
A |
2984 |
2908 |
31.90 |
|
|
|
| 9 |
A |
2977 |
2901 |
37.83 |
|
|
|
| 10 |
A |
2975 |
2899 |
58.42 |
|
|
|
| 11 |
A |
2860 |
2787 |
107.60 |
|
|
|
| 12 |
A |
1691 |
1648 |
52.60 |
|
|
|
| 13 |
A |
1532 |
1493 |
9.74 |
|
|
|
| 14 |
A |
1520 |
1482 |
6.87 |
|
|
|
| 15 |
A |
1519 |
1480 |
4.13 |
|
|
|
| 16 |
A |
1511 |
1473 |
5.54 |
|
|
|
| 17 |
A |
1497 |
1459 |
1.00 |
|
|
|
| 18 |
A |
1424 |
1388 |
17.19 |
|
|
|
| 19 |
A |
1423 |
1386 |
2.23 |
|
|
|
| 20 |
A |
1406 |
1371 |
4.13 |
|
|
|
| 21 |
A |
1382 |
1347 |
27.17 |
|
|
|
| 22 |
A |
1341 |
1307 |
4.64 |
|
|
|
| 23 |
A |
1297 |
1264 |
2.40 |
|
|
|
| 24 |
A |
1243 |
1211 |
2.43 |
|
|
|
| 25 |
A |
1194 |
1164 |
4.43 |
|
|
|
| 26 |
A |
1162 |
1133 |
4.40 |
|
|
|
| 27 |
A |
1057 |
1031 |
5.38 |
|
|
|
| 28 |
A |
1029 |
1003 |
7.71 |
|
|
|
| 29 |
A |
1003 |
978 |
1.29 |
|
|
|
| 30 |
A |
989 |
964 |
4.20 |
|
|
|
| 31 |
A |
929 |
906 |
5.45 |
|
|
|
| 32 |
A |
820 |
799 |
7.01 |
|
|
|
| 33 |
A |
771 |
751 |
1.79 |
|
|
|
| 34 |
A |
648 |
631 |
190.63 |
|
|
|
| 35 |
A |
473 |
461 |
13.44 |
|
|
|
| 36 |
A |
446 |
434 |
7.25 |
|
|
|
| 37 |
A |
366 |
357 |
1.19 |
|
|
|
| 38 |
A |
258 |
251 |
23.95 |
|
|
|
| 39 |
A |
248 |
242 |
0.13 |
|
|
|
| 40 |
A |
218 |
213 |
0.92 |
|
|
|
| 41 |
A |
196 |
191 |
33.01 |
|
|
|
| 42 |
A |
96 |
93 |
2.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32437.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 31609.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.617 |
|
|
|
| 2 |
H |
0.261 |
|
|
|
| 3 |
H |
0.259 |
|
|
|
| 4 |
C |
-0.518 |
|
|
|
| 5 |
H |
0.154 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
H |
0.137 |
|
|
|
| 8 |
C |
0.117 |
|
|
|
| 9 |
H |
0.096 |
|
|
|
| 10 |
C |
-0.188 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.113 |
|
|
|
| 13 |
C |
-0.508 |
|
|
|
| 14 |
H |
0.149 |
|
|
|
| 15 |
H |
0.142 |
|
|
|
| 16 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.047 |
0.261 |
1.155 |
1.185 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.273 |
0.510 |
0.891 |
| y |
0.510 |
-32.296 |
1.701 |
| z |
0.891 |
1.701 |
-36.483 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.117 |
0.510 |
0.891 |
| y |
0.510 |
2.582 |
1.701 |
| z |
0.891 |
1.701 |
-3.699 |
|
| Polar |
|---|
| 3z2-r2 | -7.398 |
|---|
| x2-y2 | -0.977 |
|---|
| xy | 0.510 |
|---|
| xz | 0.891 |
|---|
| yz | 1.701 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.366 |
0.005 |
0.023 |
| y |
0.005 |
8.377 |
-0.084 |
| z |
0.023 |
-0.084 |
7.530 |
<r2> (average value of r
2) Å
2
| <r2> |
130.698 |
| (<r2>)1/2 |
11.432 |