Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -41.331774 |
| Energy at 298.15K | -41.340828 |
| HF Energy | -41.331774 |
| Nuclear repulsion energy | 78.359212 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3622 |
3530 |
0.91 |
|
|
|
| 2 |
A |
3526 |
3436 |
33.13 |
|
|
|
| 3 |
A |
3496 |
3407 |
0.67 |
|
|
|
| 4 |
A |
3079 |
3001 |
62.35 |
|
|
|
| 5 |
A |
3059 |
2981 |
39.31 |
|
|
|
| 6 |
A |
2958 |
2883 |
123.63 |
|
|
|
| 7 |
A |
2943 |
2868 |
65.44 |
|
|
|
| 8 |
A |
1693 |
1650 |
39.83 |
|
|
|
| 9 |
A |
1524 |
1486 |
1.13 |
|
|
|
| 10 |
A |
1508 |
1470 |
6.26 |
|
|
|
| 11 |
A |
1423 |
1387 |
62.33 |
|
|
|
| 12 |
A |
1403 |
1367 |
6.50 |
|
|
|
| 13 |
A |
1353 |
1319 |
14.36 |
|
|
|
| 14 |
A |
1313 |
1280 |
4.26 |
|
|
|
| 15 |
A |
1231 |
1199 |
26.31 |
|
|
|
| 16 |
A |
1169 |
1139 |
15.53 |
|
|
|
| 17 |
A |
1096 |
1068 |
22.25 |
|
|
|
| 18 |
A |
1033 |
1007 |
67.99 |
|
|
|
| 19 |
A |
984 |
959 |
2.77 |
|
|
|
| 20 |
A |
876 |
854 |
30.26 |
|
|
|
| 21 |
A |
871 |
849 |
17.13 |
|
|
|
| 22 |
A |
703 |
685 |
247.74 |
|
|
|
| 23 |
A |
621 |
605 |
136.10 |
|
|
|
| 24 |
A |
529 |
515 |
16.93 |
|
|
|
| 25 |
A |
315 |
307 |
0.28 |
|
|
|
| 26 |
A |
257 |
250 |
9.57 |
|
|
|
| 27 |
A |
167 |
162 |
7.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21375.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 20830.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.372 |
-0.547 |
0.131 |
| C2 |
-0.640 |
0.688 |
-0.265 |
| C3 |
0.814 |
0.588 |
0.257 |
| O4 |
1.394 |
-0.684 |
-0.162 |
| H5 |
-1.688 |
-0.565 |
1.098 |
| H6 |
-2.101 |
-0.842 |
-0.511 |
| H7 |
-1.115 |
1.617 |
0.095 |
| H8 |
-0.619 |
0.719 |
-1.362 |
| H9 |
1.440 |
1.385 |
-0.159 |
| H10 |
0.827 |
0.673 |
1.359 |
| H11 |
0.658 |
-1.339 |
-0.092 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4892 | 2.4664 | 2.7851 | 1.0176 | 1.0154 | 2.1801 | 2.0977 | 3.4239 | 2.7987 | 2.1898 |
C2 | 1.4892 | | 1.5482 | 2.4558 | 2.1277 | 2.1298 | 1.1044 | 1.0972 | 2.1959 | 2.1888 | 2.4126 | C3 | 2.4664 | 1.5482 | | 1.4601 | 2.8806 | 3.3368 | 2.1924 | 2.1662 | 1.0956 | 1.1050 | 1.9649 | O4 | 2.7851 | 2.4558 | 1.4601 | | 3.3324 | 3.5163 | 3.4147 | 2.7318 | 2.0698 | 2.1162 | 0.9883 | H5 | 1.0176 | 2.1277 | 2.8806 | 3.3324 | | 1.6841 | 2.4695 | 2.9740 | 3.8946 | 2.8157 | 2.7420 | H6 | 1.0154 | 2.1298 | 3.3368 | 3.5163 | 1.6841 | | 2.7184 | 2.3151 | 4.1978 | 3.7904 | 2.8340 | H7 | 2.1801 | 1.1044 | 2.1924 | 3.4147 | 2.4695 | 2.7184 | | 1.7816 | 2.5776 | 2.5023 | 3.4521 | H8 | 2.0977 | 1.0972 | 2.1662 | 2.7318 | 2.9740 | 2.3151 | 1.7816 | | 2.4751 | 3.0816 | 2.7347 | H9 | 3.4239 | 2.1959 | 1.0956 | 2.0698 | 3.8946 | 4.1978 | 2.5776 | 2.4751 | | 1.7855 | 2.8350 | H10 | 2.7987 | 2.1888 | 1.1050 | 2.1162 | 2.8157 | 3.7904 | 2.5023 | 3.0816 | 1.7855 | | 2.4867 | H11 | 2.1898 | 2.4126 | 1.9649 | 0.9883 | 2.7420 | 2.8340 | 3.4521 | 2.7347 | 2.8350 | 2.4867 | |
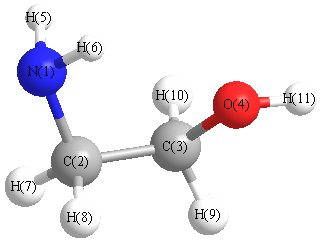 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.569 |
|
N1 |
C2 |
H7 |
113.572 |
| N1 |
C2 |
H8 |
107.428 |
|
C2 |
N1 |
H5 |
114.848 |
| C2 |
N1 |
H6 |
115.190 |
|
C2 |
C3 |
O4 |
109.405 |
| C2 |
C3 |
H9 |
111.164 |
|
C2 |
C3 |
H10 |
110.049 |
| C3 |
C2 |
H7 |
110.358 |
|
C3 |
C2 |
H8 |
108.742 |
| C3 |
O4 |
H11 |
105.111 |
|
O4 |
C3 |
H9 |
107.312 |
| O4 |
C3 |
H10 |
110.413 |
|
H5 |
N1 |
H6 |
111.874 |
| H7 |
C2 |
H8 |
108.036 |
|
H9 |
C3 |
H10 |
108.460 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.620 |
|
|
|
| 2 |
C |
-0.274 |
|
|
|
| 3 |
C |
-0.078 |
|
|
|
| 4 |
O |
-0.467 |
|
|
|
| 5 |
H |
0.276 |
|
|
|
| 6 |
H |
0.286 |
|
|
|
| 7 |
H |
0.127 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
H |
0.143 |
|
|
|
| 10 |
H |
0.112 |
|
|
|
| 11 |
H |
0.339 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.537 |
1.209 |
0.594 |
3.785 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.733 |
3.419 |
0.678 |
| y |
3.419 |
-25.292 |
-0.267 |
| z |
0.678 |
-0.267 |
-24.529 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.823 |
3.419 |
0.678 |
| y |
3.419 |
0.840 |
-0.267 |
| z |
0.678 |
-0.267 |
1.983 |
|
| Polar |
|---|
| 3z2-r2 | 3.967 |
|---|
| x2-y2 | -2.442 |
|---|
| xy | 3.419 |
|---|
| xz | 0.678 |
|---|
| yz | -0.267 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.595 |
0.212 |
-0.001 |
| y |
0.212 |
5.556 |
0.127 |
| z |
-0.001 |
0.127 |
4.951 |
<r2> (average value of r
2) Å
2
| <r2> |
75.500 |
| (<r2>)1/2 |
8.689 |
