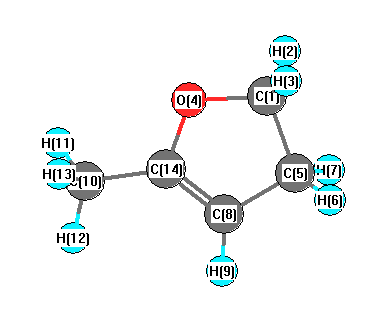
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3212 |
3130 |
8.98 |
|
|
|
| 2 |
A |
3107 |
3028 |
26.10 |
|
|
|
| 3 |
A |
3101 |
3022 |
55.07 |
|
|
|
| 4 |
A |
3069 |
2991 |
24.99 |
|
|
|
| 5 |
A |
3036 |
2958 |
73.38 |
|
|
|
| 6 |
A |
3021 |
2944 |
30.82 |
|
|
|
| 7 |
A |
2998 |
2922 |
35.56 |
|
|
|
| 8 |
A |
2984 |
2908 |
70.48 |
|
|
|
| 9 |
A |
1698 |
1655 |
47.98 |
|
|
|
| 10 |
A |
1533 |
1494 |
0.79 |
|
|
|
| 11 |
A |
1517 |
1479 |
1.02 |
|
|
|
| 12 |
A |
1510 |
1472 |
9.55 |
|
|
|
| 13 |
A |
1494 |
1456 |
10.87 |
|
|
|
| 14 |
A |
1439 |
1402 |
10.20 |
|
|
|
| 15 |
A |
1381 |
1346 |
5.22 |
|
|
|
| 16 |
A |
1318 |
1285 |
5.61 |
|
|
|
| 17 |
A |
1262 |
1229 |
37.39 |
|
|
|
| 18 |
A |
1238 |
1206 |
4.30 |
|
|
|
| 19 |
A |
1197 |
1166 |
39.89 |
|
|
|
| 20 |
A |
1187 |
1156 |
16.16 |
|
|
|
| 21 |
A |
1090 |
1063 |
1.24 |
|
|
|
| 22 |
A |
1075 |
1047 |
3.58 |
|
|
|
| 23 |
A |
1041 |
1015 |
5.83 |
|
|
|
| 24 |
A |
1011 |
986 |
18.02 |
|
|
|
| 25 |
A |
946 |
922 |
46.73 |
|
|
|
| 26 |
A |
917 |
894 |
20.71 |
|
|
|
| 27 |
A |
868 |
845 |
25.65 |
|
|
|
| 28 |
A |
837 |
816 |
6.29 |
|
|
|
| 29 |
A |
751 |
732 |
38.54 |
|
|
|
| 30 |
A |
704 |
686 |
1.32 |
|
|
|
| 31 |
A |
614 |
599 |
1.38 |
|
|
|
| 32 |
A |
530 |
516 |
0.69 |
|
|
|
| 33 |
A |
324 |
316 |
3.06 |
|
|
|
| 34 |
A |
221 |
215 |
10.27 |
|
|
|
| 35 |
A |
158 |
154 |
2.35 |
|
|
|
| 36 |
A |
65 |
64 |
2.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26226.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 25557.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.135 |
|
|
|
| 2 |
H |
0.152 |
|
|
|
| 3 |
H |
0.149 |
|
|
|
| 4 |
O |
-0.256 |
|
|
|
| 5 |
C |
-0.372 |
|
|
|
| 6 |
H |
0.144 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
| 8 |
C |
-0.131 |
|
|
|
| 9 |
H |
0.183 |
|
|
|
| 10 |
C |
-0.653 |
|
|
|
| 11 |
H |
0.171 |
|
|
|
| 12 |
H |
0.166 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
C |
0.265 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.652 |
1.142 |
0.111 |
1.320 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.612 |
1.467 |
-0.008 |
| y |
1.467 |
-38.827 |
-0.199 |
| z |
-0.008 |
-0.199 |
-37.790 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.696 |
1.467 |
-0.008 |
| y |
1.467 |
-4.126 |
-0.199 |
| z |
-0.008 |
-0.199 |
-2.570 |
|
| Polar |
|---|
| 3z2-r2 | -5.140 |
|---|
| x2-y2 | 7.215 |
|---|
| xy | 1.467 |
|---|
| xz | -0.008 |
|---|
| yz | -0.199 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.533 |
-0.642 |
0.001 |
| y |
-0.642 |
9.109 |
0.001 |
| z |
0.001 |
0.001 |
6.423 |
<r2> (average value of r
2) Å
2
| <r2> |
133.937 |
| (<r2>)1/2 |
11.573 |