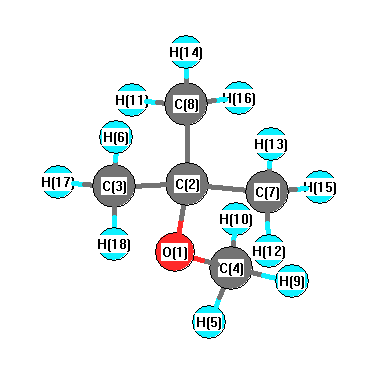
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3103 |
3023 |
44.00 |
|
|
|
| 2 |
A |
3091 |
3013 |
75.82 |
|
|
|
| 3 |
A |
3084 |
3005 |
45.97 |
|
|
|
| 4 |
A |
3075 |
2997 |
58.66 |
|
|
|
| 5 |
A |
2999 |
2923 |
11.50 |
|
|
|
| 6 |
A |
2990 |
2914 |
41.87 |
|
|
|
| 7 |
A |
2960 |
2884 |
60.29 |
|
|
|
| 8 |
A |
1538 |
1499 |
15.10 |
|
|
|
| 9 |
A |
1524 |
1485 |
9.29 |
|
|
|
| 10 |
A |
1515 |
1476 |
3.50 |
|
|
|
| 11 |
A |
1508 |
1470 |
2.36 |
|
|
|
| 12 |
A |
1472 |
1434 |
0.24 |
|
|
|
| 13 |
A |
1436 |
1399 |
9.71 |
|
|
|
| 14 |
A |
1413 |
1377 |
18.94 |
|
|
|
| 15 |
A |
1291 |
1259 |
15.85 |
|
|
|
| 16 |
A |
1223 |
1192 |
80.72 |
|
|
|
| 17 |
A |
1185 |
1155 |
4.68 |
|
|
|
| 18 |
A |
1064 |
1037 |
44.82 |
|
|
|
| 19 |
A |
1037 |
1011 |
55.00 |
|
|
|
| 20 |
A |
929 |
905 |
0.08 |
|
|
|
| 21 |
A |
824 |
803 |
13.23 |
|
|
|
| 22 |
A |
693 |
675 |
8.71 |
|
|
|
| 23 |
A |
490 |
477 |
2.16 |
|
|
|
| 24 |
A |
400 |
389 |
1.14 |
|
|
|
| 25 |
A |
349 |
340 |
0.88 |
|
|
|
| 26 |
A |
267 |
260 |
1.01 |
|
|
|
| 27 |
A |
242 |
235 |
0.38 |
|
|
|
| 28 |
A |
3098 |
3019 |
52.82 |
|
|
|
| 29 |
A |
3084 |
3005 |
9.66 |
|
|
|
| 30 |
A |
3069 |
2991 |
2.30 |
|
|
|
| 31 |
A |
3036 |
2958 |
84.86 |
|
|
|
| 32 |
A |
2986 |
2910 |
22.02 |
|
|
|
| 33 |
A |
1522 |
1483 |
0.03 |
|
|
|
| 34 |
A |
1508 |
1469 |
4.06 |
|
|
|
| 35 |
A |
1504 |
1466 |
4.16 |
|
|
|
| 36 |
A |
1490 |
1452 |
0.01 |
|
|
|
| 37 |
A |
1412 |
1376 |
19.73 |
|
|
|
| 38 |
A |
1266 |
1233 |
28.31 |
|
|
|
| 39 |
A |
1143 |
1114 |
0.89 |
|
|
|
| 40 |
A |
1040 |
1014 |
2.50 |
|
|
|
| 41 |
A |
968 |
943 |
0.06 |
|
|
|
| 42 |
A |
919 |
896 |
0.07 |
|
|
|
| 43 |
A |
440 |
428 |
7.41 |
|
|
|
| 44 |
A |
327 |
319 |
3.09 |
|
|
|
| 45 |
A |
258 |
252 |
1.54 |
|
|
|
| 46 |
A |
210 |
205 |
1.98 |
|
|
|
| 47 |
A |
173 |
169 |
0.84 |
|
|
|
| 48 |
A |
66 |
64 |
5.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35609.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 34701.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.323 |
|
|
|
| 2 |
C |
0.443 |
|
|
|
| 3 |
C |
-0.490 |
|
|
|
| 4 |
C |
-0.308 |
|
|
|
| 5 |
H |
0.167 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
C |
-0.545 |
|
|
|
| 8 |
C |
-0.545 |
|
|
|
| 9 |
H |
0.133 |
|
|
|
| 10 |
H |
0.133 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
| 12 |
H |
0.162 |
|
|
|
| 13 |
H |
0.141 |
|
|
|
| 14 |
H |
0.141 |
|
|
|
| 15 |
H |
0.144 |
|
|
|
| 16 |
H |
0.144 |
|
|
|
| 17 |
H |
0.155 |
|
|
|
| 18 |
H |
0.155 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.664 |
0.085 |
0.000 |
1.666 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.321 |
-1.163 |
0.000 |
| y |
-1.163 |
-36.897 |
0.000 |
| z |
0.000 |
0.000 |
-40.210 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.768 |
-1.163 |
0.000 |
| y |
-1.163 |
4.369 |
0.000 |
| z |
0.000 |
0.000 |
-0.601 |
|
| Polar |
|---|
| 3z2-r2 | -1.202 |
|---|
| x2-y2 | -5.424 |
|---|
| xy | -1.163 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.030 |
-0.045 |
0.000 |
| y |
-0.045 |
10.539 |
0.000 |
| z |
0.000 |
0.000 |
9.121 |
<r2> (average value of r
2) Å
2
| <r2> |
156.986 |
| (<r2>)1/2 |
12.529 |