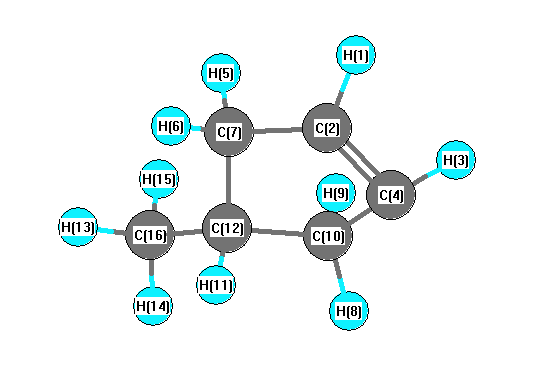
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3174 |
3093 |
49.12 |
|
|
|
| 2 |
A |
3141 |
3061 |
14.63 |
|
|
|
| 3 |
A |
3059 |
2981 |
98.13 |
|
|
|
| 4 |
A |
3058 |
2980 |
47.62 |
|
|
|
| 5 |
A |
3025 |
2947 |
55.59 |
|
|
|
| 6 |
A |
3017 |
2940 |
42.78 |
|
|
|
| 7 |
A |
2978 |
2902 |
10.50 |
|
|
|
| 8 |
A |
2973 |
2897 |
66.16 |
|
|
|
| 9 |
A |
2958 |
2883 |
20.52 |
|
|
|
| 10 |
A |
2957 |
2881 |
74.55 |
|
|
|
| 11 |
A |
1646 |
1604 |
4.45 |
|
|
|
| 12 |
A |
1524 |
1485 |
7.49 |
|
|
|
| 13 |
A |
1520 |
1481 |
2.23 |
|
|
|
| 14 |
A |
1517 |
1479 |
1.75 |
|
|
|
| 15 |
A |
1513 |
1474 |
4.68 |
|
|
|
| 16 |
A |
1430 |
1393 |
4.95 |
|
|
|
| 17 |
A |
1371 |
1336 |
4.83 |
|
|
|
| 18 |
A |
1370 |
1335 |
0.18 |
|
|
|
| 19 |
A |
1332 |
1298 |
0.44 |
|
|
|
| 20 |
A |
1311 |
1278 |
0.85 |
|
|
|
| 21 |
A |
1302 |
1269 |
5.36 |
|
|
|
| 22 |
A |
1238 |
1206 |
0.82 |
|
|
|
| 23 |
A |
1170 |
1141 |
0.01 |
|
|
|
| 24 |
A |
1152 |
1122 |
2.90 |
|
|
|
| 25 |
A |
1130 |
1101 |
0.13 |
|
|
|
| 26 |
A |
1112 |
1083 |
2.28 |
|
|
|
| 27 |
A |
1069 |
1041 |
2.24 |
|
|
|
| 28 |
A |
997 |
972 |
6.92 |
|
|
|
| 29 |
A |
977 |
952 |
0.21 |
|
|
|
| 30 |
A |
971 |
947 |
1.76 |
|
|
|
| 31 |
A |
939 |
915 |
4.98 |
|
|
|
| 32 |
A |
925 |
901 |
4.51 |
|
|
|
| 33 |
A |
906 |
883 |
2.06 |
|
|
|
| 34 |
A |
808 |
787 |
1.71 |
|
|
|
| 35 |
A |
774 |
754 |
0.87 |
|
|
|
| 36 |
A |
713 |
695 |
49.26 |
|
|
|
| 37 |
A |
580 |
565 |
4.60 |
|
|
|
| 38 |
A |
424 |
414 |
1.19 |
|
|
|
| 39 |
A |
388 |
378 |
0.06 |
|
|
|
| 40 |
A |
316 |
308 |
0.12 |
|
|
|
| 41 |
A |
244 |
238 |
0.01 |
|
|
|
| 42 |
A |
95 |
93 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31548.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30744.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.148 |
|
|
|
| 2 |
C |
-0.076 |
|
|
|
| 3 |
H |
0.148 |
|
|
|
| 4 |
C |
-0.076 |
|
|
|
| 5 |
H |
0.140 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
C |
-0.386 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
H |
0.140 |
|
|
|
| 10 |
C |
-0.386 |
|
|
|
| 11 |
H |
0.134 |
|
|
|
| 12 |
C |
0.069 |
|
|
|
| 13 |
H |
0.146 |
|
|
|
| 14 |
H |
0.146 |
|
|
|
| 15 |
H |
0.142 |
|
|
|
| 16 |
C |
-0.558 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.243 |
0.000 |
0.037 |
0.246 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.801 |
0.000 |
0.504 |
| y |
0.000 |
-37.466 |
-0.000 |
| z |
0.504 |
-0.000 |
-39.771 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.182 |
0.000 |
0.504 |
| y |
0.000 |
1.819 |
-0.000 |
| z |
0.504 |
-0.000 |
-1.637 |
|
| Polar |
|---|
| 3z2-r2 | -3.275 |
|---|
| x2-y2 | -1.334 |
|---|
| xy | 0.000 |
|---|
| xz | 0.504 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.541 |
-0.003 |
0.093 |
| y |
-0.003 |
11.230 |
-0.001 |
| z |
0.093 |
-0.001 |
7.693 |
<r2> (average value of r
2) Å
2
| <r2> |
139.334 |
| (<r2>)1/2 |
11.804 |