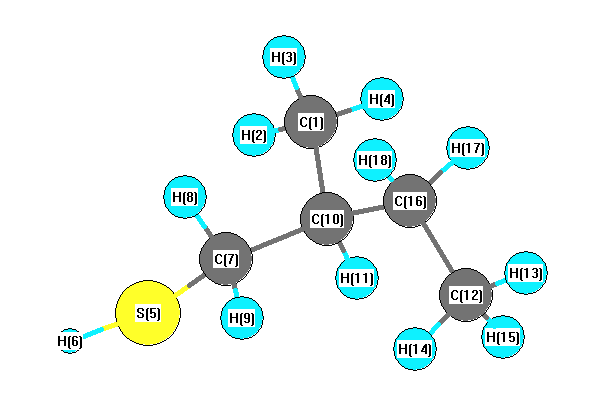
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3109 |
3030 |
27.19 |
|
|
|
| 2 |
A |
3082 |
3004 |
46.28 |
|
|
|
| 3 |
A |
3070 |
2991 |
127.69 |
|
|
|
| 4 |
A |
3066 |
2988 |
68.30 |
|
|
|
| 5 |
A |
3062 |
2984 |
11.82 |
|
|
|
| 6 |
A |
3026 |
2949 |
32.40 |
|
|
|
| 7 |
A |
3017 |
2940 |
15.59 |
|
|
|
| 8 |
A |
2983 |
2907 |
49.01 |
|
|
|
| 9 |
A |
2983 |
2907 |
27.02 |
|
|
|
| 10 |
A |
2980 |
2904 |
27.54 |
|
|
|
| 11 |
A |
2968 |
2892 |
23.04 |
|
|
|
| 12 |
A |
2477 |
2414 |
40.19 |
|
|
|
| 13 |
A |
1530 |
1491 |
11.75 |
|
|
|
| 14 |
A |
1524 |
1485 |
16.60 |
|
|
|
| 15 |
A |
1522 |
1483 |
1.49 |
|
|
|
| 16 |
A |
1521 |
1482 |
2.95 |
|
|
|
| 17 |
A |
1511 |
1472 |
1.67 |
|
|
|
| 18 |
A |
1492 |
1454 |
6.62 |
|
|
|
| 19 |
A |
1433 |
1396 |
14.90 |
|
|
|
| 20 |
A |
1432 |
1395 |
3.77 |
|
|
|
| 21 |
A |
1389 |
1354 |
3.25 |
|
|
|
| 22 |
A |
1369 |
1334 |
2.24 |
|
|
|
| 23 |
A |
1339 |
1305 |
0.90 |
|
|
|
| 24 |
A |
1309 |
1276 |
17.12 |
|
|
|
| 25 |
A |
1277 |
1245 |
11.92 |
|
|
|
| 26 |
A |
1231 |
1200 |
6.77 |
|
|
|
| 27 |
A |
1182 |
1152 |
0.66 |
|
|
|
| 28 |
A |
1146 |
1117 |
3.74 |
|
|
|
| 29 |
A |
1084 |
1057 |
1.28 |
|
|
|
| 30 |
A |
1038 |
1012 |
1.76 |
|
|
|
| 31 |
A |
1035 |
1009 |
2.13 |
|
|
|
| 32 |
A |
989 |
964 |
8.52 |
|
|
|
| 33 |
A |
926 |
903 |
0.56 |
|
|
|
| 34 |
A |
882 |
860 |
5.28 |
|
|
|
| 35 |
A |
858 |
837 |
1.49 |
|
|
|
| 36 |
A |
797 |
777 |
2.07 |
|
|
|
| 37 |
A |
756 |
736 |
10.43 |
|
|
|
| 38 |
A |
682 |
665 |
5.69 |
|
|
|
| 39 |
A |
451 |
439 |
0.28 |
|
|
|
| 40 |
A |
388 |
378 |
0.16 |
|
|
|
| 41 |
A |
373 |
363 |
1.62 |
|
|
|
| 42 |
A |
247 |
241 |
0.08 |
|
|
|
| 43 |
A |
225 |
220 |
1.20 |
|
|
|
| 44 |
A |
207 |
202 |
0.11 |
|
|
|
| 45 |
A |
189 |
184 |
1.78 |
|
|
|
| 46 |
A |
131 |
128 |
17.71 |
|
|
|
| 47 |
A |
92 |
89 |
1.34 |
|
|
|
| 48 |
A |
57 |
55 |
10.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34717.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 33831.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.489 |
|
|
|
| 2 |
H |
0.163 |
|
|
|
| 3 |
H |
0.138 |
|
|
|
| 4 |
H |
0.144 |
|
|
|
| 5 |
S |
0.051 |
|
|
|
| 6 |
H |
0.037 |
|
|
|
| 7 |
C |
-0.614 |
|
|
|
| 8 |
H |
0.175 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
C |
0.091 |
|
|
|
| 11 |
H |
0.132 |
|
|
|
| 12 |
C |
-0.497 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
| 14 |
H |
0.142 |
|
|
|
| 15 |
H |
0.147 |
|
|
|
| 16 |
C |
-0.217 |
|
|
|
| 17 |
H |
0.138 |
|
|
|
| 18 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.668 |
-0.666 |
0.952 |
2.033 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.750 |
2.375 |
-2.149 |
| y |
2.375 |
-45.228 |
-1.823 |
| z |
-2.149 |
-1.823 |
-48.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.993 |
2.375 |
-2.149 |
| y |
2.375 |
4.790 |
-1.823 |
| z |
-2.149 |
-1.823 |
0.202 |
|
| Polar |
|---|
| 3z2-r2 | 0.405 |
|---|
| x2-y2 | -6.522 |
|---|
| xy | 2.375 |
|---|
| xz | -2.149 |
|---|
| yz | -1.823 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.783 |
0.700 |
0.501 |
| y |
0.700 |
11.230 |
-0.222 |
| z |
0.501 |
-0.222 |
9.651 |
<r2> (average value of r
2) Å
2
| <r2> |
206.629 |
| (<r2>)1/2 |
14.375 |