Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -24.156756 |
| Energy at 298.15K | -24.162240 |
| HF Energy | -24.156756 |
| Nuclear repulsion energy | 39.967961 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3350 |
3265 |
21.22 |
|
|
|
| 2 |
A' |
3119 |
3040 |
21.64 |
|
|
|
| 3 |
A' |
2989 |
2913 |
31.98 |
|
|
|
| 4 |
A' |
2970 |
2895 |
69.95 |
|
|
|
| 5 |
A' |
1666 |
1624 |
57.26 |
|
|
|
| 6 |
A' |
1496 |
1458 |
16.74 |
|
|
|
| 7 |
A' |
1430 |
1393 |
14.39 |
|
|
|
| 8 |
A' |
1396 |
1361 |
33.69 |
|
|
|
| 9 |
A' |
1271 |
1239 |
42.96 |
|
|
|
| 10 |
A' |
1065 |
1038 |
37.45 |
|
|
|
| 11 |
A' |
930 |
906 |
7.76 |
|
|
|
| 12 |
A' |
485 |
472 |
18.71 |
|
|
|
| 13 |
A" |
3055 |
2977 |
32.55 |
|
|
|
| 14 |
A" |
1502 |
1464 |
15.04 |
|
|
|
| 15 |
A" |
1132 |
1103 |
1.72 |
|
|
|
| 16 |
A" |
1089 |
1061 |
17.44 |
|
|
|
| 17 |
A" |
698 |
680 |
79.55 |
|
|
|
| 18 |
A" |
192 |
187 |
1.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14918.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 14537.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
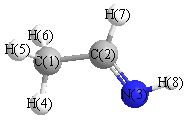
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.061 |
-0.621 |
0.000 |
| C2 |
0.000 |
0.460 |
0.000 |
| N3 |
1.266 |
0.161 |
0.000 |
| H4 |
-0.585 |
-1.606 |
0.000 |
| H5 |
-1.707 |
-0.528 |
0.885 |
| H6 |
-1.707 |
-0.528 |
-0.885 |
| H7 |
-0.372 |
1.500 |
0.000 |
| H8 |
1.874 |
0.998 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5149 | 2.4549 | 1.0938 | 1.0999 | 1.0999 | 2.2301 | 3.3516 |
C2 | 1.5149 | | 1.3009 | 2.1478 | 2.1623 | 2.1623 | 1.1039 | 1.9493 | N3 | 2.4549 | 1.3009 | | 2.5596 | 3.1781 | 3.1781 | 2.1151 | 1.0342 | H4 | 1.0938 | 2.1478 | 2.5596 | | 1.7902 | 1.7902 | 3.1133 | 3.5818 | H5 | 1.0999 | 2.1623 | 3.1781 | 1.7902 | | 1.7696 | 2.5845 | 3.9921 | H6 | 1.0999 | 2.1623 | 3.1781 | 1.7902 | 1.7696 | | 2.5845 | 3.9921 | H7 | 2.2301 | 1.1039 | 2.1151 | 3.1133 | 2.5845 | 2.5845 | | 2.3006 | H8 | 3.3516 | 1.9493 | 1.0342 | 3.5818 | 3.9921 | 3.9921 | 2.3006 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.158 |
|
C1 |
C2 |
H7 |
115.879 |
| C2 |
C1 |
H4 |
109.783 |
|
C2 |
C1 |
H5 |
110.565 |
| C2 |
C1 |
H6 |
110.565 |
|
C2 |
N3 |
H8 |
112.686 |
| N3 |
C2 |
H7 |
122.963 |
|
H4 |
C1 |
H5 |
109.387 |
| H4 |
C1 |
H6 |
109.387 |
|
H5 |
C1 |
H6 |
107.106 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.461 |
|
|
|
| 2 |
C |
-0.195 |
|
|
|
| 3 |
N |
-0.158 |
|
|
|
| 4 |
H |
0.171 |
|
|
|
| 5 |
H |
0.159 |
|
|
|
| 6 |
H |
0.159 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.473 |
1.689 |
0.000 |
2.241 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.783 |
3.091 |
0.000 |
| y |
3.091 |
-18.053 |
0.000 |
| z |
0.000 |
0.000 |
-20.079 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.717 |
3.091 |
0.000 |
| y |
3.091 |
1.878 |
0.000 |
| z |
0.000 |
0.000 |
-1.161 |
|
| Polar |
|---|
| 3z2-r2 | -2.323 |
|---|
| x2-y2 | -1.730 |
|---|
| xy | 3.091 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.291 |
0.190 |
0.000 |
| y |
0.190 |
4.498 |
0.000 |
| z |
0.000 |
0.000 |
3.214 |
<r2> (average value of r
2) Å
2
| <r2> |
44.866 |
| (<r2>)1/2 |
6.698 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -24.155213 |
| Energy at 298.15K | -24.160654 |
| HF Energy | -24.155213 |
| Nuclear repulsion energy | 39.960295 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3305 |
3221 |
33.60 |
|
|
|
| 2 |
A' |
3092 |
3013 |
67.32 |
|
|
|
| 3 |
A' |
3068 |
2989 |
5.51 |
|
|
|
| 4 |
A' |
2980 |
2904 |
17.94 |
|
|
|
| 5 |
A' |
1648 |
1606 |
50.63 |
|
|
|
| 6 |
A' |
1501 |
1462 |
32.00 |
|
|
|
| 7 |
A' |
1427 |
1391 |
22.15 |
|
|
|
| 8 |
A' |
1418 |
1382 |
5.12 |
|
|
|
| 9 |
A' |
1264 |
1232 |
79.71 |
|
|
|
| 10 |
A' |
1066 |
1039 |
29.84 |
|
|
|
| 11 |
A' |
906 |
882 |
2.73 |
|
|
|
| 12 |
A' |
479 |
467 |
9.12 |
|
|
|
| 13 |
A" |
3059 |
2981 |
26.27 |
|
|
|
| 14 |
A" |
1495 |
1457 |
13.94 |
|
|
|
| 15 |
A" |
1152 |
1123 |
57.47 |
|
|
|
| 16 |
A" |
1083 |
1056 |
13.78 |
|
|
|
| 17 |
A" |
695 |
678 |
9.90 |
|
|
|
| 18 |
A" |
170 |
166 |
1.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14903.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 14523.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.049 |
-0.618 |
0.000 |
| C2 |
0.000 |
0.488 |
0.000 |
| N3 |
1.295 |
0.366 |
0.000 |
| H4 |
-0.585 |
-1.612 |
0.000 |
| H5 |
-1.697 |
-0.531 |
0.884 |
| H6 |
-1.697 |
-0.531 |
-0.884 |
| H7 |
-0.386 |
1.513 |
0.000 |
| H8 |
1.597 |
-0.626 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5239 | 2.5418 | 1.0978 | 1.0996 | 1.0996 | 2.2317 | 2.6463 |
C2 | 1.5239 | | 1.3003 | 2.1802 | 2.1677 | 2.1677 | 1.0959 | 1.9474 | N3 | 2.5418 | 1.3003 | | 2.7291 | 3.2460 | 3.2460 | 2.0349 | 1.0378 | H4 | 1.0978 | 2.1802 | 2.7291 | | 1.7859 | 1.7859 | 3.1322 | 2.3946 | H5 | 1.0996 | 2.1677 | 3.2460 | 1.7859 | | 1.7681 | 2.5841 | 3.4123 | H6 | 1.0996 | 2.1677 | 3.2460 | 1.7859 | 1.7681 | | 2.5841 | 3.4123 | H7 | 2.2317 | 1.0959 | 2.0349 | 3.1322 | 2.5841 | 2.5841 | | 2.9176 | H8 | 2.6463 | 1.9474 | 1.0378 | 2.3946 | 3.4123 | 3.4123 | 2.9176 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.143 |
|
C1 |
C2 |
H7 |
115.865 |
| C2 |
C1 |
H4 |
111.484 |
|
C2 |
C1 |
H5 |
110.381 |
| C2 |
C1 |
H6 |
110.381 |
|
C2 |
N3 |
H8 |
112.313 |
| N3 |
C2 |
H7 |
115.992 |
|
H4 |
C1 |
H5 |
108.725 |
| H4 |
C1 |
H6 |
108.725 |
|
H5 |
C1 |
H6 |
107.015 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.503 |
|
|
|
| 2 |
C |
-0.166 |
|
|
|
| 3 |
N |
-0.160 |
|
|
|
| 4 |
H |
0.147 |
|
|
|
| 5 |
H |
0.170 |
|
|
|
| 6 |
H |
0.170 |
|
|
|
| 7 |
H |
0.142 |
|
|
|
| 8 |
H |
0.200 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.240 |
-1.682 |
0.000 |
2.801 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.852 |
-3.749 |
0.000 |
| y |
-3.749 |
-19.338 |
0.000 |
| z |
0.000 |
0.000 |
-20.014 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.176 |
-3.749 |
0.000 |
| y |
-3.749 |
1.595 |
0.000 |
| z |
0.000 |
0.000 |
0.582 |
|
| Polar |
|---|
| 3z2-r2 | 1.163 |
|---|
| x2-y2 | -2.514 |
|---|
| xy | -3.749 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.116 |
-0.163 |
0.000 |
| y |
-0.163 |
4.479 |
0.000 |
| z |
0.000 |
0.000 |
3.184 |
<r2> (average value of r
2) Å
2
| <r2> |
44.884 |
| (<r2>)1/2 |
6.700 |
