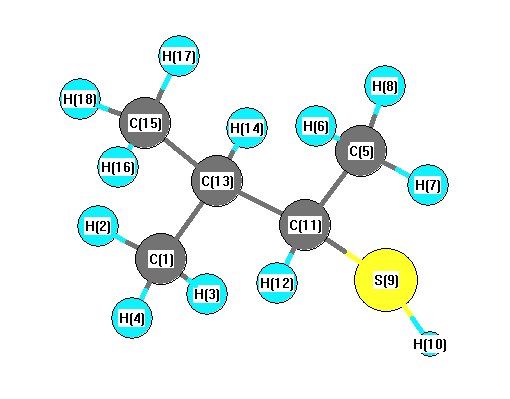
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3092 |
3013 |
40.39 |
|
|
|
| 2 |
A |
3088 |
3009 |
47.99 |
|
|
|
| 3 |
A |
3085 |
3006 |
48.80 |
|
|
|
| 4 |
A |
3071 |
2993 |
45.47 |
|
|
|
| 5 |
A |
3068 |
2990 |
103.18 |
|
|
|
| 6 |
A |
3057 |
2979 |
1.52 |
|
|
|
| 7 |
A |
3022 |
2945 |
21.64 |
|
|
|
| 8 |
A |
2991 |
2915 |
31.35 |
|
|
|
| 9 |
A |
2984 |
2908 |
47.20 |
|
|
|
| 10 |
A |
2981 |
2905 |
5.32 |
|
|
|
| 11 |
A |
2979 |
2903 |
23.61 |
|
|
|
| 12 |
A |
2481 |
2418 |
40.24 |
|
|
|
| 13 |
A |
1540 |
1501 |
4.36 |
|
|
|
| 14 |
A |
1526 |
1487 |
7.23 |
|
|
|
| 15 |
A |
1523 |
1484 |
22.74 |
|
|
|
| 16 |
A |
1514 |
1475 |
3.18 |
|
|
|
| 17 |
A |
1512 |
1473 |
2.37 |
|
|
|
| 18 |
A |
1508 |
1469 |
4.12 |
|
|
|
| 19 |
A |
1440 |
1403 |
7.68 |
|
|
|
| 20 |
A |
1432 |
1395 |
16.66 |
|
|
|
| 21 |
A |
1420 |
1383 |
6.23 |
|
|
|
| 22 |
A |
1381 |
1346 |
0.42 |
|
|
|
| 23 |
A |
1349 |
1315 |
0.49 |
|
|
|
| 24 |
A |
1324 |
1291 |
2.38 |
|
|
|
| 25 |
A |
1247 |
1215 |
33.33 |
|
|
|
| 26 |
A |
1203 |
1172 |
3.86 |
|
|
|
| 27 |
A |
1180 |
1150 |
14.68 |
|
|
|
| 28 |
A |
1145 |
1116 |
0.83 |
|
|
|
| 29 |
A |
1083 |
1056 |
3.18 |
|
|
|
| 30 |
A |
1041 |
1015 |
14.25 |
|
|
|
| 31 |
A |
992 |
966 |
3.76 |
|
|
|
| 32 |
A |
970 |
945 |
0.44 |
|
|
|
| 33 |
A |
941 |
917 |
0.90 |
|
|
|
| 34 |
A |
914 |
891 |
3.57 |
|
|
|
| 35 |
A |
852 |
830 |
2.80 |
|
|
|
| 36 |
A |
768 |
748 |
4.57 |
|
|
|
| 37 |
A |
646 |
629 |
7.63 |
|
|
|
| 38 |
A |
467 |
455 |
0.93 |
|
|
|
| 39 |
A |
394 |
384 |
0.61 |
|
|
|
| 40 |
A |
362 |
352 |
0.24 |
|
|
|
| 41 |
A |
348 |
339 |
0.21 |
|
|
|
| 42 |
A |
321 |
312 |
2.61 |
|
|
|
| 43 |
A |
242 |
236 |
0.81 |
|
|
|
| 44 |
A |
224 |
218 |
1.35 |
|
|
|
| 45 |
A |
215 |
210 |
1.61 |
|
|
|
| 46 |
A |
189 |
184 |
1.32 |
|
|
|
| 47 |
A |
153 |
149 |
19.62 |
|
|
|
| 48 |
A |
59 |
58 |
2.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34659.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 33775.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.488 |
|
|
|
| 2 |
H |
0.142 |
|
|
|
| 3 |
H |
0.165 |
|
|
|
| 4 |
H |
0.137 |
|
|
|
| 5 |
C |
-0.474 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
H |
0.145 |
|
|
|
| 8 |
H |
0.166 |
|
|
|
| 9 |
S |
0.031 |
|
|
|
| 10 |
H |
0.034 |
|
|
|
| 11 |
C |
-0.331 |
|
|
|
| 12 |
H |
0.165 |
|
|
|
| 13 |
C |
0.095 |
|
|
|
| 14 |
H |
0.136 |
|
|
|
| 15 |
C |
-0.527 |
|
|
|
| 16 |
H |
0.142 |
|
|
|
| 17 |
H |
0.156 |
|
|
|
| 18 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.522 |
1.839 |
0.773 |
2.062 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.531 |
2.603 |
2.330 |
| y |
2.603 |
-50.574 |
-0.231 |
| z |
2.330 |
-0.231 |
-48.183 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.848 |
2.603 |
2.330 |
| y |
2.603 |
-3.718 |
-0.231 |
| z |
2.330 |
-0.231 |
-0.130 |
|
| Polar |
|---|
| 3z2-r2 | -0.260 |
|---|
| x2-y2 | 5.043 |
|---|
| xy | 2.603 |
|---|
| xz | 2.330 |
|---|
| yz | -0.231 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.193 |
-0.880 |
0.050 |
| y |
-0.880 |
12.235 |
0.395 |
| z |
0.050 |
0.395 |
9.458 |
<r2> (average value of r
2) Å
2
| <r2> |
188.487 |
| (<r2>)1/2 |
13.729 |