Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -31.137330 |
| Energy at 298.15K | |
| HF Energy | -31.137330 |
| Nuclear repulsion energy | 74.541541 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3231 |
3149 |
0.00 |
|
|
|
| 2 |
A1 |
3218 |
3136 |
26.54 |
|
|
|
| 3 |
A1 |
3179 |
3098 |
2.33 |
|
|
|
| 4 |
A1 |
1544 |
1505 |
0.04 |
|
|
|
| 5 |
A1 |
1431 |
1394 |
1.55 |
|
|
|
| 6 |
A1 |
1132 |
1104 |
0.19 |
|
|
|
| 7 |
A1 |
1075 |
1048 |
14.10 |
|
|
|
| 8 |
A1 |
1029 |
1003 |
12.13 |
|
|
|
| 9 |
A1 |
845 |
824 |
0.27 |
|
|
|
| 10 |
A2 |
954 |
930 |
0.00 |
|
|
|
| 11 |
A2 |
843 |
822 |
0.00 |
|
|
|
| 12 |
A2 |
528 |
514 |
0.00 |
|
|
|
| 13 |
B1 |
930 |
907 |
10.78 |
|
|
|
| 14 |
B1 |
763 |
743 |
0.34 |
|
|
|
| 15 |
B1 |
704 |
686 |
138.12 |
|
|
|
| 16 |
B1 |
503 |
490 |
9.25 |
|
|
|
| 17 |
B2 |
3199 |
3117 |
29.50 |
|
|
|
| 18 |
B2 |
3181 |
3099 |
14.59 |
|
|
|
| 19 |
B2 |
1390 |
1354 |
41.19 |
|
|
|
| 20 |
B2 |
1302 |
1269 |
0.01 |
|
|
|
| 21 |
B2 |
1207 |
1176 |
0.88 |
|
|
|
| 22 |
B2 |
957 |
932 |
0.35 |
|
|
|
| 23 |
B2 |
913 |
890 |
1.97 |
|
|
|
| 24 |
B2 |
75i |
73i |
2.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16990.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 16557.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.242 |
| C2 |
0.000 |
1.144 |
0.400 |
| C3 |
0.000 |
0.690 |
-1.021 |
| C4 |
0.000 |
-0.690 |
-1.021 |
| C5 |
0.000 |
-1.144 |
0.400 |
| H6 |
0.000 |
0.000 |
2.325 |
| H7 |
0.000 |
2.182 |
0.723 |
| H8 |
0.000 |
1.346 |
-1.884 |
| H9 |
0.000 |
-1.346 |
-1.884 |
| H10 |
0.000 |
-2.182 |
0.723 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4206 | 2.3659 | 2.3659 | 1.4206 | 1.0830 | 2.2427 | 3.4036 | 3.4036 | 2.2427 |
C2 | 1.4206 | | 1.4919 | 2.3199 | 2.2881 | 2.2394 | 1.0869 | 2.2928 | 3.3790 | 3.3416 | C3 | 2.3659 | 1.4919 | | 1.3793 | 2.3199 | 3.4165 | 2.2954 | 1.0842 | 2.2111 | 3.3598 | C4 | 2.3659 | 2.3199 | 1.3793 | | 1.4919 | 3.4165 | 3.3598 | 2.2111 | 1.0842 | 2.2954 | C5 | 1.4206 | 2.2881 | 2.3199 | 1.4919 | | 2.2394 | 3.3416 | 3.3790 | 2.2928 | 1.0869 | H6 | 1.0830 | 2.2394 | 3.4165 | 3.4165 | 2.2394 | | 2.7068 | 4.4191 | 4.4191 | 2.7068 | H7 | 2.2427 | 1.0869 | 2.2954 | 3.3598 | 3.3416 | 2.7068 | | 2.7378 | 4.3868 | 4.3637 | H8 | 3.4036 | 2.2928 | 1.0842 | 2.2111 | 3.3790 | 4.4191 | 2.7378 | | 2.6924 | 4.3868 | H9 | 3.4036 | 3.3790 | 2.2111 | 1.0842 | 2.2928 | 4.4191 | 4.3868 | 2.6924 | | 2.7378 | H10 | 2.2427 | 3.3416 | 3.3598 | 2.2954 | 1.0869 | 2.7068 | 4.3637 | 4.3868 | 2.7378 | |
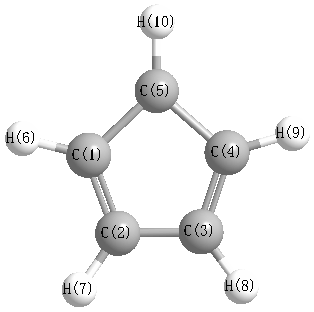 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.624 |
|
C1 |
C2 |
H7 |
126.345 |
| C1 |
C5 |
C4 |
108.624 |
|
C1 |
C5 |
H10 |
126.345 |
| C2 |
C1 |
C5 |
107.287 |
|
C2 |
C1 |
H6 |
126.356 |
| C2 |
C3 |
C4 |
107.733 |
|
C2 |
C3 |
H8 |
125.001 |
| C3 |
C2 |
H7 |
125.031 |
|
C3 |
C4 |
C5 |
107.733 |
| C3 |
C4 |
H9 |
127.266 |
|
C4 |
C3 |
H8 |
127.266 |
| C4 |
C5 |
H10 |
125.031 |
|
C5 |
C1 |
H6 |
126.356 |
| C5 |
C4 |
H9 |
125.001 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.167 |
|
|
|
| 2 |
C |
-0.167 |
|
|
|
| 3 |
C |
-0.171 |
|
|
|
| 4 |
C |
-0.171 |
|
|
|
| 5 |
C |
-0.167 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.170 |
|
|
|
| 9 |
H |
0.170 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.030 |
0.030 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.699 |
0.000 |
0.000 |
| y |
0.000 |
-25.690 |
0.000 |
| z |
0.000 |
0.000 |
-27.859 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.925 |
0.000 |
0.000 |
| y |
0.000 |
5.089 |
0.000 |
| z |
0.000 |
0.000 |
1.836 |
|
| Polar |
|---|
| 3z2-r2 | 3.672 |
|---|
| x2-y2 | -8.009 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.783 |
0.000 |
0.000 |
| y |
0.000 |
9.566 |
0.000 |
| z |
0.000 |
0.000 |
8.519 |
<r2> (average value of r
2) Å
2
| <r2> |
74.928 |
| (<r2>)1/2 |
8.656 |
