Jump to
S2C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -12.337338 |
| Energy at 298.15K | -12.337199 |
| Nuclear repulsion energy | 12.524924 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3091 |
3013 |
35.37 |
|
|
|
| 2 |
A1 |
1618 |
1576 |
98.27 |
|
|
|
| 3 |
A1 |
1249 |
1217 |
22.72 |
|
|
|
| 4 |
B1 |
817 |
797 |
141.65 |
|
|
|
| 5 |
B2 |
3187 |
3105 |
5.09 |
|
|
|
| 6 |
B2 |
492 |
480 |
3.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5226.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 5093.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.491 |
| C2 |
0.000 |
0.000 |
0.841 |
| H3 |
0.000 |
0.938 |
-1.050 |
| H4 |
0.000 |
-0.938 |
-1.050 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3312 | 1.0920 | 1.0920 |
C2 | 1.3312 | | 2.1104 | 2.1104 | H3 | 1.0920 | 2.1104 | | 1.8758 | H4 | 1.0920 | 2.1104 | 1.8758 | |
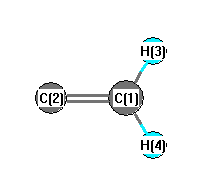 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C2 |
C1 |
H3 |
120.811 |
|
C2 |
C1 |
H4 |
120.811 |
| H3 |
C1 |
H4 |
118.379 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.412 |
|
|
|
| 2 |
C |
-0.005 |
|
|
|
| 3 |
H |
0.209 |
|
|
|
| 4 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.470 |
2.470 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.434 |
0.000 |
0.000 |
| y |
0.000 |
-10.283 |
0.000 |
| z |
0.000 |
0.000 |
-14.485 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.050 |
0.000 |
0.000 |
| y |
0.000 |
4.177 |
0.000 |
| z |
0.000 |
0.000 |
-2.126 |
|
| Polar |
|---|
| 3z2-r2 | -4.253 |
|---|
| x2-y2 | -4.151 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.920 |
0.000 |
0.000 |
| y |
0.000 |
2.702 |
0.000 |
| z |
0.000 |
0.000 |
4.345 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -12.269722 |
| Energy at 298.15K | -12.269644 |
| HF Energy | -12.269722 |
| Nuclear repulsion energy | 12.415408 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3008 |
2931 |
5.75 |
|
|
|
| 2 |
A1 |
1500 |
1461 |
3.73 |
|
|
|
| 3 |
A1 |
1409 |
1373 |
2.92 |
|
|
|
| 4 |
B1 |
504 |
491 |
76.80 |
|
|
|
| 5 |
B2 |
3104 |
3025 |
15.69 |
|
|
|
| 6 |
B2 |
1007 |
981 |
8.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5264.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 5130.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.497 |
| C2 |
0.000 |
0.000 |
0.851 |
| H3 |
0.000 |
0.938 |
-1.062 |
| H4 |
0.000 |
-0.938 |
-1.062 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
| C1 | | 1.3476 | 1.0956 | 1.0956 |
C2 | 1.3476 | | 2.1310 | 2.1310 | H3 | 1.0956 | 2.1310 | | 1.8765 | H4 | 1.0956 | 2.1310 | 1.8765 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.315 |
|
|
|
| 2 |
C |
0.001 |
|
|
|
| 3 |
H |
0.157 |
|
|
|
| 4 |
H |
0.157 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.625 |
0.625 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.079 |
0.000 |
0.000 |
| y |
0.000 |
-12.019 |
0.000 |
| z |
0.000 |
0.000 |
-11.303 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.418 |
0.000 |
0.000 |
| y |
0.000 |
0.672 |
0.000 |
| z |
0.000 |
0.000 |
1.746 |
|
| Polar |
|---|
| 3z2-r2 | 3.491 |
|---|
| x2-y2 | -2.060 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.908 |
0.000 |
0.000 |
| y |
0.000 |
2.284 |
0.000 |
| z |
0.000 |
0.000 |
4.512 |
<r2> (average value of r
2) Å
2
| <r2> |
15.688 |
| (<r2>)1/2 |
3.961 |
