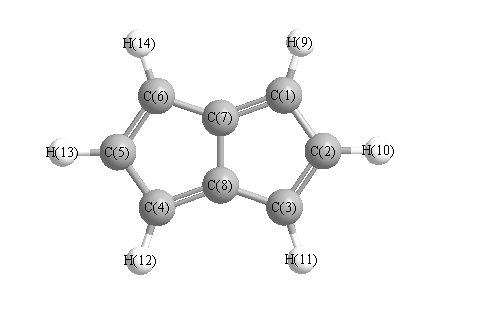
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3218 |
3136 |
0.00 |
|
|
|
| 2 |
Ag |
3192 |
3111 |
0.00 |
|
|
|
| 3 |
Ag |
3179 |
3098 |
0.00 |
|
|
|
| 4 |
Ag |
1618 |
1577 |
0.00 |
|
|
|
| 5 |
Ag |
1520 |
1481 |
0.00 |
|
|
|
| 6 |
Ag |
1377 |
1342 |
0.00 |
|
|
|
| 7 |
Ag |
1312 |
1279 |
0.00 |
|
|
|
| 8 |
Ag |
1187 |
1156 |
0.00 |
|
|
|
| 9 |
Ag |
1111 |
1082 |
0.00 |
|
|
|
| 10 |
Ag |
944 |
920 |
0.00 |
|
|
|
| 11 |
Ag |
839 |
818 |
0.00 |
|
|
|
| 12 |
Ag |
710 |
692 |
0.00 |
|
|
|
| 13 |
Ag |
648 |
632 |
0.00 |
|
|
|
| 14 |
Au |
949 |
925 |
14.11 |
|
|
|
| 15 |
Au |
903 |
880 |
19.84 |
|
|
|
| 16 |
Au |
756 |
737 |
131.83 |
|
|
|
| 17 |
Au |
516 |
503 |
11.04 |
|
|
|
| 18 |
Au |
299 |
291 |
1.72 |
|
|
|
| 19 |
Au |
190 |
185 |
5.73 |
|
|
|
| 20 |
Bg |
956 |
931 |
0.00 |
|
|
|
| 21 |
Bg |
937 |
913 |
0.00 |
|
|
|
| 22 |
Bg |
823 |
802 |
0.00 |
|
|
|
| 23 |
Bg |
690 |
672 |
0.00 |
|
|
|
| 24 |
Bg |
550 |
536 |
0.00 |
|
|
|
| 25 |
Bu |
3218 |
3136 |
56.91 |
|
|
|
| 26 |
Bu |
3193 |
3111 |
35.40 |
|
|
|
| 27 |
Bu |
3180 |
3099 |
17.81 |
|
|
|
| 28 |
Bu |
1622 |
1580 |
63.83 |
|
|
|
| 29 |
Bu |
1504 |
1466 |
19.64 |
|
|
|
| 30 |
Bu |
1337 |
1303 |
17.05 |
|
|
|
| 31 |
Bu |
1267 |
1235 |
20.69 |
|
|
|
| 32 |
Bu |
1130 |
1101 |
54.45 |
|
|
|
| 33 |
Bu |
1043 |
1016 |
12.98 |
|
|
|
| 34 |
Bu |
938 |
914 |
7.17 |
|
|
|
| 35 |
Bu |
851 |
829 |
8.66 |
|
|
|
| 36 |
Bu |
443 |
431 |
4.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24074.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 23460.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.245 |
|
|
|
| 2 |
C |
-0.293 |
|
|
|
| 3 |
C |
0.047 |
|
|
|
| 4 |
C |
-0.245 |
|
|
|
| 5 |
C |
-0.293 |
|
|
|
| 6 |
C |
0.047 |
|
|
|
| 7 |
C |
-0.052 |
|
|
|
| 8 |
C |
-0.052 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.158 |
|
|
|
| 11 |
H |
0.196 |
|
|
|
| 12 |
H |
0.189 |
|
|
|
| 13 |
H |
0.158 |
|
|
|
| 14 |
H |
0.196 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.203 |
1.479 |
0.000 |
| y |
1.479 |
-40.734 |
0.000 |
| z |
0.000 |
0.000 |
-51.490 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.909 |
1.479 |
0.000 |
| y |
1.479 |
5.612 |
0.000 |
| z |
0.000 |
0.000 |
-10.521 |
|
| Polar |
|---|
| 3z2-r2 | -21.042 |
|---|
| x2-y2 | -0.469 |
|---|
| xy | 1.479 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.826 |
1.359 |
0.000 |
| y |
1.359 |
12.955 |
0.000 |
| z |
0.000 |
0.000 |
5.585 |
<r2> (average value of r
2) Å
2
| <r2> |
175.903 |
| (<r2>)1/2 |
13.263 |