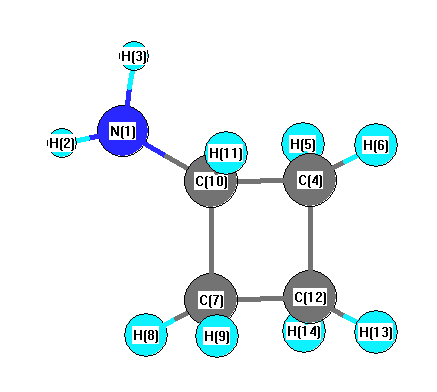
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3465 |
3377 |
6.52 |
|
|
|
| 2 |
A' |
3105 |
3025 |
103.56 |
|
|
|
| 3 |
A' |
3071 |
2993 |
23.40 |
|
|
|
| 4 |
A' |
3033 |
2956 |
45.43 |
|
|
|
| 5 |
A' |
3012 |
2936 |
45.11 |
|
|
|
| 6 |
A' |
2916 |
2842 |
133.23 |
|
|
|
| 7 |
A' |
1689 |
1645 |
34.01 |
|
|
|
| 8 |
A' |
1514 |
1475 |
4.40 |
|
|
|
| 9 |
A' |
1493 |
1455 |
2.00 |
|
|
|
| 10 |
A' |
1372 |
1337 |
29.82 |
|
|
|
| 11 |
A' |
1289 |
1257 |
6.27 |
|
|
|
| 12 |
A' |
1236 |
1204 |
0.38 |
|
|
|
| 13 |
A' |
1165 |
1135 |
5.19 |
|
|
|
| 14 |
A' |
1072 |
1045 |
8.87 |
|
|
|
| 15 |
A' |
946 |
922 |
0.57 |
|
|
|
| 16 |
A' |
878 |
855 |
3.18 |
|
|
|
| 17 |
A' |
822 |
801 |
6.14 |
|
|
|
| 18 |
A' |
702 |
684 |
52.03 |
|
|
|
| 19 |
A' |
643 |
626 |
164.27 |
|
|
|
| 20 |
A' |
406 |
396 |
6.46 |
|
|
|
| 21 |
A' |
129 |
125 |
1.21 |
|
|
|
| 22 |
A" |
3591 |
3499 |
0.75 |
|
|
|
| 23 |
A" |
3078 |
2999 |
15.11 |
|
|
|
| 24 |
A" |
3009 |
2933 |
109.51 |
|
|
|
| 25 |
A" |
1487 |
1449 |
3.74 |
|
|
|
| 26 |
A" |
1328 |
1294 |
1.02 |
|
|
|
| 27 |
A" |
1285 |
1252 |
0.08 |
|
|
|
| 28 |
A" |
1264 |
1231 |
1.43 |
|
|
|
| 29 |
A" |
1210 |
1179 |
1.64 |
|
|
|
| 30 |
A" |
1164 |
1134 |
0.13 |
|
|
|
| 31 |
A" |
1025 |
998 |
0.01 |
|
|
|
| 32 |
A" |
946 |
922 |
1.21 |
|
|
|
| 33 |
A" |
908 |
884 |
2.89 |
|
|
|
| 34 |
A" |
775 |
755 |
0.05 |
|
|
|
| 35 |
A" |
391 |
381 |
8.67 |
|
|
|
| 36 |
A" |
239 |
233 |
38.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27827.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 27117.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.635 |
|
|
|
| 2 |
H |
0.263 |
|
|
|
| 3 |
H |
0.263 |
|
|
|
| 4 |
C |
-0.310 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.116 |
|
|
|
| 7 |
C |
-0.310 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
H |
0.116 |
|
|
|
| 10 |
C |
0.074 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
C |
-0.224 |
|
|
|
| 13 |
H |
0.131 |
|
|
|
| 14 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.228 |
1.175 |
0.000 |
1.197 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.719 |
-2.447 |
0.000 |
| y |
-2.447 |
-31.313 |
0.000 |
| z |
0.000 |
0.000 |
-31.641 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.242 |
-2.447 |
0.000 |
| y |
-2.447 |
0.867 |
0.000 |
| z |
0.000 |
0.000 |
0.375 |
|
| Polar |
|---|
| 3z2-r2 | 0.749 |
|---|
| x2-y2 | -1.406 |
|---|
| xy | -2.447 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.851 |
-0.515 |
0.000 |
| y |
-0.515 |
7.753 |
0.000 |
| z |
0.000 |
0.000 |
7.919 |
<r2> (average value of r
2) Å
2
| <r2> |
100.065 |
| (<r2>)1/2 |
10.003 |