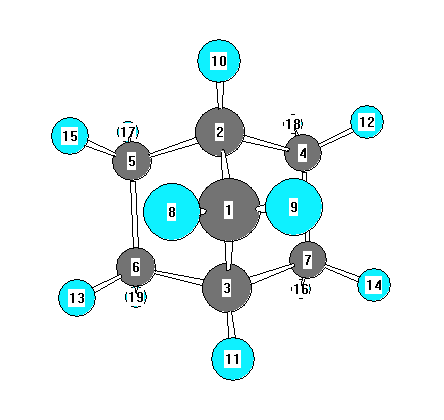
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3071 |
2993 |
152.58 |
|
|
|
| 2 |
A1 |
3065 |
2987 |
4.83 |
|
|
|
| 3 |
A1 |
3009 |
2933 |
15.38 |
|
|
|
| 4 |
A1 |
3001 |
2924 |
71.47 |
|
|
|
| 5 |
A1 |
1543 |
1504 |
0.48 |
|
|
|
| 6 |
A1 |
1511 |
1472 |
8.70 |
|
|
|
| 7 |
A1 |
1342 |
1308 |
0.81 |
|
|
|
| 8 |
A1 |
1278 |
1245 |
1.51 |
|
|
|
| 9 |
A1 |
1159 |
1129 |
1.28 |
|
|
|
| 10 |
A1 |
1005 |
979 |
0.10 |
|
|
|
| 11 |
A1 |
924 |
900 |
2.77 |
|
|
|
| 12 |
A1 |
875 |
853 |
1.91 |
|
|
|
| 13 |
A1 |
818 |
797 |
0.12 |
|
|
|
| 14 |
A1 |
753 |
734 |
0.97 |
|
|
|
| 15 |
A1 |
400 |
390 |
0.01 |
|
|
|
| 16 |
A2 |
3037 |
2959 |
0.00 |
|
|
|
| 17 |
A2 |
2992 |
2916 |
0.00 |
|
|
|
| 18 |
A2 |
1507 |
1469 |
0.00 |
|
|
|
| 19 |
A2 |
1323 |
1290 |
0.00 |
|
|
|
| 20 |
A2 |
1310 |
1277 |
0.00 |
|
|
|
| 21 |
A2 |
1236 |
1204 |
0.00 |
|
|
|
| 22 |
A2 |
1137 |
1108 |
0.00 |
|
|
|
| 23 |
A2 |
971 |
946 |
0.00 |
|
|
|
| 24 |
A2 |
959 |
935 |
0.00 |
|
|
|
| 25 |
A2 |
549 |
535 |
0.00 |
|
|
|
| 26 |
A2 |
172 |
167 |
0.00 |
|
|
|
| 27 |
B1 |
3067 |
2989 |
104.70 |
|
|
|
| 28 |
B1 |
3057 |
2979 |
1.26 |
|
|
|
| 29 |
B1 |
3008 |
2931 |
133.63 |
|
|
|
| 30 |
B1 |
1522 |
1484 |
7.08 |
|
|
|
| 31 |
B1 |
1349 |
1315 |
0.01 |
|
|
|
| 32 |
B1 |
1233 |
1202 |
4.12 |
|
|
|
| 33 |
B1 |
1128 |
1099 |
0.12 |
|
|
|
| 34 |
B1 |
1040 |
1013 |
0.23 |
|
|
|
| 35 |
B1 |
960 |
935 |
0.73 |
|
|
|
| 36 |
B1 |
887 |
864 |
2.35 |
|
|
|
| 37 |
B1 |
808 |
787 |
0.54 |
|
|
|
| 38 |
B1 |
449 |
437 |
0.02 |
|
|
|
| 39 |
B2 |
3055 |
2977 |
134.76 |
|
|
|
| 40 |
B2 |
3041 |
2964 |
19.93 |
|
|
|
| 41 |
B2 |
2992 |
2916 |
78.52 |
|
|
|
| 42 |
B2 |
1519 |
1480 |
2.34 |
|
|
|
| 43 |
B2 |
1341 |
1307 |
0.96 |
|
|
|
| 44 |
B2 |
1298 |
1265 |
0.33 |
|
|
|
| 45 |
B2 |
1270 |
1238 |
0.65 |
|
|
|
| 46 |
B2 |
1181 |
1151 |
0.29 |
|
|
|
| 47 |
B2 |
1096 |
1068 |
0.33 |
|
|
|
| 48 |
B2 |
972 |
947 |
0.54 |
|
|
|
| 49 |
B2 |
813 |
792 |
3.36 |
|
|
|
| 50 |
B2 |
760 |
740 |
0.02 |
|
|
|
| 51 |
B2 |
341 |
333 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38566.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 37583.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.362 |
|
|
|
| 2 |
C |
0.039 |
|
|
|
| 3 |
C |
0.039 |
|
|
|
| 4 |
C |
-0.323 |
|
|
|
| 5 |
C |
-0.323 |
|
|
|
| 6 |
C |
-0.323 |
|
|
|
| 7 |
C |
-0.323 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
H |
0.135 |
|
|
|
| 10 |
H |
0.143 |
|
|
|
| 11 |
H |
0.143 |
|
|
|
| 12 |
H |
0.135 |
|
|
|
| 13 |
H |
0.135 |
|
|
|
| 14 |
H |
0.135 |
|
|
|
| 15 |
H |
0.135 |
|
|
|
| 16 |
H |
0.121 |
|
|
|
| 17 |
H |
0.121 |
|
|
|
| 18 |
H |
0.121 |
|
|
|
| 19 |
H |
0.121 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.067 |
0.067 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.582 |
0.000 |
0.000 |
| y |
0.000 |
-45.888 |
0.000 |
| z |
0.000 |
0.000 |
-45.391 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.057 |
0.000 |
0.000 |
| y |
0.000 |
-0.401 |
0.000 |
| z |
0.000 |
0.000 |
0.343 |
|
| Polar |
|---|
| 3z2-r2 | 0.687 |
|---|
| x2-y2 | 0.305 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.451 |
0.000 |
0.000 |
| y |
0.000 |
10.980 |
0.000 |
| z |
0.000 |
0.000 |
10.774 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |