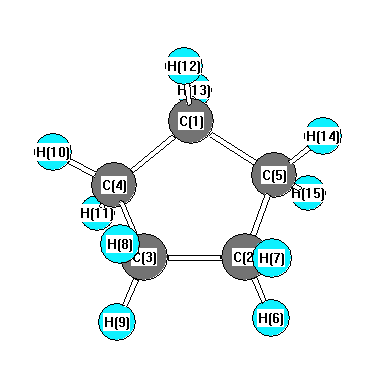
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3072 |
2994 |
190.40 |
|
|
|
| 2 |
A |
3052 |
2974 |
29.47 |
|
|
|
| 3 |
A |
3051 |
2973 |
40.94 |
|
|
|
| 4 |
A |
3046 |
2968 |
18.07 |
|
|
|
| 5 |
A |
3031 |
2954 |
49.24 |
|
|
|
| 6 |
A |
3010 |
2933 |
78.69 |
|
|
|
| 7 |
A |
2990 |
2914 |
82.87 |
|
|
|
| 8 |
A |
2987 |
2911 |
12.60 |
|
|
|
| 9 |
A |
2982 |
2906 |
50.74 |
|
|
|
| 10 |
A |
2981 |
2905 |
32.05 |
|
|
|
| 11 |
A |
1542 |
1502 |
0.18 |
|
|
|
| 12 |
A |
1521 |
1482 |
6.09 |
|
|
|
| 13 |
A |
1520 |
1481 |
4.58 |
|
|
|
| 14 |
A |
1515 |
1477 |
5.64 |
|
|
|
| 15 |
A |
1511 |
1472 |
0.72 |
|
|
|
| 16 |
A |
1350 |
1316 |
0.00 |
|
|
|
| 17 |
A |
1350 |
1315 |
0.02 |
|
|
|
| 18 |
A |
1337 |
1303 |
0.11 |
|
|
|
| 19 |
A |
1321 |
1287 |
0.19 |
|
|
|
| 20 |
A |
1318 |
1284 |
0.58 |
|
|
|
| 21 |
A |
1277 |
1244 |
0.90 |
|
|
|
| 22 |
A |
1238 |
1207 |
0.30 |
|
|
|
| 23 |
A |
1222 |
1191 |
0.67 |
|
|
|
| 24 |
A |
1209 |
1178 |
0.46 |
|
|
|
| 25 |
A |
1171 |
1141 |
0.00 |
|
|
|
| 26 |
A |
1062 |
1035 |
0.04 |
|
|
|
| 27 |
A |
1052 |
1025 |
0.00 |
|
|
|
| 28 |
A |
1006 |
981 |
0.99 |
|
|
|
| 29 |
A |
978 |
953 |
1.98 |
|
|
|
| 30 |
A |
904 |
881 |
4.37 |
|
|
|
| 31 |
A |
899 |
876 |
4.34 |
|
|
|
| 32 |
A |
872 |
850 |
0.54 |
|
|
|
| 33 |
A |
867 |
845 |
1.00 |
|
|
|
| 34 |
A |
823 |
802 |
0.12 |
|
|
|
| 35 |
A |
773 |
754 |
1.41 |
|
|
|
| 36 |
A |
621 |
606 |
0.12 |
|
|
|
| 37 |
A |
555 |
541 |
1.51 |
|
|
|
| 38 |
A |
255 |
248 |
0.01 |
|
|
|
| 39 |
A |
28 |
27 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30649.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29867.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.254 |
|
|
|
| 2 |
C |
-0.260 |
|
|
|
| 3 |
C |
-0.267 |
|
|
|
| 4 |
C |
-0.257 |
|
|
|
| 5 |
C |
-0.254 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
H |
0.129 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
H |
0.130 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.130 |
|
|
|
| 12 |
H |
0.135 |
|
|
|
| 13 |
H |
0.124 |
|
|
|
| 14 |
H |
0.125 |
|
|
|
| 15 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.001 |
0.000 |
0.001 |
0.001 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.172 |
-0.004 |
0.000 |
| y |
-0.004 |
-34.183 |
-0.020 |
| z |
0.000 |
-0.020 |
-32.916 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.623 |
-0.004 |
0.000 |
| y |
-0.004 |
-0.639 |
-0.020 |
| z |
0.000 |
-0.020 |
1.262 |
|
| Polar |
|---|
| 3z2-r2 | 2.524 |
|---|
| x2-y2 | 0.010 |
|---|
| xy | -0.004 |
|---|
| xz | 0.000 |
|---|
| yz | -0.020 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.619 |
0.015 |
-0.005 |
| y |
0.015 |
8.613 |
0.004 |
| z |
-0.005 |
0.004 |
7.773 |
<r2> (average value of r
2) Å
2
| <r2> |
101.306 |
| (<r2>)1/2 |
10.065 |