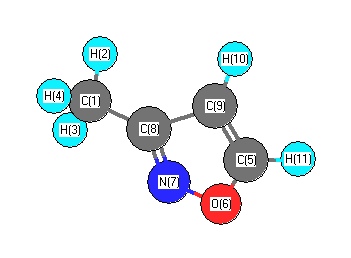
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3272 |
3188 |
1.31 |
|
|
|
| 2 |
A |
3245 |
3163 |
0.37 |
|
|
|
| 3 |
A |
3119 |
3039 |
12.34 |
|
|
|
| 4 |
A |
3080 |
3001 |
23.32 |
|
|
|
| 5 |
A |
3006 |
2929 |
23.55 |
|
|
|
| 6 |
A |
1563 |
1523 |
25.38 |
|
|
|
| 7 |
A |
1507 |
1469 |
12.36 |
|
|
|
| 8 |
A |
1505 |
1467 |
17.93 |
|
|
|
| 9 |
A |
1445 |
1408 |
20.54 |
|
|
|
| 10 |
A |
1433 |
1397 |
1.05 |
|
|
|
| 11 |
A |
1393 |
1357 |
24.08 |
|
|
|
| 12 |
A |
1272 |
1240 |
0.51 |
|
|
|
| 13 |
A |
1126 |
1097 |
14.41 |
|
|
|
| 14 |
A |
1080 |
1052 |
2.70 |
|
|
|
| 15 |
A |
1038 |
1012 |
18.08 |
|
|
|
| 16 |
A |
1026 |
1000 |
1.29 |
|
|
|
| 17 |
A |
969 |
945 |
2.26 |
|
|
|
| 18 |
A |
896 |
873 |
1.46 |
|
|
|
| 19 |
A |
883 |
861 |
1.00 |
|
|
|
| 20 |
A |
831 |
810 |
23.15 |
|
|
|
| 21 |
A |
797 |
776 |
66.72 |
|
|
|
| 22 |
A |
643 |
626 |
0.03 |
|
|
|
| 23 |
A |
639 |
623 |
8.34 |
|
|
|
| 24 |
A |
568 |
554 |
4.54 |
|
|
|
| 25 |
A |
328 |
320 |
7.28 |
|
|
|
| 26 |
A |
263 |
257 |
4.73 |
|
|
|
| 27 |
A |
129 |
126 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18527.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 18055.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.625 |
|
|
|
| 2 |
H |
0.173 |
|
|
|
| 3 |
H |
0.189 |
|
|
|
| 4 |
H |
0.173 |
|
|
|
| 5 |
C |
-0.161 |
|
|
|
| 6 |
O |
-0.185 |
|
|
|
| 7 |
N |
0.106 |
|
|
|
| 8 |
C |
-0.097 |
|
|
|
| 9 |
C |
0.019 |
|
|
|
| 10 |
H |
0.206 |
|
|
|
| 11 |
H |
0.200 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.016 |
3.615 |
0.000 |
3.755 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.996 |
-3.017 |
0.000 |
| y |
-3.017 |
-37.686 |
-0.000 |
| z |
0.000 |
-0.000 |
-36.692 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.193 |
-3.017 |
0.000 |
| y |
-3.017 |
-4.342 |
-0.000 |
| z |
0.000 |
-0.000 |
-2.851 |
|
| Polar |
|---|
| 3z2-r2 | -5.701 |
|---|
| x2-y2 | 7.690 |
|---|
| xy | -3.017 |
|---|
| xz | 0.000 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.946 |
-0.112 |
0.003 |
| y |
-0.112 |
7.918 |
0.001 |
| z |
0.003 |
0.001 |
4.696 |
<r2> (average value of r
2) Å
2
| <r2> |
115.879 |
| (<r2>)1/2 |
10.765 |