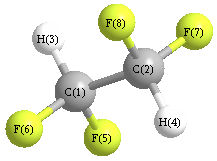
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3131 |
3051 |
0.00 |
|
|
|
| 2 |
Ag |
1425 |
1389 |
0.00 |
|
|
|
| 3 |
Ag |
1129 |
1100 |
0.00 |
|
|
|
| 4 |
Ag |
1034 |
1008 |
0.00 |
|
|
|
| 5 |
Ag |
566 |
551 |
0.00 |
|
|
|
| 6 |
Ag |
331 |
322 |
0.00 |
|
|
|
| 7 |
Au |
1317 |
1284 |
12.16 |
|
|
|
| 8 |
Au |
1051 |
1024 |
354.29 |
|
|
|
| 9 |
Au |
195 |
190 |
4.17 |
|
|
|
| 10 |
Au |
66 |
64 |
4.08 |
|
|
|
| 11 |
Bg |
1346 |
1312 |
0.00 |
|
|
|
| 12 |
Bg |
1014 |
988 |
0.00 |
|
|
|
| 13 |
Bg |
447 |
435 |
0.00 |
|
|
|
| 14 |
Bu |
3141 |
3061 |
24.68 |
|
|
|
| 15 |
Bu |
1291 |
1258 |
19.45 |
|
|
|
| 16 |
Bu |
1038 |
1012 |
185.15 |
|
|
|
| 17 |
Bu |
478 |
466 |
13.88 |
|
|
|
| 18 |
Bu |
394 |
384 |
73.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9696.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 9449.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.190 |
|
|
|
| 2 |
C |
0.190 |
|
|
|
| 3 |
H |
0.181 |
|
|
|
| 4 |
H |
0.181 |
|
|
|
| 5 |
F |
-0.185 |
|
|
|
| 6 |
F |
-0.185 |
|
|
|
| 7 |
F |
-0.185 |
|
|
|
| 8 |
F |
-0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.521 |
-4.101 |
0.000 |
| y |
-4.101 |
-37.507 |
0.000 |
| z |
0.000 |
0.000 |
-38.055 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.260 |
-4.101 |
0.000 |
| y |
-4.101 |
-4.220 |
0.000 |
| z |
0.000 |
0.000 |
-5.040 |
|
| Polar |
|---|
| 3z2-r2 | -10.081 |
|---|
| x2-y2 | 8.986 |
|---|
| xy | -4.101 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.328 |
0.166 |
0.000 |
| y |
0.166 |
3.565 |
0.000 |
| z |
0.000 |
0.000 |
4.076 |
<r2> (average value of r
2) Å
2
| <r2> |
123.614 |
| (<r2>)1/2 |
11.118 |