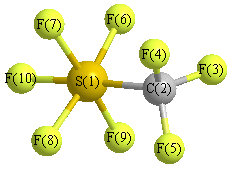
Vibrational Frequencies calculated at HF/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
1354 |
1235 |
231.23 |
|
|
|
| 2 |
A' |
1058 |
965 |
366.37 |
|
|
|
| 3 |
A' |
905 |
826 |
352.24 |
|
|
|
| 4 |
A' |
855 |
780 |
216.59 |
|
|
|
| 5 |
A' |
728 |
664 |
193.29 |
|
|
|
| 6 |
A' |
609 |
555 |
9.18 |
|
|
|
| 7 |
A' |
604 |
552 |
0.07 |
|
|
|
| 8 |
A' |
543 |
496 |
2.23 |
|
|
|
| 9 |
A' |
502 |
458 |
116.90 |
|
|
|
| 10 |
A' |
437 |
399 |
26.20 |
|
|
|
| 11 |
A' |
338 |
309 |
1.13 |
|
|
|
| 12 |
A' |
301 |
275 |
3.09 |
|
|
|
| 13 |
A' |
283 |
258 |
12.59 |
|
|
|
| 14 |
A' |
259 |
236 |
1.56 |
|
|
|
| 15 |
A' |
173 |
158 |
1.75 |
|
|
|
| 16 |
A" |
1350 |
1232 |
227.71 |
|
|
|
| 17 |
A" |
906 |
827 |
351.61 |
|
|
|
| 18 |
A" |
540 |
493 |
2.11 |
|
|
|
| 19 |
A" |
439 |
400 |
26.09 |
|
|
|
| 20 |
A" |
363 |
332 |
0.06 |
|
|
|
| 21 |
A" |
336 |
307 |
1.56 |
|
|
|
| 22 |
A" |
288 |
262 |
3.39 |
|
|
|
| 23 |
A" |
183 |
167 |
1.40 |
|
|
|
| 24 |
A" |
9 |
8 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6680.6 cm
-1
Scaled (by 0.9125) Zero Point Vibrational Energy (zpe) 6096.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
1.655 |
|
|
|
| 2 |
C |
0.464 |
|
|
|
| 3 |
F |
-0.117 |
|
|
|
| 4 |
F |
-0.117 |
|
|
|
| 5 |
F |
-0.117 |
|
|
|
| 6 |
F |
-0.362 |
|
|
|
| 7 |
F |
-0.366 |
|
|
|
| 8 |
F |
-0.372 |
|
|
|
| 9 |
F |
-0.366 |
|
|
|
| 10 |
F |
-0.303 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.859 |
2.508 |
0.000 |
2.651 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-65.960 |
-1.387 |
0.000 |
| y |
-1.387 |
-62.137 |
0.000 |
| z |
0.000 |
0.000 |
-66.340 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.722 |
-1.387 |
0.000 |
| y |
-1.387 |
4.013 |
0.000 |
| z |
0.000 |
0.000 |
-2.291 |
|
| Polar |
|---|
| 3z2-r2 | -4.583 |
|---|
| x2-y2 | -3.823 |
|---|
| xy | -1.387 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.278 |
-0.074 |
0.000 |
| y |
-0.074 |
6.880 |
0.000 |
| z |
0.000 |
0.000 |
6.274 |
<r2> (average value of r
2) Å
2
| <r2> |
285.835 |
| (<r2>)1/2 |
16.907 |