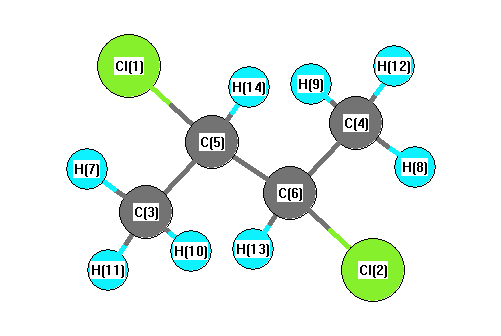
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3116 |
3036 |
0.00 |
|
|
|
| 2 |
Ag |
3108 |
3029 |
0.00 |
|
|
|
| 3 |
Ag |
3087 |
3009 |
0.00 |
|
|
|
| 4 |
Ag |
3014 |
2937 |
0.00 |
|
|
|
| 5 |
Ag |
1514 |
1476 |
0.00 |
|
|
|
| 6 |
Ag |
1510 |
1472 |
0.00 |
|
|
|
| 7 |
Ag |
1439 |
1402 |
0.00 |
|
|
|
| 8 |
Ag |
1401 |
1365 |
0.00 |
|
|
|
| 9 |
Ag |
1264 |
1232 |
0.00 |
|
|
|
| 10 |
Ag |
1189 |
1158 |
0.00 |
|
|
|
| 11 |
Ag |
1145 |
1116 |
0.00 |
|
|
|
| 12 |
Ag |
1033 |
1007 |
0.00 |
|
|
|
| 13 |
Ag |
843 |
822 |
0.00 |
|
|
|
| 14 |
Ag |
637 |
620 |
0.00 |
|
|
|
| 15 |
Ag |
464 |
452 |
0.00 |
|
|
|
| 16 |
Ag |
322 |
314 |
0.00 |
|
|
|
| 17 |
Ag |
260 |
254 |
0.00 |
|
|
|
| 18 |
Ag |
217 |
211 |
0.00 |
|
|
|
| 19 |
Au |
3121 |
3042 |
55.28 |
|
|
|
| 20 |
Au |
3113 |
3034 |
30.09 |
|
|
|
| 21 |
Au |
3095 |
3016 |
5.35 |
|
|
|
| 22 |
Au |
3013 |
2936 |
26.72 |
|
|
|
| 23 |
Au |
1515 |
1476 |
18.50 |
|
|
|
| 24 |
Au |
1507 |
1469 |
18.62 |
|
|
|
| 25 |
Au |
1440 |
1403 |
31.60 |
|
|
|
| 26 |
Au |
1326 |
1292 |
3.25 |
|
|
|
| 27 |
Au |
1209 |
1178 |
46.62 |
|
|
|
| 28 |
Au |
1098 |
1070 |
13.35 |
|
|
|
| 29 |
Au |
1034 |
1008 |
24.82 |
|
|
|
| 30 |
Au |
983 |
958 |
22.03 |
|
|
|
| 31 |
Au |
576 |
562 |
108.57 |
|
|
|
| 32 |
Au |
340 |
331 |
4.85 |
|
|
|
| 33 |
Au |
320 |
312 |
4.86 |
|
|
|
| 34 |
Au |
231 |
225 |
4.22 |
|
|
|
| 35 |
Au |
191 |
186 |
5.31 |
|
|
|
| 36 |
Au |
62 |
60 |
4.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24868.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 24234.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Cl |
-0.046 |
|
|
|
| 2 |
Cl |
-0.046 |
|
|
|
| 3 |
C |
-0.423 |
|
|
|
| 4 |
C |
-0.423 |
|
|
|
| 5 |
C |
-0.257 |
|
|
|
| 6 |
C |
-0.257 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
H |
0.177 |
|
|
|
| 10 |
H |
0.177 |
|
|
|
| 11 |
H |
0.165 |
|
|
|
| 12 |
H |
0.165 |
|
|
|
| 13 |
H |
0.207 |
|
|
|
| 14 |
H |
0.207 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.357 |
2.800 |
-4.882 |
| y |
2.800 |
-53.036 |
4.814 |
| z |
-4.882 |
4.814 |
-55.642 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.983 |
2.800 |
-4.882 |
| y |
2.800 |
-0.537 |
4.814 |
| z |
-4.882 |
4.814 |
-4.446 |
|
| Polar |
|---|
| 3z2-r2 | -8.891 |
|---|
| x2-y2 | 3.680 |
|---|
| xy | 2.800 |
|---|
| xz | -4.882 |
|---|
| yz | 4.814 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.785 |
-0.637 |
0.880 |
| y |
-0.637 |
9.277 |
-2.925 |
| z |
0.880 |
-2.925 |
11.771 |
<r2> (average value of r
2) Å
2
| <r2> |
186.906 |
| (<r2>)1/2 |
13.671 |