Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -41.062551 |
| Energy at 298.15K | -41.074723 |
| HF Energy | -41.062551 |
| Nuclear repulsion energy | 126.168034 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3091 |
3013 |
101.61 |
|
|
|
| 2 |
A |
3077 |
2998 |
29.51 |
|
|
|
| 3 |
A |
3066 |
2988 |
4.14 |
|
|
|
| 4 |
A |
3065 |
2986 |
67.38 |
|
|
|
| 5 |
A |
3063 |
2985 |
96.55 |
|
|
|
| 6 |
A |
3034 |
2956 |
43.78 |
|
|
|
| 7 |
A |
3030 |
2952 |
19.39 |
|
|
|
| 8 |
A |
3006 |
2930 |
7.75 |
|
|
|
| 9 |
A |
2980 |
2904 |
19.21 |
|
|
|
| 10 |
A |
2975 |
2899 |
82.84 |
|
|
|
| 11 |
A |
2970 |
2895 |
63.43 |
|
|
|
| 12 |
A |
2957 |
2881 |
36.68 |
|
|
|
| 13 |
A |
1703 |
1660 |
0.54 |
|
|
|
| 14 |
A |
1532 |
1493 |
9.33 |
|
|
|
| 15 |
A |
1521 |
1482 |
6.67 |
|
|
|
| 16 |
A |
1517 |
1479 |
3.86 |
|
|
|
| 17 |
A |
1517 |
1478 |
9.16 |
|
|
|
| 18 |
A |
1506 |
1468 |
1.12 |
|
|
|
| 19 |
A |
1505 |
1466 |
10.18 |
|
|
|
| 20 |
A |
1435 |
1399 |
3.40 |
|
|
|
| 21 |
A |
1430 |
1393 |
4.52 |
|
|
|
| 22 |
A |
1382 |
1347 |
1.14 |
|
|
|
| 23 |
A |
1348 |
1314 |
0.53 |
|
|
|
| 24 |
A |
1336 |
1302 |
0.39 |
|
|
|
| 25 |
A |
1315 |
1282 |
0.14 |
|
|
|
| 26 |
A |
1301 |
1268 |
3.88 |
|
|
|
| 27 |
A |
1256 |
1224 |
0.08 |
|
|
|
| 28 |
A |
1182 |
1152 |
0.05 |
|
|
|
| 29 |
A |
1115 |
1087 |
1.12 |
|
|
|
| 30 |
A |
1088 |
1061 |
0.70 |
|
|
|
| 31 |
A |
1079 |
1052 |
0.63 |
|
|
|
| 32 |
A |
1060 |
1033 |
9.30 |
|
|
|
| 33 |
A |
1030 |
1004 |
2.48 |
|
|
|
| 34 |
A |
1005 |
979 |
44.40 |
|
|
|
| 35 |
A |
929 |
905 |
4.64 |
|
|
|
| 36 |
A |
897 |
874 |
2.96 |
|
|
|
| 37 |
A |
877 |
854 |
0.62 |
|
|
|
| 38 |
A |
786 |
766 |
0.86 |
|
|
|
| 39 |
A |
751 |
731 |
2.67 |
|
|
|
| 40 |
A |
526 |
513 |
0.19 |
|
|
|
| 41 |
A |
380 |
371 |
1.44 |
|
|
|
| 42 |
A |
307 |
299 |
0.83 |
|
|
|
| 43 |
A |
279 |
272 |
1.75 |
|
|
|
| 44 |
A |
232 |
226 |
0.04 |
|
|
|
| 45 |
A |
198 |
193 |
0.92 |
|
|
|
| 46 |
A |
140 |
137 |
0.35 |
|
|
|
| 47 |
A |
92 |
90 |
0.08 |
|
|
|
| 48 |
A |
74 |
72 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35971.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 35053.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
3.232 |
0.217 |
-0.159 |
| C2 |
1.832 |
-0.373 |
-0.221 |
| C3 |
0.752 |
0.111 |
0.440 |
| C4 |
-0.651 |
-0.475 |
0.375 |
| C5 |
-1.710 |
0.529 |
-0.175 |
| C6 |
-3.144 |
-0.060 |
-0.190 |
| H7 |
3.264 |
1.103 |
0.489 |
| H8 |
3.957 |
-0.515 |
0.228 |
| H9 |
3.584 |
0.514 |
-1.159 |
| H10 |
1.709 |
-1.260 |
-0.851 |
| H11 |
0.874 |
0.999 |
1.070 |
| H12 |
-0.967 |
-0.792 |
1.384 |
| H13 |
-0.647 |
-1.381 |
-0.252 |
| H14 |
-1.419 |
0.833 |
-1.192 |
| H15 |
-1.694 |
1.444 |
0.439 |
| H16 |
-3.869 |
0.667 |
-0.582 |
| H17 |
-3.196 |
-0.959 |
-0.820 |
| H18 |
-3.466 |
-0.343 |
0.823 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5207 | 2.5537 | 3.9808 | 4.9518 | 6.3822 | 1.0977 | 1.1008 | 1.1008 | 2.2319 | 2.7721 | 4.5866 | 4.1960 | 4.8041 | 5.1112 | 7.1279 | 6.5680 | 6.7926 |
C2 | 1.5207 | | 1.3560 | 2.5562 | 3.6555 | 4.9862 | 2.1755 | 2.1760 | 2.1760 | 1.0950 | 2.1143 | 3.2542 | 2.6759 | 3.6013 | 4.0211 | 5.8066 | 5.0973 | 5.3999 | C3 | 2.5537 | 1.3560 | | 1.5225 | 2.5719 | 3.9504 | 2.7010 | 3.2720 | 3.2770 | 2.1131 | 1.0958 | 2.1594 | 2.1588 | 2.8107 | 2.7852 | 4.7653 | 4.2802 | 4.2594 | C4 | 3.9808 | 2.5562 | 1.5225 | | 1.5595 | 2.5894 | 4.2230 | 4.6105 | 4.6117 | 2.7736 | 2.2326 | 1.1040 | 1.1011 | 2.1809 | 2.1848 | 3.5458 | 2.8524 | 2.8528 | C5 | 4.9518 | 3.6555 | 2.5719 | 1.5595 | | 1.5505 | 5.0506 | 5.7760 | 5.3841 | 3.9179 | 2.9064 | 2.1747 | 2.1873 | 1.1004 | 1.1019 | 2.2015 | 2.2000 | 2.2000 | C6 | 6.3822 | 4.9862 | 3.9504 | 2.5894 | 1.5505 | | 6.5479 | 7.1275 | 6.8212 | 5.0432 | 4.3420 | 2.7844 | 2.8260 | 2.1854 | 2.1818 | 1.0987 | 1.0996 | 1.0998 | H7 | 1.0977 | 2.1755 | 2.7010 | 4.2230 | 5.0506 | 6.5479 | | 1.7791 | 1.7791 | 3.1300 | 2.4620 | 4.7219 | 4.6913 | 4.9830 | 4.9695 | 7.2261 | 6.9061 | 6.8913 | H8 | 1.1008 | 2.1760 | 3.2720 | 4.6105 | 5.7760 | 7.1275 | 1.7791 | | 1.7676 | 2.6021 | 3.5364 | 5.0653 | 4.7084 | 5.7213 | 5.9838 | 7.9557 | 7.2424 | 7.4481 | H9 | 1.1008 | 2.1760 | 3.2770 | 4.6117 | 5.3841 | 6.8212 | 1.7791 | 1.7676 | | 2.5993 | 3.5425 | 5.3746 | 4.7232 | 5.0129 | 5.5917 | 7.4764 | 6.9458 | 7.3726 | H10 | 2.2319 | 1.0950 | 2.1131 | 2.7736 | 3.9179 | 5.0432 | 3.1300 | 2.6021 | 2.5993 | | 3.0816 | 3.5186 | 2.4340 | 3.7795 | 4.5340 | 5.9080 | 4.9144 | 5.5159 | H11 | 2.7721 | 2.1143 | 1.0958 | 2.2326 | 2.9064 | 4.3420 | 2.4620 | 3.5364 | 3.5425 | 3.0816 | | 2.5881 | 3.1183 | 3.2256 | 2.6809 | 5.0334 | 4.8961 | 4.5491 | H12 | 4.5866 | 3.2542 | 2.1594 | 1.1040 | 2.1747 | 2.7844 | 4.7219 | 5.0653 | 5.3746 | 3.5186 | 2.5881 | | 1.7679 | 3.0794 | 2.5339 | 3.7967 | 3.1390 | 2.5999 | H13 | 4.1960 | 2.6759 | 2.1588 | 1.1011 | 2.1873 | 2.8260 | 4.6913 | 4.7084 | 4.7232 | 2.4340 | 3.1183 | 1.7679 | | 2.5260 | 3.0906 | 3.8321 | 2.6455 | 3.1905 | H14 | 4.8041 | 3.6013 | 2.8107 | 2.1809 | 1.1004 | 2.1854 | 4.9830 | 5.7213 | 5.0129 | 3.7795 | 3.2256 | 3.0794 | 2.5260 | | 1.7635 | 2.5302 | 2.5508 | 3.1034 | H15 | 5.1112 | 4.0211 | 2.7852 | 2.1848 | 1.1019 | 2.1818 | 4.9695 | 5.9838 | 5.5917 | 4.5340 | 2.6809 | 2.5339 | 3.0906 | 1.7635 | | 2.5257 | 3.1013 | 2.5456 | H16 | 7.1279 | 5.8066 | 4.7653 | 3.5458 | 2.2015 | 1.0987 | 7.2261 | 7.9557 | 7.4764 | 5.9080 | 5.0334 | 3.7967 | 3.8321 | 2.5302 | 2.5257 | | 1.7761 | 1.7764 | H17 | 6.5680 | 5.0973 | 4.2802 | 2.8524 | 2.2000 | 1.0996 | 6.9061 | 7.2424 | 6.9458 | 4.9144 | 4.8961 | 3.1390 | 2.6455 | 2.5508 | 3.1013 | 1.7761 | | 1.7755 | H18 | 6.7926 | 5.3999 | 4.2594 | 2.8528 | 2.2000 | 1.0998 | 6.8913 | 7.4481 | 7.3726 | 5.5159 | 4.5491 | 2.5999 | 3.1905 | 3.1034 | 2.5456 | 1.7764 | 1.7755 | |
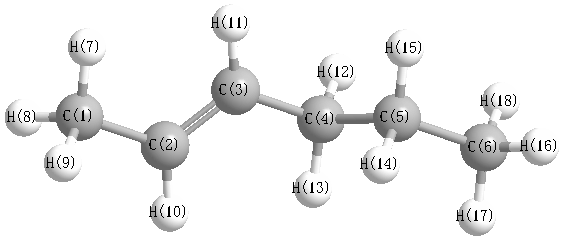 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.081 |
|
C1 |
C2 |
H10 |
116.193 |
| C2 |
C1 |
H7 |
111.348 |
|
C2 |
C1 |
H8 |
111.203 |
| C2 |
C1 |
H9 |
111.205 |
|
C2 |
C3 |
C4 |
125.157 |
| C2 |
C3 |
H11 |
118.780 |
|
C3 |
C2 |
H10 |
118.725 |
| C3 |
C4 |
C5 |
113.122 |
|
C3 |
C4 |
H12 |
109.577 |
| C3 |
C4 |
H13 |
109.700 |
|
C4 |
C3 |
H11 |
116.061 |
| C4 |
C5 |
C6 |
112.734 |
|
C4 |
C5 |
H14 |
108.928 |
| C4 |
C5 |
H15 |
109.148 |
|
C5 |
C4 |
H12 |
108.252 |
| C5 |
C4 |
H13 |
109.385 |
|
C5 |
C6 |
H16 |
111.260 |
| C5 |
C6 |
H17 |
111.085 |
|
C5 |
C6 |
H18 |
111.075 |
| C6 |
C5 |
H14 |
109.894 |
|
C6 |
C5 |
H15 |
109.529 |
| H7 |
C1 |
H8 |
108.041 |
|
H7 |
C1 |
H9 |
108.041 |
| H8 |
C1 |
H9 |
106.813 |
|
H12 |
C4 |
H13 |
106.587 |
| H14 |
C5 |
H15 |
106.406 |
|
H16 |
C6 |
H17 |
107.790 |
| H16 |
C6 |
H18 |
107.807 |
|
H17 |
C6 |
H18 |
107.659 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.595 |
|
|
|
| 2 |
C |
-0.019 |
|
|
|
| 3 |
C |
-0.002 |
|
|
|
| 4 |
C |
-0.361 |
|
|
|
| 5 |
C |
-0.203 |
|
|
|
| 6 |
C |
-0.472 |
|
|
|
| 7 |
H |
0.159 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
| 9 |
H |
0.152 |
|
|
|
| 10 |
H |
0.130 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
H |
0.125 |
|
|
|
| 13 |
H |
0.131 |
|
|
|
| 14 |
H |
0.129 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
| 16 |
H |
0.146 |
|
|
|
| 17 |
H |
0.139 |
|
|
|
| 18 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.023 |
0.043 |
-0.048 |
0.068 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.319 |
-0.039 |
-0.711 |
| y |
-0.039 |
-40.409 |
1.362 |
| z |
-0.711 |
1.362 |
-40.569 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.169 |
-0.039 |
-0.711 |
| y |
-0.039 |
-0.464 |
1.362 |
| z |
-0.711 |
1.362 |
-0.705 |
|
| Polar |
|---|
| 3z2-r2 | -1.410 |
|---|
| x2-y2 | 1.089 |
|---|
| xy | -0.039 |
|---|
| xz | -0.711 |
|---|
| yz | 1.362 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.363 |
-0.349 |
-1.015 |
| y |
-0.349 |
9.222 |
0.664 |
| z |
-1.015 |
0.664 |
9.035 |
<r2> (average value of r
2) Å
2
| <r2> |
246.368 |
| (<r2>)1/2 |
15.696 |
