Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -85.599560 |
| Energy at 298.15K | |
| HF Energy | -85.599560 |
| Nuclear repulsion energy | 105.426448 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3073 |
20.26 |
32.19 |
0.66 |
0.80 |
| 2 |
A |
3125 |
3045 |
15.00 |
99.33 |
0.31 |
0.47 |
| 3 |
A |
3066 |
2988 |
13.65 |
110.37 |
0.07 |
0.14 |
| 4 |
A |
1497 |
1459 |
8.24 |
8.07 |
0.75 |
0.86 |
| 5 |
A |
1432 |
1396 |
6.20 |
3.80 |
0.75 |
0.86 |
| 6 |
A |
1375 |
1340 |
18.42 |
6.71 |
0.75 |
0.86 |
| 7 |
A |
1316 |
1282 |
11.33 |
3.05 |
0.73 |
0.84 |
| 8 |
A |
1228 |
1197 |
8.24 |
10.07 |
0.73 |
0.85 |
| 9 |
A |
1125 |
1097 |
17.89 |
4.74 |
0.71 |
0.83 |
| 10 |
A |
1094 |
1066 |
55.82 |
4.91 |
0.52 |
0.69 |
| 11 |
A |
1010 |
984 |
209.47 |
0.81 |
0.27 |
0.42 |
| 12 |
A |
993 |
968 |
14.94 |
11.97 |
0.39 |
0.56 |
| 13 |
A |
878 |
855 |
59.55 |
7.57 |
0.35 |
0.52 |
| 14 |
A |
527 |
513 |
6.00 |
3.63 |
0.33 |
0.50 |
| 15 |
A |
440 |
429 |
27.55 |
1.11 |
0.73 |
0.84 |
| 16 |
A |
390 |
380 |
7.17 |
2.52 |
0.59 |
0.74 |
| 17 |
A |
238 |
232 |
14.15 |
0.15 |
0.59 |
0.74 |
| 18 |
A |
99 |
97 |
14.34 |
0.10 |
0.60 |
0.75 |
Unscaled Zero Point Vibrational Energy (zpe) 11492.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 11199.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
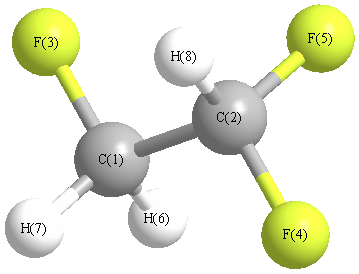
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.782 |
-0.568 |
-0.353 |
| C2 |
0.450 |
0.017 |
0.338 |
| F3 |
-1.951 |
0.093 |
0.190 |
| F4 |
1.567 |
-0.820 |
0.019 |
| F5 |
0.743 |
1.308 |
-0.184 |
| H6 |
-0.749 |
-0.378 |
-1.430 |
| H7 |
-0.869 |
-1.636 |
-0.135 |
| H8 |
0.379 |
0.100 |
1.426 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5292 | 1.4489 | 2.3920 | 2.4232 | 1.0935 | 1.0939 | 2.2271 |
C2 | 1.5292 | | 2.4073 | 1.4318 | 1.4227 | 2.1726 | 2.1671 | 1.0931 | F3 | 1.4489 | 2.4073 | | 3.6391 | 2.9790 | 2.0713 | 2.0655 | 2.6378 | F4 | 2.3920 | 1.4318 | 3.6391 | | 2.2911 | 2.7681 | 2.5738 | 2.0583 | F5 | 2.4232 | 1.4227 | 2.9790 | 2.2911 | | 2.5732 | 3.3565 | 2.0446 | H6 | 1.0935 | 2.1726 | 2.0713 | 2.7681 | 2.5732 | | 1.8096 | 3.1074 | H7 | 1.0939 | 2.1671 | 2.0655 | 2.5738 | 3.3565 | 1.8096 | | 2.6471 | H8 | 2.2271 | 1.0931 | 2.6378 | 2.0583 | 2.0446 | 3.1074 | 2.6471 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.721 |
|
C1 |
C2 |
F5 |
110.296 |
| C1 |
C2 |
H8 |
115.254 |
|
C2 |
C1 |
F3 |
107.838 |
| C2 |
C1 |
H6 |
110.770 |
|
C2 |
C1 |
H7 |
110.305 |
| F3 |
C1 |
H6 |
108.309 |
|
F3 |
C1 |
H7 |
107.830 |
| F4 |
C2 |
F5 |
106.760 |
|
F4 |
C2 |
H8 |
108.470 |
| F5 |
C2 |
H8 |
108.006 |
|
H6 |
C1 |
H7 |
111.644 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.064 |
|
|
|
| 2 |
C |
0.208 |
|
|
|
| 3 |
F |
-0.255 |
|
|
|
| 4 |
F |
-0.214 |
|
|
|
| 5 |
F |
-0.198 |
|
|
|
| 6 |
H |
0.182 |
|
|
|
| 7 |
H |
0.174 |
|
|
|
| 8 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.493 |
-1.834 |
0.307 |
1.924 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.215 |
2.081 |
2.200 |
| y |
2.081 |
-28.950 |
0.817 |
| z |
2.200 |
0.817 |
-24.480 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.500 |
2.081 |
2.200 |
| y |
2.081 |
0.897 |
0.817 |
| z |
2.200 |
0.817 |
7.602 |
|
| Polar |
|---|
| 3z2-r2 | 15.205 |
|---|
| x2-y2 | -6.264 |
|---|
| xy | 2.081 |
|---|
| xz | 2.200 |
|---|
| yz | 0.817 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.756 |
-0.170 |
-0.117 |
| y |
-0.170 |
3.720 |
0.023 |
| z |
-0.117 |
0.023 |
3.364 |
<r2> (average value of r
2) Å
2
| <r2> |
97.289 |
| (<r2>)1/2 |
9.864 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -85.595791 |
| Energy at 298.15K | |
| HF Energy | -85.595791 |
| Nuclear repulsion energy | 107.107657 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3086 |
3008 |
42.16 |
108.49 |
0.12 |
0.22 |
| 2 |
A' |
3048 |
2970 |
11.27 |
94.54 |
0.15 |
0.26 |
| 3 |
A' |
1495 |
1456 |
9.21 |
7.89 |
0.75 |
0.86 |
| 4 |
A' |
1420 |
1384 |
4.37 |
2.09 |
0.67 |
0.80 |
| 5 |
A' |
1379 |
1344 |
24.74 |
6.81 |
0.74 |
0.85 |
| 6 |
A' |
1130 |
1101 |
82.95 |
6.74 |
0.36 |
0.53 |
| 7 |
A' |
1020 |
994 |
16.67 |
4.46 |
0.74 |
0.85 |
| 8 |
A' |
844 |
823 |
43.79 |
10.39 |
0.07 |
0.13 |
| 9 |
A' |
704 |
686 |
58.63 |
4.64 |
0.59 |
0.74 |
| 10 |
A' |
453 |
442 |
16.24 |
2.62 |
0.74 |
0.85 |
| 11 |
A' |
224 |
218 |
2.65 |
0.36 |
0.56 |
0.72 |
| 12 |
A" |
3127 |
3047 |
17.27 |
54.43 |
0.75 |
0.86 |
| 13 |
A" |
1365 |
1330 |
18.75 |
2.16 |
0.75 |
0.86 |
| 14 |
A" |
1242 |
1210 |
11.05 |
19.63 |
0.75 |
0.86 |
| 15 |
A" |
1091 |
1063 |
68.03 |
0.74 |
0.75 |
0.86 |
| 16 |
A" |
908 |
885 |
100.99 |
6.33 |
0.75 |
0.86 |
| 17 |
A" |
347 |
339 |
0.26 |
0.49 |
0.75 |
0.86 |
| 18 |
A" |
105 |
102 |
3.41 |
0.18 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11493.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 11200.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.775 |
-0.839 |
0.000 |
| C2 |
0.383 |
0.638 |
0.000 |
| F3 |
-0.403 |
-1.668 |
0.000 |
| F4 |
-0.403 |
0.948 |
1.145 |
| F5 |
-0.403 |
0.948 |
-1.145 |
| H6 |
1.349 |
-1.071 |
-0.904 |
| H7 |
1.349 |
-1.071 |
0.904 |
| H8 |
1.249 |
1.310 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5279 | 1.4401 | 2.4276 | 2.4276 | 1.0954 | 1.0954 | 2.2012 |
C2 | 1.5279 | | 2.4360 | 1.4238 | 1.4238 | 2.1606 | 2.1606 | 1.0965 | F3 | 1.4401 | 2.4360 | | 2.8553 | 2.8553 | 2.0599 | 2.0599 | 3.4058 | F4 | 2.4276 | 1.4238 | 2.8553 | | 2.2909 | 3.3681 | 2.6840 | 2.0434 | F5 | 2.4276 | 1.4238 | 2.8553 | 2.2909 | | 2.6840 | 3.3681 | 2.0434 | H6 | 1.0954 | 2.1606 | 2.0599 | 3.3681 | 2.6840 | | 1.8072 | 2.5486 | H7 | 1.0954 | 2.1606 | 2.0599 | 2.6840 | 3.3681 | 1.8072 | | 2.5486 | H8 | 2.2012 | 1.0965 | 3.4058 | 2.0434 | 2.0434 | 2.5486 | 2.5486 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.608 |
|
C1 |
C2 |
F5 |
110.608 |
| C1 |
C2 |
H8 |
112.981 |
|
C2 |
C1 |
F3 |
110.282 |
| C2 |
C1 |
H6 |
109.795 |
|
C2 |
C1 |
H7 |
109.795 |
| F3 |
C1 |
H6 |
107.892 |
|
F3 |
C1 |
H7 |
107.892 |
| F4 |
C2 |
F5 |
107.133 |
|
F4 |
C2 |
H8 |
107.636 |
| F5 |
C2 |
H8 |
107.636 |
|
H6 |
C1 |
H7 |
111.147 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.058 |
|
|
|
| 2 |
C |
0.215 |
|
|
|
| 3 |
F |
-0.240 |
|
|
|
| 4 |
F |
-0.199 |
|
|
|
| 5 |
F |
-0.199 |
|
|
|
| 6 |
H |
0.169 |
|
|
|
| 7 |
H |
0.169 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.317 |
0.105 |
0.000 |
4.318 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.153 |
-0.438 |
0.000 |
| y |
-0.438 |
-29.973 |
0.000 |
| z |
0.000 |
0.000 |
-29.847 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.757 |
-0.438 |
0.000 |
| y |
-0.438 |
-2.473 |
0.000 |
| z |
0.000 |
0.000 |
-2.284 |
|
| Polar |
|---|
| 3z2-r2 | -4.568 |
|---|
| x2-y2 | 4.820 |
|---|
| xy | -0.438 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.675 |
0.053 |
0.000 |
| y |
0.053 |
3.538 |
0.000 |
| z |
0.000 |
0.000 |
3.714 |
<r2> (average value of r
2) Å
2
| <r2> |
89.797 |
| (<r2>)1/2 |
9.476 |
