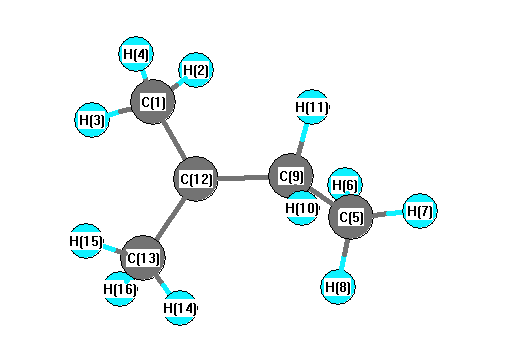
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3076 |
2997 |
68.02 |
|
|
|
| 2 |
A |
3069 |
2990 |
28.45 |
|
|
|
| 3 |
A |
3065 |
2987 |
108.17 |
|
|
|
| 4 |
A |
3056 |
2978 |
26.07 |
|
|
|
| 5 |
A |
3016 |
2939 |
44.88 |
|
|
|
| 6 |
A |
3005 |
2929 |
51.90 |
|
|
|
| 7 |
A |
3002 |
2925 |
18.57 |
|
|
|
| 8 |
A |
2984 |
2908 |
43.65 |
|
|
|
| 9 |
A |
2899 |
2825 |
116.03 |
|
|
|
| 10 |
A |
2891 |
2817 |
51.84 |
|
|
|
| 11 |
A |
2876 |
2803 |
56.74 |
|
|
|
| 12 |
A |
1528 |
1489 |
4.57 |
|
|
|
| 13 |
A |
1522 |
1484 |
8.56 |
|
|
|
| 14 |
A |
1515 |
1476 |
4.13 |
|
|
|
| 15 |
A |
1509 |
1470 |
13.26 |
|
|
|
| 16 |
A |
1508 |
1469 |
9.66 |
|
|
|
| 17 |
A |
1496 |
1458 |
1.89 |
|
|
|
| 18 |
A |
1495 |
1457 |
1.84 |
|
|
|
| 19 |
A |
1434 |
1398 |
3.28 |
|
|
|
| 20 |
A |
1427 |
1390 |
7.44 |
|
|
|
| 21 |
A |
1417 |
1381 |
6.37 |
|
|
|
| 22 |
A |
1377 |
1342 |
0.70 |
|
|
|
| 23 |
A |
1304 |
1271 |
1.44 |
|
|
|
| 24 |
A |
1284 |
1251 |
8.45 |
|
|
|
| 25 |
A |
1247 |
1215 |
5.03 |
|
|
|
| 26 |
A |
1105 |
1077 |
1.28 |
|
|
|
| 27 |
A |
1062 |
1035 |
3.05 |
|
|
|
| 28 |
A |
1040 |
1013 |
4.50 |
|
|
|
| 29 |
A |
1005 |
979 |
0.82 |
|
|
|
| 30 |
A |
986 |
961 |
1.77 |
|
|
|
| 31 |
A |
948 |
924 |
0.50 |
|
|
|
| 32 |
A |
931 |
907 |
0.35 |
|
|
|
| 33 |
A |
780 |
760 |
2.01 |
|
|
|
| 34 |
A |
717 |
699 |
1.69 |
|
|
|
| 35 |
A |
440 |
429 |
0.94 |
|
|
|
| 36 |
A |
369 |
360 |
0.06 |
|
|
|
| 37 |
A |
311 |
303 |
2.01 |
|
|
|
| 38 |
A |
252 |
246 |
0.45 |
|
|
|
| 39 |
A |
196 |
191 |
1.22 |
|
|
|
| 40 |
A |
121 |
118 |
0.02 |
|
|
|
| 41 |
A |
109 |
106 |
0.09 |
|
|
|
| 42 |
A |
28 |
27 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31700.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30891.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.579 |
|
|
|
| 2 |
H |
0.160 |
|
|
|
| 3 |
H |
0.148 |
|
|
|
| 4 |
H |
0.139 |
|
|
|
| 5 |
C |
-0.487 |
|
|
|
| 6 |
H |
0.143 |
|
|
|
| 7 |
H |
0.141 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
C |
-0.266 |
|
|
|
| 10 |
H |
0.132 |
|
|
|
| 11 |
H |
0.122 |
|
|
|
| 12 |
C |
0.336 |
|
|
|
| 13 |
C |
-0.577 |
|
|
|
| 14 |
H |
0.152 |
|
|
|
| 15 |
H |
0.142 |
|
|
|
| 16 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.054 |
-0.012 |
0.253 |
0.259 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.576 |
-0.179 |
0.197 |
| y |
-0.179 |
-34.312 |
0.087 |
| z |
0.197 |
0.087 |
-35.230 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.196 |
-0.179 |
0.197 |
| y |
-0.179 |
0.591 |
0.087 |
| z |
0.197 |
0.087 |
-0.787 |
|
| Polar |
|---|
| 3z2-r2 | -1.574 |
|---|
| x2-y2 | -0.264 |
|---|
| xy | -0.179 |
|---|
| xz | 0.197 |
|---|
| yz | 0.087 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.013 |
-0.001 |
0.091 |
| y |
-0.001 |
9.073 |
0.002 |
| z |
0.091 |
0.002 |
7.833 |
<r2> (average value of r
2) Å
2
| <r2> |
135.411 |
| (<r2>)1/2 |
11.637 |