Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -45.317377 |
| Energy at 298.15K | -45.325136 |
| HF Energy | -45.317377 |
| Nuclear repulsion energy | 73.478708 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3080 |
3002 |
52.52 |
|
|
|
| 2 |
A' |
3020 |
2943 |
68.03 |
|
|
|
| 3 |
A' |
3012 |
2935 |
8.64 |
|
|
|
| 4 |
A' |
2984 |
2908 |
34.94 |
|
|
|
| 5 |
A' |
1537 |
1498 |
10.30 |
|
|
|
| 6 |
A' |
1522 |
1483 |
1.43 |
|
|
|
| 7 |
A' |
1515 |
1476 |
0.92 |
|
|
|
| 8 |
A' |
1437 |
1400 |
10.05 |
|
|
|
| 9 |
A' |
1415 |
1379 |
6.96 |
|
|
|
| 10 |
A' |
1341 |
1307 |
1.21 |
|
|
|
| 11 |
A' |
1142 |
1113 |
0.39 |
|
|
|
| 12 |
A' |
1041 |
1015 |
7.86 |
|
|
|
| 13 |
A' |
966 |
942 |
87.24 |
|
|
|
| 14 |
A' |
891 |
868 |
15.19 |
|
|
|
| 15 |
A' |
428 |
417 |
7.73 |
|
|
|
| 16 |
A' |
258 |
251 |
5.80 |
|
|
|
| 17 |
A" |
3101 |
3022 |
109.31 |
|
|
|
| 18 |
A" |
3070 |
2992 |
20.92 |
|
|
|
| 19 |
A" |
3048 |
2971 |
2.91 |
|
|
|
| 20 |
A" |
1525 |
1486 |
9.37 |
|
|
|
| 21 |
A" |
1313 |
1279 |
0.43 |
|
|
|
| 22 |
A" |
1254 |
1222 |
0.01 |
|
|
|
| 23 |
A" |
1172 |
1142 |
1.21 |
|
|
|
| 24 |
A" |
900 |
877 |
1.62 |
|
|
|
| 25 |
A" |
777 |
757 |
2.78 |
|
|
|
| 26 |
A" |
226 |
220 |
0.00 |
|
|
|
| 27 |
A" |
112 |
109 |
5.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21042.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 20506.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.091 |
-0.796 |
0.000 |
| C2 |
0.000 |
0.736 |
0.000 |
| C3 |
-1.475 |
1.220 |
0.000 |
| F4 |
1.501 |
-1.197 |
0.000 |
| H5 |
-0.354 |
-1.240 |
0.900 |
| H6 |
-0.354 |
-1.240 |
-0.900 |
| H7 |
0.526 |
1.122 |
-0.884 |
| H8 |
0.526 |
1.122 |
0.884 |
| H9 |
-1.520 |
2.316 |
0.000 |
| H10 |
-2.015 |
0.864 |
-0.889 |
| H11 |
-2.015 |
0.864 |
0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5340 | 2.5526 | 1.4657 | 1.0975 | 1.0975 | 2.1557 | 2.1557 | 3.5043 | 2.8249 | 2.8249 |
C2 | 1.5340 | | 1.5525 | 2.4467 | 2.1994 | 2.1994 | 1.0983 | 1.0983 | 2.1929 | 2.2060 | 2.2060 | C3 | 2.5526 | 1.5525 | | 3.8336 | 2.8493 | 2.8493 | 2.1891 | 2.1891 | 1.0971 | 1.0991 | 1.0991 | F4 | 1.4657 | 2.4467 | 3.8336 | | 2.0618 | 2.0618 | 2.6661 | 2.6661 | 4.6332 | 4.1711 | 4.1711 | H5 | 1.0975 | 2.1994 | 2.8493 | 2.0618 | | 1.7990 | 3.0873 | 2.5204 | 3.8492 | 3.2224 | 2.6808 | H6 | 1.0975 | 2.1994 | 2.8493 | 2.0618 | 1.7990 | | 2.5204 | 3.0873 | 3.8492 | 2.6808 | 3.2224 | H7 | 2.1557 | 1.0983 | 2.1891 | 2.6661 | 3.0873 | 2.5204 | | 1.7671 | 2.5281 | 2.5537 | 3.1083 | H8 | 2.1557 | 1.0983 | 2.1891 | 2.6661 | 2.5204 | 3.0873 | 1.7671 | | 2.5281 | 3.1083 | 2.5537 | H9 | 3.5043 | 2.1929 | 1.0971 | 4.6332 | 3.8492 | 3.8492 | 2.5281 | 2.5281 | | 1.7731 | 1.7731 | H10 | 2.8249 | 2.2060 | 1.0991 | 4.1711 | 3.2224 | 2.6808 | 2.5537 | 3.1083 | 1.7731 | | 1.7771 | H11 | 2.8249 | 2.2060 | 1.0991 | 4.1711 | 2.6808 | 3.2224 | 3.1083 | 2.5537 | 1.7731 | 1.7771 | |
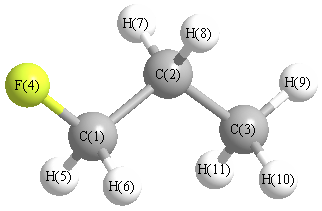 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.590 |
|
C1 |
C2 |
H7 |
108.832 |
| C1 |
C2 |
H8 |
108.832 |
|
C2 |
C1 |
F4 |
109.278 |
| C2 |
C1 |
H5 |
112.337 |
|
C2 |
C1 |
H6 |
112.337 |
| C2 |
C3 |
H9 |
110.538 |
|
C2 |
C3 |
H10 |
111.453 |
| C2 |
C3 |
H11 |
111.453 |
|
C3 |
C2 |
H7 |
110.174 |
| C3 |
C2 |
H8 |
110.174 |
|
F4 |
C1 |
H5 |
106.210 |
| F4 |
C1 |
H6 |
106.210 |
|
H5 |
C1 |
H6 |
110.096 |
| H7 |
C2 |
H8 |
107.119 |
|
H9 |
C3 |
H10 |
107.674 |
| H9 |
C3 |
H11 |
107.674 |
|
H10 |
C3 |
H11 |
107.879 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.031 |
|
|
|
| 2 |
C |
-0.215 |
|
|
|
| 3 |
C |
-0.488 |
|
|
|
| 4 |
F |
-0.288 |
|
|
|
| 5 |
H |
0.133 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
H |
0.151 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
| 10 |
H |
0.146 |
|
|
|
| 11 |
H |
0.146 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.405 |
1.057 |
0.000 |
2.627 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.709 |
3.057 |
0.000 |
| y |
3.057 |
-26.315 |
0.000 |
| z |
0.000 |
0.000 |
-24.740 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.181 |
3.057 |
0.000 |
| y |
3.057 |
0.910 |
0.000 |
| z |
0.000 |
0.000 |
3.271 |
|
| Polar |
|---|
| 3z2-r2 | 6.543 |
|---|
| x2-y2 | -3.394 |
|---|
| xy | 3.057 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.683 |
-0.349 |
0.000 |
| y |
-0.349 |
5.507 |
0.000 |
| z |
0.000 |
0.000 |
5.063 |
<r2> (average value of r
2) Å
2
| <r2> |
90.438 |
| (<r2>)1/2 |
9.510 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -45.317614 |
| Energy at 298.15K | |
| HF Energy | -45.317614 |
| Nuclear repulsion energy | 74.804904 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3098 |
3019 |
71.94 |
|
|
|
| 2 |
A |
3095 |
3016 |
39.01 |
|
|
|
| 3 |
A |
3072 |
2994 |
63.53 |
|
|
|
| 4 |
A |
3041 |
2963 |
21.19 |
|
|
|
| 5 |
A |
3021 |
2944 |
38.21 |
|
|
|
| 6 |
A |
2991 |
2915 |
21.84 |
|
|
|
| 7 |
A |
2985 |
2908 |
45.68 |
|
|
|
| 8 |
A |
1531 |
1492 |
9.92 |
|
|
|
| 9 |
A |
1524 |
1485 |
7.97 |
|
|
|
| 10 |
A |
1513 |
1474 |
4.26 |
|
|
|
| 11 |
A |
1498 |
1460 |
4.46 |
|
|
|
| 12 |
A |
1435 |
1399 |
7.65 |
|
|
|
| 13 |
A |
1404 |
1369 |
10.33 |
|
|
|
| 14 |
A |
1391 |
1355 |
0.68 |
|
|
|
| 15 |
A |
1297 |
1264 |
0.48 |
|
|
|
| 16 |
A |
1246 |
1214 |
0.56 |
|
|
|
| 17 |
A |
1163 |
1133 |
1.62 |
|
|
|
| 18 |
A |
1119 |
1090 |
4.04 |
|
|
|
| 19 |
A |
1073 |
1045 |
16.95 |
|
|
|
| 20 |
A |
941 |
917 |
39.52 |
|
|
|
| 21 |
A |
913 |
889 |
33.53 |
|
|
|
| 22 |
A |
861 |
839 |
9.48 |
|
|
|
| 23 |
A |
766 |
746 |
2.34 |
|
|
|
| 24 |
A |
460 |
448 |
5.64 |
|
|
|
| 25 |
A |
307 |
299 |
1.95 |
|
|
|
| 26 |
A |
211 |
206 |
2.21 |
|
|
|
| 27 |
A |
132 |
128 |
4.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21041.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 20505.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.790 |
0.553 |
0.312 |
| C2 |
-0.612 |
0.655 |
-0.302 |
| C3 |
-1.553 |
-0.499 |
0.126 |
| F4 |
1.441 |
-0.671 |
-0.171 |
| H5 |
1.441 |
1.381 |
0.008 |
| H6 |
0.763 |
0.470 |
1.406 |
| H7 |
-0.514 |
0.678 |
-1.396 |
| H8 |
-1.037 |
1.625 |
0.002 |
| H9 |
-2.536 |
-0.398 |
-0.353 |
| H10 |
-1.130 |
-1.468 |
-0.160 |
| H11 |
-1.707 |
-0.503 |
1.215 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5335 | 2.5754 | 1.4686 | 1.0967 | 1.0976 | 2.1527 | 2.1412 | 3.5225 | 2.8280 | 2.8575 |
C2 | 1.5335 | | 1.5492 | 2.4478 | 2.1998 | 2.2003 | 1.0991 | 1.1023 | 2.1938 | 2.1903 | 2.1995 | C3 | 2.5754 | 1.5492 | | 3.0145 | 3.5380 | 2.8187 | 2.1863 | 2.1896 | 1.0977 | 1.0961 | 1.0995 | F4 | 1.4686 | 2.4478 | 3.0145 | | 2.0603 | 2.0612 | 2.6732 | 3.3836 | 3.9909 | 2.6926 | 3.4442 | H5 | 1.0967 | 2.1998 | 3.5380 | 2.0603 | | 1.8015 | 2.5081 | 2.4907 | 4.3720 | 3.8423 | 3.8625 | H6 | 1.0976 | 2.2003 | 2.8187 | 2.0612 | 1.8015 | | 3.0867 | 2.5594 | 3.8384 | 3.1300 | 2.6620 | H7 | 2.1527 | 1.0991 | 2.1863 | 2.6732 | 2.5081 | 3.0867 | | 1.7677 | 2.5162 | 2.5521 | 3.1034 | H8 | 2.1412 | 1.1023 | 2.1896 | 3.3836 | 2.4907 | 2.5594 | 1.7677 | | 2.5428 | 3.0996 | 2.5394 | H9 | 3.5225 | 2.1938 | 1.0977 | 3.9909 | 4.3720 | 3.8384 | 2.5162 | 2.5428 | | 1.7773 | 1.7764 | H10 | 2.8280 | 2.1903 | 1.0961 | 2.6926 | 3.8423 | 3.1300 | 2.5521 | 3.0996 | 1.7773 | | 1.7766 | H11 | 2.8575 | 2.1995 | 1.0995 | 3.4442 | 3.8625 | 2.6620 | 3.1034 | 2.5394 | 1.7764 | 1.7766 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.319 |
|
C1 |
C2 |
H7 |
108.586 |
| C1 |
C2 |
H8 |
107.526 |
|
C2 |
C1 |
F4 |
109.223 |
| C2 |
C1 |
H5 |
112.457 |
|
C2 |
C1 |
H6 |
112.440 |
| C2 |
C3 |
H9 |
110.806 |
|
C2 |
C3 |
H10 |
110.620 |
| C2 |
C3 |
H11 |
111.147 |
|
C3 |
C2 |
H7 |
110.133 |
| C3 |
C2 |
H8 |
110.199 |
|
F4 |
C1 |
H5 |
105.951 |
| F4 |
C1 |
H6 |
105.974 |
|
H5 |
C1 |
H6 |
110.374 |
| H7 |
C2 |
H8 |
106.837 |
|
H9 |
C3 |
H10 |
108.230 |
| H9 |
C3 |
H11 |
107.895 |
|
H10 |
C3 |
H11 |
108.025 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.031 |
|
|
|
| 2 |
C |
-0.220 |
|
|
|
| 3 |
C |
-0.473 |
|
|
|
| 4 |
F |
-0.293 |
|
|
|
| 5 |
H |
0.142 |
|
|
|
| 6 |
H |
0.133 |
|
|
|
| 7 |
H |
0.152 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
H |
0.150 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
| 11 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.353 |
1.958 |
0.639 |
2.464 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.862 |
2.510 |
0.959 |
| y |
2.510 |
-25.720 |
-0.026 |
| z |
0.959 |
-0.026 |
-24.897 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.554 |
2.510 |
0.959 |
| y |
2.510 |
0.660 |
-0.026 |
| z |
0.959 |
-0.026 |
1.894 |
|
| Polar |
|---|
| 3z2-r2 | 3.788 |
|---|
| x2-y2 | -2.143 |
|---|
| xy | 2.510 |
|---|
| xz | 0.959 |
|---|
| yz | -0.026 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.660 |
0.139 |
0.009 |
| y |
0.139 |
5.398 |
0.017 |
| z |
0.009 |
0.017 |
5.164 |
<r2> (average value of r
2) Å
2
| <r2> |
80.612 |
| (<r2>)1/2 |
8.978 |
