Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -70.850212 |
| Energy at 298.15K | -70.855354 |
| Nuclear repulsion energy | 122.728118 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3719 |
3624 |
50.05 |
|
|
|
| 2 |
A' |
3614 |
3522 |
49.81 |
|
|
|
| 3 |
A' |
3559 |
3468 |
52.95 |
|
|
|
| 4 |
A' |
1692 |
1649 |
148.07 |
|
|
|
| 5 |
A' |
1649 |
1607 |
358.86 |
|
|
|
| 6 |
A' |
1623 |
1582 |
142.91 |
|
|
|
| 7 |
A' |
1393 |
1357 |
1.95 |
|
|
|
| 8 |
A' |
1297 |
1264 |
86.12 |
|
|
|
| 9 |
A' |
1110 |
1082 |
176.02 |
|
|
|
| 10 |
A' |
1088 |
1060 |
115.53 |
|
|
|
| 11 |
A' |
746 |
727 |
2.47 |
|
|
|
| 12 |
A' |
580 |
565 |
75.07 |
|
|
|
| 13 |
A' |
518 |
505 |
1.25 |
|
|
|
| 14 |
A' |
400 |
389 |
7.64 |
|
|
|
| 15 |
A' |
259 |
252 |
14.35 |
|
|
|
| 16 |
A" |
786 |
766 |
0.78 |
|
|
|
| 17 |
A" |
648 |
631 |
506.68 |
|
|
|
| 18 |
A" |
622 |
606 |
24.16 |
|
|
|
| 19 |
A" |
605 |
590 |
0.01 |
|
|
|
| 20 |
A" |
407 |
397 |
15.17 |
|
|
|
| 21 |
A" |
65 |
64 |
9.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13188.8 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12852.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.757 |
0.000 |
| C2 |
-0.051 |
-0.795 |
0.000 |
| O3 |
-1.122 |
-1.449 |
0.000 |
| O4 |
1.071 |
1.403 |
0.000 |
| O5 |
-1.245 |
1.335 |
0.000 |
| N6 |
1.215 |
-1.333 |
0.000 |
| H7 |
1.317 |
-2.339 |
0.000 |
| H8 |
2.028 |
-0.731 |
0.000 |
| H9 |
-1.174 |
2.318 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5526 | 2.4748 | 1.2515 | 1.3730 | 2.4175 | 3.3640 | 2.5147 | 1.9533 |
C2 | 1.5526 | | 1.2549 | 2.4688 | 2.4421 | 1.3762 | 2.0629 | 2.0803 | 3.3093 | O3 | 2.4748 | 1.2549 | | 3.5986 | 2.7869 | 2.3406 | 2.5970 | 3.2313 | 3.7673 | O4 | 1.2515 | 2.4688 | 3.5986 | | 2.3174 | 2.7406 | 3.7504 | 2.3386 | 2.4240 | O5 | 1.3730 | 2.4421 | 2.7869 | 2.3174 | | 3.6297 | 4.4793 | 3.8704 | 0.9853 | N6 | 2.4175 | 1.3762 | 2.3406 | 2.7406 | 3.6297 | | 1.0107 | 1.0118 | 4.3633 | H7 | 3.3640 | 2.0629 | 2.5970 | 3.7504 | 4.4793 | 1.0107 | | 1.7583 | 5.2810 | H8 | 2.5147 | 2.0803 | 3.2313 | 2.3386 | 3.8704 | 1.0118 | 1.7583 | | 4.4207 | H9 | 1.9533 | 3.3093 | 3.7673 | 2.4240 | 0.9853 | 4.3633 | 5.2810 | 4.4207 | |
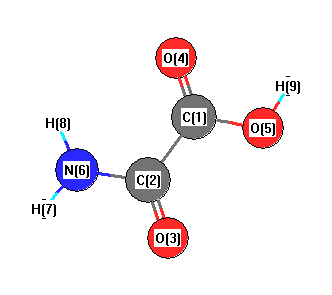 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
123.294 |
|
C1 |
C2 |
N6 |
111.119 |
| C1 |
O5 |
H9 |
110.767 |
|
C2 |
C1 |
O4 |
123.025 |
| C2 |
C1 |
O5 |
113.032 |
|
C2 |
N6 |
H7 |
118.806 |
| C2 |
N6 |
H8 |
120.418 |
|
O3 |
C2 |
N6 |
125.587 |
| O4 |
C1 |
O5 |
123.943 |
|
H7 |
N6 |
H8 |
120.776 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.052 |
|
|
|
| 2 |
C |
0.034 |
|
|
|
| 3 |
O |
-0.119 |
|
|
|
| 4 |
O |
-0.155 |
|
|
|
| 5 |
O |
-0.295 |
|
|
|
| 6 |
N |
-0.495 |
|
|
|
| 7 |
H |
0.341 |
|
|
|
| 8 |
H |
0.356 |
|
|
|
| 9 |
H |
0.385 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.584 |
0.777 |
0.000 |
2.699 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.418 |
8.140 |
-0.002 |
| y |
8.140 |
-42.315 |
-0.011 |
| z |
-0.002 |
-0.011 |
-34.238 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
14.858 |
8.140 |
-0.002 |
| y |
8.140 |
-13.486 |
-0.011 |
| z |
-0.002 |
-0.011 |
-1.372 |
|
| Polar |
|---|
| 3z2-r2 | -2.744 |
|---|
| x2-y2 | 18.896 |
|---|
| xy | 8.140 |
|---|
| xz | -0.002 |
|---|
| yz | -0.011 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.367 |
0.255 |
0.000 |
| y |
0.255 |
7.302 |
0.001 |
| z |
0.000 |
0.001 |
2.947 |
<r2> (average value of r
2) Å
2
| <r2> |
119.379 |
| (<r2>)1/2 |
10.926 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -70.855516 |
| Energy at 298.15K | -70.860813 |
| Nuclear repulsion energy | 123.577718 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3709 |
3614 |
55.49 |
|
|
|
| 2 |
A' |
3553 |
3462 |
64.85 |
|
|
|
| 3 |
A' |
3505 |
3416 |
85.83 |
|
|
|
| 4 |
A' |
1723 |
1679 |
142.10 |
|
|
|
| 5 |
A' |
1679 |
1636 |
400.14 |
|
|
|
| 6 |
A' |
1622 |
1580 |
24.33 |
|
|
|
| 7 |
A' |
1394 |
1359 |
27.80 |
|
|
|
| 8 |
A' |
1257 |
1225 |
522.77 |
|
|
|
| 9 |
A' |
1154 |
1125 |
20.80 |
|
|
|
| 10 |
A' |
1097 |
1069 |
0.23 |
|
|
|
| 11 |
A' |
763 |
743 |
16.02 |
|
|
|
| 12 |
A' |
596 |
581 |
11.84 |
|
|
|
| 13 |
A' |
530 |
517 |
1.95 |
|
|
|
| 14 |
A' |
387 |
377 |
6.49 |
|
|
|
| 15 |
A' |
252 |
245 |
44.21 |
|
|
|
| 16 |
A" |
773 |
753 |
18.51 |
|
|
|
| 17 |
A" |
702 |
684 |
205.95 |
|
|
|
| 18 |
A" |
668 |
651 |
297.82 |
|
|
|
| 19 |
A" |
631 |
615 |
12.21 |
|
|
|
| 20 |
A" |
418 |
408 |
3.46 |
|
|
|
| 21 |
A" |
108 |
105 |
12.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13259.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 12921.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.025 |
-0.798 |
0.000 |
| C2 |
0.000 |
0.758 |
0.000 |
| O3 |
-1.117 |
1.355 |
0.000 |
| O4 |
1.073 |
-1.465 |
0.000 |
| O5 |
-1.237 |
-1.338 |
0.000 |
| N6 |
1.233 |
1.336 |
0.000 |
| H7 |
1.317 |
2.343 |
0.000 |
| H8 |
2.059 |
0.749 |
0.000 |
| H9 |
-1.910 |
-0.612 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5565 | 2.4377 | 1.2422 | 1.3724 | 2.4523 | 3.3968 | 2.5561 | 1.9444 |
C2 | 1.5565 | | 1.2669 | 2.4687 | 2.4340 | 1.3620 | 2.0608 | 2.0592 | 2.3507 | O3 | 2.4377 | 1.2669 | | 3.5710 | 2.6962 | 2.3509 | 2.6270 | 3.2338 | 2.1207 | O4 | 1.2422 | 2.4687 | 3.5710 | | 2.3130 | 2.8056 | 3.8165 | 2.4245 | 3.1030 | O5 | 1.3724 | 2.4340 | 2.6962 | 2.3130 | | 3.6403 | 4.4805 | 3.9015 | 0.9910 | N6 | 2.4523 | 1.3620 | 2.3509 | 2.8056 | 3.6403 | | 1.0112 | 1.0126 | 3.6981 | H7 | 3.3968 | 2.0608 | 2.6270 | 3.8165 | 4.4805 | 1.0112 | | 1.7584 | 4.3757 | H8 | 2.5561 | 2.0592 | 3.2338 | 2.4245 | 3.9015 | 1.0126 | 1.7584 | | 4.1965 | H9 | 1.9444 | 2.3507 | 2.1207 | 3.1030 | 0.9910 | 3.6981 | 4.3757 | 4.1965 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.039 |
|
C1 |
C2 |
N6 |
114.172 |
| C1 |
O5 |
H9 |
109.662 |
|
C2 |
C1 |
O4 |
123.400 |
| C2 |
C1 |
O5 |
112.256 |
|
C2 |
N6 |
H7 |
119.816 |
| C2 |
N6 |
H8 |
119.539 |
|
O3 |
C2 |
N6 |
126.789 |
| O4 |
C1 |
O5 |
124.344 |
|
H7 |
N6 |
H8 |
120.645 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.015 |
|
|
|
| 2 |
C |
-0.031 |
|
|
|
| 3 |
O |
-0.199 |
|
|
|
| 4 |
O |
-0.126 |
|
|
|
| 5 |
O |
-0.317 |
|
|
|
| 6 |
N |
-0.448 |
|
|
|
| 7 |
H |
0.345 |
|
|
|
| 8 |
H |
0.363 |
|
|
|
| 9 |
H |
0.399 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.729 |
3.084 |
0.000 |
3.535 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.057 |
8.361 |
0.000 |
| y |
8.361 |
-39.849 |
0.000 |
| z |
0.000 |
0.000 |
-34.175 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.955 |
8.361 |
0.000 |
| y |
8.361 |
-7.233 |
0.000 |
| z |
0.000 |
0.000 |
1.278 |
|
| Polar |
|---|
| 3z2-r2 | 2.557 |
|---|
| x2-y2 | 8.792 |
|---|
| xy | 8.361 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.701 |
-0.171 |
0.000 |
| y |
-0.171 |
5.745 |
0.000 |
| z |
0.000 |
0.000 |
2.912 |
<r2> (average value of r
2) Å
2
| <r2> |
117.527 |
| (<r2>)1/2 |
10.841 |
