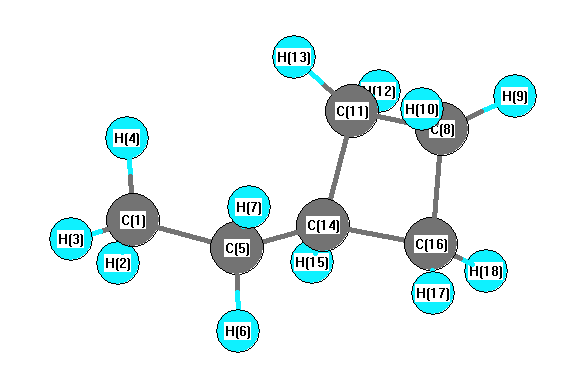
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3086 |
3008 |
146.84 |
|
|
|
| 2 |
A |
3065 |
2987 |
81.45 |
|
|
|
| 3 |
A |
3064 |
2985 |
15.94 |
|
|
|
| 4 |
A |
3062 |
2984 |
80.24 |
|
|
|
| 5 |
A |
3058 |
2980 |
13.99 |
|
|
|
| 6 |
A |
3016 |
2940 |
48.31 |
|
|
|
| 7 |
A |
3007 |
2930 |
37.17 |
|
|
|
| 8 |
A |
3000 |
2924 |
44.66 |
|
|
|
| 9 |
A |
2997 |
2921 |
95.16 |
|
|
|
| 10 |
A |
2980 |
2904 |
31.11 |
|
|
|
| 11 |
A |
2977 |
2901 |
23.20 |
|
|
|
| 12 |
A |
2961 |
2886 |
43.90 |
|
|
|
| 13 |
A |
1528 |
1489 |
7.07 |
|
|
|
| 14 |
A |
1523 |
1484 |
1.26 |
|
|
|
| 15 |
A |
1522 |
1483 |
8.40 |
|
|
|
| 16 |
A |
1511 |
1473 |
1.11 |
|
|
|
| 17 |
A |
1502 |
1464 |
2.03 |
|
|
|
| 18 |
A |
1496 |
1458 |
2.92 |
|
|
|
| 19 |
A |
1428 |
1392 |
6.21 |
|
|
|
| 20 |
A |
1383 |
1348 |
0.95 |
|
|
|
| 21 |
A |
1345 |
1311 |
5.66 |
|
|
|
| 22 |
A |
1305 |
1272 |
1.87 |
|
|
|
| 23 |
A |
1283 |
1250 |
0.28 |
|
|
|
| 24 |
A |
1275 |
1243 |
0.88 |
|
|
|
| 25 |
A |
1268 |
1236 |
0.04 |
|
|
|
| 26 |
A |
1254 |
1222 |
0.56 |
|
|
|
| 27 |
A |
1237 |
1205 |
0.25 |
|
|
|
| 28 |
A |
1203 |
1173 |
0.41 |
|
|
|
| 29 |
A |
1194 |
1164 |
0.05 |
|
|
|
| 30 |
A |
1138 |
1109 |
1.03 |
|
|
|
| 31 |
A |
1103 |
1075 |
0.32 |
|
|
|
| 32 |
A |
1018 |
992 |
0.67 |
|
|
|
| 33 |
A |
994 |
969 |
0.80 |
|
|
|
| 34 |
A |
986 |
960 |
3.08 |
|
|
|
| 35 |
A |
947 |
923 |
0.32 |
|
|
|
| 36 |
A |
932 |
908 |
1.50 |
|
|
|
| 37 |
A |
923 |
900 |
5.04 |
|
|
|
| 38 |
A |
875 |
853 |
1.15 |
|
|
|
| 39 |
A |
799 |
779 |
1.86 |
|
|
|
| 40 |
A |
790 |
770 |
1.21 |
|
|
|
| 41 |
A |
746 |
727 |
0.97 |
|
|
|
| 42 |
A |
625 |
609 |
1.66 |
|
|
|
| 43 |
A |
401 |
391 |
0.20 |
|
|
|
| 44 |
A |
394 |
384 |
0.01 |
|
|
|
| 45 |
A |
238 |
232 |
0.01 |
|
|
|
| 46 |
A |
198 |
193 |
0.03 |
|
|
|
| 47 |
A |
117 |
114 |
0.02 |
|
|
|
| 48 |
A |
100 |
98 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36427.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 35498.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.478 |
|
|
|
| 2 |
H |
0.138 |
|
|
|
| 3 |
H |
0.144 |
|
|
|
| 4 |
H |
0.146 |
|
|
|
| 5 |
C |
-0.289 |
|
|
|
| 6 |
H |
0.128 |
|
|
|
| 7 |
H |
0.124 |
|
|
|
| 8 |
C |
-0.215 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
C |
-0.358 |
|
|
|
| 12 |
H |
0.131 |
|
|
|
| 13 |
H |
0.126 |
|
|
|
| 14 |
C |
0.133 |
|
|
|
| 15 |
H |
0.114 |
|
|
|
| 16 |
C |
-0.350 |
|
|
|
| 17 |
H |
0.127 |
|
|
|
| 18 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.065 |
-0.035 |
0.064 |
0.097 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.345 |
-0.216 |
-0.172 |
| y |
-0.216 |
-41.068 |
-0.054 |
| z |
-0.172 |
-0.054 |
-39.588 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.017 |
-0.216 |
-0.172 |
| y |
-0.216 |
-0.602 |
-0.054 |
| z |
-0.172 |
-0.054 |
1.619 |
|
| Polar |
|---|
| 3z2-r2 | 3.238 |
|---|
| x2-y2 | -0.277 |
|---|
| xy | -0.216 |
|---|
| xz | -0.172 |
|---|
| yz | -0.054 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.462 |
0.020 |
0.012 |
| y |
0.020 |
9.948 |
0.006 |
| z |
0.012 |
0.006 |
9.244 |
<r2> (average value of r
2) Å
2
| <r2> |
171.770 |
| (<r2>)1/2 |
13.106 |