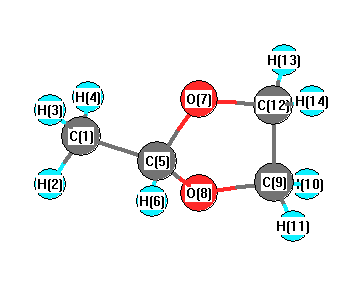
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3121 |
3041 |
51.08 |
|
|
|
| 2 |
A |
3119 |
3040 |
24.98 |
|
|
|
| 3 |
A |
3107 |
3028 |
25.21 |
|
|
|
| 4 |
A |
3087 |
3008 |
41.90 |
|
|
|
| 5 |
A |
3026 |
2949 |
62.38 |
|
|
|
| 6 |
A |
3014 |
2937 |
19.15 |
|
|
|
| 7 |
A |
2995 |
2919 |
73.34 |
|
|
|
| 8 |
A |
2966 |
2890 |
86.15 |
|
|
|
| 9 |
A |
1533 |
1494 |
0.41 |
|
|
|
| 10 |
A |
1518 |
1479 |
3.64 |
|
|
|
| 11 |
A |
1504 |
1466 |
5.30 |
|
|
|
| 12 |
A |
1502 |
1463 |
5.75 |
|
|
|
| 13 |
A |
1437 |
1401 |
38.49 |
|
|
|
| 14 |
A |
1399 |
1363 |
30.82 |
|
|
|
| 15 |
A |
1371 |
1336 |
3.33 |
|
|
|
| 16 |
A |
1361 |
1327 |
1.63 |
|
|
|
| 17 |
A |
1319 |
1285 |
4.93 |
|
|
|
| 18 |
A |
1226 |
1195 |
12.98 |
|
|
|
| 19 |
A |
1197 |
1166 |
1.37 |
|
|
|
| 20 |
A |
1171 |
1141 |
25.73 |
|
|
|
| 21 |
A |
1135 |
1106 |
9.91 |
|
|
|
| 22 |
A |
1109 |
1080 |
81.42 |
|
|
|
| 23 |
A |
1102 |
1074 |
52.10 |
|
|
|
| 24 |
A |
1009 |
983 |
31.25 |
|
|
|
| 25 |
A |
991 |
966 |
39.32 |
|
|
|
| 26 |
A |
933 |
909 |
41.37 |
|
|
|
| 27 |
A |
866 |
844 |
2.20 |
|
|
|
| 28 |
A |
825 |
804 |
39.54 |
|
|
|
| 29 |
A |
799 |
779 |
29.42 |
|
|
|
| 30 |
A |
666 |
649 |
0.88 |
|
|
|
| 31 |
A |
651 |
634 |
3.96 |
|
|
|
| 32 |
A |
478 |
466 |
8.01 |
|
|
|
| 33 |
A |
319 |
311 |
8.34 |
|
|
|
| 34 |
A |
206 |
200 |
1.51 |
|
|
|
| 35 |
A |
192 |
187 |
4.72 |
|
|
|
| 36 |
A |
50 |
49 |
9.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26152.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 25485.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.453 |
|
|
|
| 2 |
H |
0.156 |
|
|
|
| 3 |
H |
0.160 |
|
|
|
| 4 |
H |
0.157 |
|
|
|
| 5 |
C |
0.125 |
|
|
|
| 6 |
H |
0.119 |
|
|
|
| 7 |
O |
-0.285 |
|
|
|
| 8 |
O |
-0.276 |
|
|
|
| 9 |
C |
-0.164 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.142 |
|
|
|
| 12 |
C |
-0.145 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
| 14 |
H |
0.145 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.451 |
-0.279 |
0.497 |
1.559 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.935 |
-0.104 |
-0.890 |
| y |
-0.104 |
-43.082 |
-1.132 |
| z |
-0.890 |
-1.132 |
-36.040 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.626 |
-0.104 |
-0.890 |
| y |
-0.104 |
-10.095 |
-1.132 |
| z |
-0.890 |
-1.132 |
0.469 |
|
| Polar |
|---|
| 3z2-r2 | 0.938 |
|---|
| x2-y2 | 13.147 |
|---|
| xy | -0.104 |
|---|
| xz | -0.890 |
|---|
| yz | -1.132 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.398 |
0.046 |
-0.251 |
| y |
0.046 |
6.960 |
-0.088 |
| z |
-0.251 |
-0.088 |
6.891 |
<r2> (average value of r
2) Å
2
| <r2> |
129.798 |
| (<r2>)1/2 |
11.393 |