Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -36.442551 |
| Energy at 298.15K | -36.449811 |
| HF Energy | -36.442551 |
| Nuclear repulsion energy | 70.414747 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3044 |
2967 |
34.66 |
|
|
|
| 2 |
A1 |
3020 |
2943 |
12.61 |
|
|
|
| 3 |
A1 |
1547 |
1508 |
0.21 |
|
|
|
| 4 |
A1 |
1514 |
1475 |
4.11 |
|
|
|
| 5 |
A1 |
1374 |
1339 |
0.63 |
|
|
|
| 6 |
A1 |
1014 |
988 |
0.40 |
|
|
|
| 7 |
A1 |
872 |
850 |
25.21 |
|
|
|
| 8 |
A1 |
794 |
774 |
6.96 |
|
|
|
| 9 |
A2 |
3081 |
3002 |
0.00 |
|
|
|
| 10 |
A2 |
1239 |
1208 |
0.00 |
|
|
|
| 11 |
A2 |
1124 |
1096 |
0.00 |
|
|
|
| 12 |
A2 |
851 |
829 |
0.00 |
|
|
|
| 13 |
B1 |
3114 |
3035 |
106.92 |
|
|
|
| 14 |
B1 |
3075 |
2996 |
43.85 |
|
|
|
| 15 |
B1 |
1198 |
1168 |
1.81 |
|
|
|
| 16 |
B1 |
1120 |
1092 |
0.71 |
|
|
|
| 17 |
B1 |
778 |
758 |
0.58 |
|
|
|
| 18 |
B1 |
96 |
93 |
5.48 |
|
|
|
| 19 |
B2 |
3013 |
2936 |
170.31 |
|
|
|
| 20 |
B2 |
1521 |
1482 |
0.26 |
|
|
|
| 21 |
B2 |
1322 |
1288 |
9.09 |
|
|
|
| 22 |
B2 |
1285 |
1252 |
8.30 |
|
|
|
| 23 |
B2 |
980 |
955 |
42.48 |
|
|
|
| 24 |
B2 |
929 |
905 |
39.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18952.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 18469.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.109 |
| C2 |
0.000 |
0.000 |
-1.088 |
| C3 |
0.000 |
1.073 |
0.051 |
| C4 |
0.000 |
-1.073 |
0.051 |
| H5 |
0.893 |
0.000 |
-1.722 |
| H6 |
-0.893 |
0.000 |
-1.722 |
| H7 |
0.899 |
1.697 |
0.123 |
| H8 |
-0.899 |
1.697 |
0.123 |
| H9 |
-0.899 |
-1.697 |
0.123 |
| H10 |
0.899 |
-1.697 |
0.123 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1968 | 1.5066 | 1.5066 | 2.9680 | 2.9680 | 2.1588 | 2.1588 | 2.1588 | 2.1588 |
C2 | 2.1968 | | 1.5644 | 1.5644 | 1.0948 | 1.0948 | 2.2702 | 2.2702 | 2.2702 | 2.2702 | C3 | 1.5066 | 1.5644 | | 2.1451 | 2.2559 | 2.2559 | 1.0968 | 1.0968 | 2.9127 | 2.9127 | C4 | 1.5066 | 1.5644 | 2.1451 | | 2.2559 | 2.2559 | 2.9127 | 2.9127 | 1.0968 | 1.0968 | H5 | 2.9680 | 1.0948 | 2.2559 | 2.2559 | | 1.7855 | 2.5064 | 3.0808 | 3.0808 | 2.5064 | H6 | 2.9680 | 1.0948 | 2.2559 | 2.2559 | 1.7855 | | 3.0808 | 2.5064 | 2.5064 | 3.0808 | H7 | 2.1588 | 2.2702 | 1.0968 | 2.9127 | 2.5064 | 3.0808 | | 1.7976 | 3.8407 | 3.3941 | H8 | 2.1588 | 2.2702 | 1.0968 | 2.9127 | 3.0808 | 2.5064 | 1.7976 | | 3.3941 | 3.8407 | H9 | 2.1588 | 2.2702 | 2.9127 | 1.0968 | 3.0808 | 2.5064 | 3.8407 | 3.3941 | | 1.7976 | H10 | 2.1588 | 2.2702 | 2.9127 | 1.0968 | 2.5064 | 3.0808 | 3.3941 | 3.8407 | 1.7976 | |
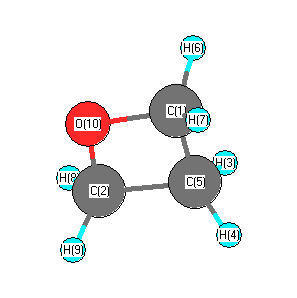 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.325 |
|
O1 |
C3 |
H7 |
111.055 |
| O1 |
C3 |
H8 |
111.055 |
|
O1 |
C4 |
C2 |
91.325 |
| O1 |
C4 |
H9 |
111.055 |
|
O1 |
C4 |
H10 |
111.055 |
| C2 |
C3 |
H7 |
115.983 |
|
C2 |
C3 |
H8 |
115.983 |
| C2 |
C4 |
H9 |
115.983 |
|
C2 |
C4 |
H10 |
115.983 |
| C3 |
O1 |
C4 |
90.780 |
|
C3 |
C2 |
C4 |
86.570 |
| C3 |
C2 |
H5 |
114.922 |
|
C3 |
C2 |
H6 |
114.922 |
| C4 |
C2 |
H5 |
114.922 |
|
C4 |
C2 |
H6 |
114.922 |
| H5 |
C2 |
H6 |
109.259 |
|
H7 |
C3 |
H8 |
110.064 |
| H9 |
C4 |
H10 |
110.064 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.276 |
|
|
|
| 2 |
C |
-0.251 |
|
|
|
| 3 |
C |
-0.154 |
|
|
|
| 4 |
C |
-0.154 |
|
|
|
| 5 |
H |
0.149 |
|
|
|
| 6 |
H |
0.149 |
|
|
|
| 7 |
H |
0.134 |
|
|
|
| 8 |
H |
0.134 |
|
|
|
| 9 |
H |
0.134 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.622 |
2.622 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.354 |
0.000 |
0.000 |
| y |
0.000 |
-22.609 |
0.000 |
| z |
0.000 |
0.000 |
-29.004 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.453 |
0.000 |
0.000 |
| y |
0.000 |
4.070 |
0.000 |
| z |
0.000 |
0.000 |
-5.523 |
|
| Polar |
|---|
| 3z2-r2 | -11.046 |
|---|
| x2-y2 | -1.745 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.929 |
0.000 |
0.000 |
| y |
0.000 |
6.199 |
0.000 |
| z |
0.000 |
0.000 |
4.963 |
<r2> (average value of r
2) Å
2
| <r2> |
59.486 |
| (<r2>)1/2 |
7.713 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -36.442551 |
| Energy at 298.15K | -36.449811 |
| HF Energy | -36.442551 |
| Nuclear repulsion energy | 70.414753 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3114 |
3034 |
107.65 |
|
|
|
| 2 |
A' |
3075 |
2996 |
43.16 |
|
|
|
| 3 |
A' |
3044 |
2966 |
34.52 |
|
|
|
| 4 |
A' |
3019 |
2942 |
12.79 |
|
|
|
| 5 |
A' |
1547 |
1508 |
0.21 |
|
|
|
| 6 |
A' |
1514 |
1475 |
4.10 |
|
|
|
| 7 |
A' |
1374 |
1339 |
0.63 |
|
|
|
| 8 |
A' |
1198 |
1168 |
1.81 |
|
|
|
| 9 |
A' |
1120 |
1092 |
0.71 |
|
|
|
| 10 |
A' |
1014 |
989 |
0.39 |
|
|
|
| 11 |
A' |
872 |
850 |
25.21 |
|
|
|
| 12 |
A' |
794 |
774 |
6.97 |
|
|
|
| 13 |
A' |
778 |
758 |
0.58 |
|
|
|
| 14 |
A' |
96 |
93 |
5.48 |
|
|
|
| 15 |
A" |
3081 |
3002 |
0.00 |
|
|
|
| 16 |
A" |
3013 |
2936 |
170.32 |
|
|
|
| 17 |
A" |
1521 |
1482 |
0.26 |
|
|
|
| 18 |
A" |
1322 |
1288 |
9.11 |
|
|
|
| 19 |
A" |
1285 |
1253 |
8.27 |
|
|
|
| 20 |
A" |
1239 |
1208 |
0.00 |
|
|
|
| 21 |
A" |
1124 |
1096 |
0.00 |
|
|
|
| 22 |
A" |
980 |
955 |
42.12 |
|
|
|
| 23 |
A" |
929 |
905 |
40.07 |
|
|
|
| 24 |
A" |
851 |
829 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18952.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 18469.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.001 |
-1.109 |
0.000 |
| C2 |
0.000 |
1.088 |
0.000 |
| C3 |
0.000 |
-0.051 |
1.073 |
| C4 |
0.000 |
-0.051 |
-1.073 |
| H5 |
0.893 |
1.722 |
0.000 |
| H6 |
-0.893 |
1.722 |
0.000 |
| H7 |
0.899 |
-0.123 |
1.697 |
| H8 |
-0.898 |
-0.122 |
1.698 |
| H9 |
0.899 |
-0.123 |
-1.697 |
| H10 |
-0.898 |
-0.122 |
-1.698 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1968 | 1.5067 | 1.5067 | 2.9686 | 2.9679 | 2.1588 | 2.1588 | 2.1588 | 2.1588 |
C2 | 2.1968 | | 1.5642 | 1.5642 | 1.0949 | 1.0949 | 2.2700 | 2.2702 | 2.2700 | 2.2702 | C3 | 1.5067 | 1.5642 | | 2.1450 | 2.2560 | 2.2559 | 1.0968 | 1.0968 | 2.9123 | 2.9130 | C4 | 1.5067 | 1.5642 | 2.1450 | | 2.2560 | 2.2559 | 2.9123 | 2.9130 | 1.0968 | 1.0968 | H5 | 2.9686 | 1.0949 | 2.2560 | 2.2560 | | 1.7852 | 2.5066 | 3.0808 | 2.5066 | 3.0808 | H6 | 2.9679 | 1.0949 | 2.2559 | 2.2559 | 1.7852 | | 3.0809 | 2.5065 | 3.0809 | 2.5065 | H7 | 2.1588 | 2.2700 | 1.0968 | 2.9123 | 2.5066 | 3.0809 | | 1.7975 | 3.3932 | 3.8407 | H8 | 2.1588 | 2.2702 | 1.0968 | 2.9130 | 3.0808 | 2.5065 | 1.7975 | | 3.8407 | 3.3950 | H9 | 2.1588 | 2.2700 | 2.9123 | 1.0968 | 2.5066 | 3.0809 | 3.3932 | 3.8407 | | 1.7975 | H10 | 2.1588 | 2.2702 | 2.9130 | 1.0968 | 3.0808 | 2.5065 | 3.8407 | 3.3950 | 1.7975 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.328 |
|
O1 |
C3 |
H7 |
111.053 |
| O1 |
C3 |
H8 |
111.052 |
|
O1 |
C4 |
C2 |
91.328 |
| O1 |
C4 |
H9 |
111.053 |
|
O1 |
C4 |
H10 |
111.052 |
| C2 |
C3 |
H7 |
115.984 |
|
C2 |
C3 |
H8 |
115.995 |
| C2 |
C4 |
H9 |
115.984 |
|
C2 |
C4 |
H10 |
115.995 |
| C3 |
O1 |
C4 |
90.770 |
|
C3 |
C2 |
C4 |
86.575 |
| C3 |
C2 |
H5 |
114.937 |
|
C3 |
C2 |
H6 |
114.925 |
| C4 |
C2 |
H5 |
114.937 |
|
C4 |
C2 |
H6 |
114.925 |
| H5 |
C2 |
H6 |
109.228 |
|
H7 |
C3 |
H8 |
110.056 |
| H9 |
C4 |
H10 |
110.056 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.276 |
|
|
|
| 2 |
C |
-0.251 |
|
|
|
| 3 |
C |
-0.154 |
|
|
|
| 4 |
C |
-0.154 |
|
|
|
| 5 |
H |
0.149 |
|
|
|
| 6 |
H |
0.149 |
|
|
|
| 7 |
H |
0.134 |
|
|
|
| 8 |
H |
0.134 |
|
|
|
| 9 |
H |
0.134 |
|
|
|
| 10 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.002 |
2.622 |
0.000 |
2.622 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.355 |
-0.003 |
0.000 |
| y |
-0.003 |
-29.003 |
0.000 |
| z |
0.000 |
0.000 |
-22.609 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.451 |
-0.003 |
0.000 |
| y |
-0.003 |
-5.521 |
0.000 |
| z |
0.000 |
0.000 |
4.071 |
|
| Polar |
|---|
| 3z2-r2 | 8.142 |
|---|
| x2-y2 | 4.648 |
|---|
| xy | -0.003 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.929 |
0.000 |
0.000 |
| y |
0.000 |
4.963 |
0.000 |
| z |
0.000 |
0.000 |
6.199 |
<r2> (average value of r
2) Å
2
| <r2> |
59.486 |
| (<r2>)1/2 |
7.713 |
