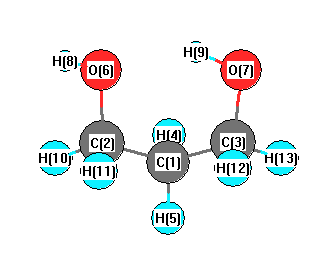
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3686 |
3592 |
6.54 |
|
|
|
| 2 |
A |
3563 |
3472 |
91.12 |
|
|
|
| 3 |
A |
3073 |
2995 |
65.66 |
|
|
|
| 4 |
A |
3062 |
2984 |
62.87 |
|
|
|
| 5 |
A |
3028 |
2951 |
63.95 |
|
|
|
| 6 |
A |
2995 |
2919 |
34.46 |
|
|
|
| 7 |
A |
2967 |
2891 |
61.92 |
|
|
|
| 8 |
A |
2944 |
2869 |
83.54 |
|
|
|
| 9 |
A |
1532 |
1493 |
2.55 |
|
|
|
| 10 |
A |
1521 |
1483 |
1.91 |
|
|
|
| 11 |
A |
1485 |
1447 |
10.78 |
|
|
|
| 12 |
A |
1446 |
1409 |
79.57 |
|
|
|
| 13 |
A |
1430 |
1393 |
4.88 |
|
|
|
| 14 |
A |
1391 |
1355 |
0.40 |
|
|
|
| 15 |
A |
1377 |
1342 |
7.54 |
|
|
|
| 16 |
A |
1291 |
1258 |
18.24 |
|
|
|
| 17 |
A |
1256 |
1224 |
1.46 |
|
|
|
| 18 |
A |
1203 |
1172 |
5.67 |
|
|
|
| 19 |
A |
1175 |
1145 |
72.05 |
|
|
|
| 20 |
A |
1129 |
1100 |
12.91 |
|
|
|
| 21 |
A |
1073 |
1046 |
13.42 |
|
|
|
| 22 |
A |
1045 |
1019 |
119.56 |
|
|
|
| 23 |
A |
928 |
904 |
3.45 |
|
|
|
| 24 |
A |
908 |
885 |
5.02 |
|
|
|
| 25 |
A |
888 |
866 |
27.17 |
|
|
|
| 26 |
A |
810 |
789 |
13.76 |
|
|
|
| 27 |
A |
597 |
581 |
184.03 |
|
|
|
| 28 |
A |
497 |
484 |
3.79 |
|
|
|
| 29 |
A |
375 |
365 |
6.31 |
|
|
|
| 30 |
A |
323 |
314 |
29.57 |
|
|
|
| 31 |
A |
275 |
268 |
125.93 |
|
|
|
| 32 |
A |
205 |
200 |
8.25 |
|
|
|
| 33 |
A |
112 |
109 |
1.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24793.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 24161.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.272 |
|
|
|
| 2 |
C |
-0.129 |
|
|
|
| 3 |
C |
-0.120 |
|
|
|
| 4 |
H |
0.159 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
O |
-0.531 |
|
|
|
| 7 |
O |
-0.468 |
|
|
|
| 8 |
H |
0.354 |
|
|
|
| 9 |
H |
0.352 |
|
|
|
| 10 |
H |
0.141 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
| 12 |
H |
0.105 |
|
|
|
| 13 |
H |
0.146 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.884 |
1.964 |
1.226 |
4.522 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.015 |
-5.518 |
0.630 |
| y |
-5.518 |
-30.847 |
-1.197 |
| z |
0.630 |
-1.197 |
-32.345 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.581 |
-5.518 |
0.630 |
| y |
-5.518 |
0.833 |
-1.197 |
| z |
0.630 |
-1.197 |
-1.414 |
|
| Polar |
|---|
| 3z2-r2 | -2.828 |
|---|
| x2-y2 | -0.168 |
|---|
| xy | -5.518 |
|---|
| xz | 0.630 |
|---|
| yz | -1.197 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.991 |
-0.204 |
0.109 |
| y |
-0.204 |
6.811 |
0.103 |
| z |
0.109 |
0.103 |
5.702 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |