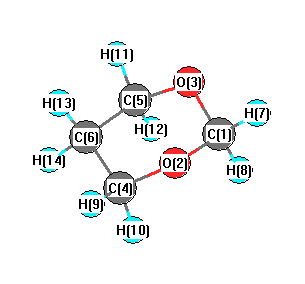
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3152 |
3071 |
28.01 |
|
|
|
| 2 |
A' |
3106 |
3027 |
38.03 |
|
|
|
| 3 |
A' |
3073 |
2995 |
39.36 |
|
|
|
| 4 |
A' |
3006 |
2929 |
36.87 |
|
|
|
| 5 |
A' |
2956 |
2881 |
147.11 |
|
|
|
| 6 |
A' |
2921 |
2847 |
77.28 |
|
|
|
| 7 |
A' |
1517 |
1478 |
6.88 |
|
|
|
| 8 |
A' |
1490 |
1452 |
0.78 |
|
|
|
| 9 |
A' |
1488 |
1450 |
7.58 |
|
|
|
| 10 |
A' |
1398 |
1362 |
16.46 |
|
|
|
| 11 |
A' |
1298 |
1265 |
4.51 |
|
|
|
| 12 |
A' |
1183 |
1153 |
12.23 |
|
|
|
| 13 |
A' |
1136 |
1107 |
106.87 |
|
|
|
| 14 |
A' |
1081 |
1054 |
47.40 |
|
|
|
| 15 |
A' |
970 |
945 |
46.31 |
|
|
|
| 16 |
A' |
890 |
867 |
4.26 |
|
|
|
| 17 |
A' |
805 |
784 |
16.54 |
|
|
|
| 18 |
A' |
619 |
603 |
6.23 |
|
|
|
| 19 |
A' |
468 |
456 |
0.95 |
|
|
|
| 20 |
A' |
392 |
382 |
13.24 |
|
|
|
| 21 |
A' |
253 |
246 |
3.84 |
|
|
|
| 22 |
A" |
3102 |
3023 |
48.53 |
|
|
|
| 23 |
A" |
2948 |
2872 |
25.40 |
|
|
|
| 24 |
A" |
1504 |
1465 |
4.28 |
|
|
|
| 25 |
A" |
1421 |
1384 |
8.01 |
|
|
|
| 26 |
A" |
1383 |
1347 |
1.02 |
|
|
|
| 27 |
A" |
1371 |
1336 |
0.73 |
|
|
|
| 28 |
A" |
1300 |
1267 |
0.13 |
|
|
|
| 29 |
A" |
1241 |
1209 |
18.56 |
|
|
|
| 30 |
A" |
1208 |
1178 |
3.07 |
|
|
|
| 31 |
A" |
1054 |
1027 |
2.80 |
|
|
|
| 32 |
A" |
979 |
954 |
108.53 |
|
|
|
| 33 |
A" |
902 |
879 |
23.97 |
|
|
|
| 34 |
A" |
840 |
819 |
28.31 |
|
|
|
| 35 |
A" |
440 |
429 |
10.27 |
|
|
|
| 36 |
A" |
240 |
234 |
2.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26566.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 25888.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.128 |
|
|
|
| 2 |
O |
-0.241 |
|
|
|
| 3 |
O |
-0.241 |
|
|
|
| 4 |
C |
-0.141 |
|
|
|
| 5 |
C |
-0.141 |
|
|
|
| 6 |
C |
-0.277 |
|
|
|
| 7 |
H |
0.180 |
|
|
|
| 8 |
H |
0.118 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
| 11 |
H |
0.168 |
|
|
|
| 12 |
H |
0.121 |
|
|
|
| 13 |
H |
0.159 |
|
|
|
| 14 |
H |
0.135 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.017 |
2.571 |
0.000 |
2.765 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.164 |
2.274 |
0.000 |
| y |
2.274 |
-35.223 |
0.000 |
| z |
0.000 |
0.000 |
-40.581 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.738 |
2.274 |
0.000 |
| y |
2.274 |
3.149 |
0.000 |
| z |
0.000 |
0.000 |
-4.888 |
|
| Polar |
|---|
| 3z2-r2 | -9.775 |
|---|
| x2-y2 | -0.941 |
|---|
| xy | 2.274 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.093 |
0.480 |
0.000 |
| y |
0.480 |
8.287 |
0.000 |
| z |
0.000 |
0.000 |
7.554 |
<r2> (average value of r
2) Å
2
| <r2> |
120.790 |
| (<r2>)1/2 |
10.990 |