Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -23.507620 |
| Energy at 298.15K | -23.511814 |
| HF Energy | -23.507620 |
| Nuclear repulsion energy | 35.415372 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3339 |
3254 |
15.78 |
|
|
|
| 2 |
A' |
3240 |
3158 |
13.36 |
|
|
|
| 3 |
A' |
3114 |
3035 |
8.26 |
|
|
|
| 4 |
A' |
3036 |
2958 |
57.82 |
|
|
|
| 5 |
A' |
1506 |
1468 |
8.74 |
|
|
|
| 6 |
A' |
1447 |
1410 |
8.59 |
|
|
|
| 7 |
A' |
1303 |
1270 |
34.86 |
|
|
|
| 8 |
A' |
1212 |
1181 |
1.65 |
|
|
|
| 9 |
A' |
1103 |
1075 |
29.51 |
|
|
|
| 10 |
A' |
984 |
959 |
12.07 |
|
|
|
| 11 |
A' |
482 |
469 |
12.94 |
|
|
|
| 12 |
A" |
1050 |
1023 |
2.37 |
|
|
|
| 13 |
A" |
837 |
816 |
27.63 |
|
|
|
| 14 |
A" |
683 |
665 |
130.80 |
|
|
|
| 15 |
A" |
544 |
531 |
13.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11939.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 11635.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.430 |
0.000 |
| C2 |
1.181 |
-0.347 |
0.000 |
| N3 |
-1.211 |
-0.175 |
0.000 |
| H4 |
0.105 |
1.523 |
0.000 |
| H5 |
2.164 |
0.118 |
0.000 |
| H6 |
1.103 |
-1.432 |
0.000 |
| H7 |
-1.978 |
0.522 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4135 | 1.3538 | 1.0985 | 2.1864 | 2.1643 | 1.9805 |
C2 | 1.4135 | | 2.3986 | 2.1577 | 1.0879 | 1.0880 | 3.2766 | N3 | 1.3538 | 2.3986 | | 2.1485 | 3.3883 | 2.6344 | 1.0360 | H4 | 1.0985 | 2.1577 | 2.1485 | | 2.4926 | 3.1195 | 2.3116 | H5 | 2.1864 | 1.0879 | 3.3883 | 2.4926 | | 1.8789 | 4.1621 | H6 | 2.1643 | 1.0880 | 2.6344 | 3.1195 | 1.8789 | | 3.6491 | H7 | 1.9805 | 3.2766 | 1.0360 | 2.3116 | 4.1621 | 3.6491 | |
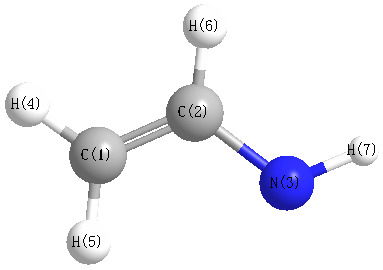 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.325 |
|
C1 |
C2 |
H6 |
119.246 |
| C1 |
N3 |
H7 |
111.244 |
|
C2 |
C1 |
N3 |
120.152 |
| C2 |
C1 |
H4 |
117.847 |
|
N3 |
C1 |
H4 |
122.001 |
| H5 |
C2 |
H6 |
119.429 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.218 |
|
|
|
| 2 |
C |
-0.205 |
|
|
|
| 3 |
N |
-0.180 |
|
|
|
| 4 |
H |
0.110 |
|
|
|
| 5 |
H |
0.137 |
|
|
|
| 6 |
H |
0.140 |
|
|
|
| 7 |
H |
0.216 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.008 |
1.896 |
0.000 |
2.147 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.625 |
-3.233 |
0.000 |
| y |
-3.233 |
-18.275 |
0.000 |
| z |
0.000 |
0.000 |
-20.569 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.797 |
-3.233 |
0.000 |
| y |
-3.233 |
0.322 |
0.000 |
| z |
0.000 |
0.000 |
-3.119 |
|
| Polar |
|---|
| 3z2-r2 | -6.239 |
|---|
| x2-y2 | 1.650 |
|---|
| xy | -3.233 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.951 |
-0.396 |
0.000 |
| y |
-0.396 |
4.456 |
0.000 |
| z |
0.000 |
0.000 |
2.408 |
<r2> (average value of r
2) Å
2
| <r2> |
40.322 |
| (<r2>)1/2 |
6.350 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -23.506330 |
| Energy at 298.15K | -23.510482 |
| HF Energy | -23.506330 |
| Nuclear repulsion energy | 35.440565 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3281 |
3198 |
38.79 |
|
|
|
| 2 |
A' |
3217 |
3135 |
21.08 |
|
|
|
| 3 |
A' |
3122 |
3042 |
17.08 |
|
|
|
| 4 |
A' |
3093 |
3014 |
11.70 |
|
|
|
| 5 |
A' |
1492 |
1454 |
6.31 |
|
|
|
| 6 |
A' |
1433 |
1396 |
4.65 |
|
|
|
| 7 |
A' |
1337 |
1303 |
22.73 |
|
|
|
| 8 |
A' |
1181 |
1151 |
7.38 |
|
|
|
| 9 |
A' |
1127 |
1099 |
51.76 |
|
|
|
| 10 |
A' |
978 |
953 |
0.76 |
|
|
|
| 11 |
A' |
475 |
463 |
9.44 |
|
|
|
| 12 |
A" |
1060 |
1033 |
73.28 |
|
|
|
| 13 |
A" |
830 |
809 |
75.86 |
|
|
|
| 14 |
A" |
676 |
659 |
17.54 |
|
|
|
| 15 |
A" |
500 |
487 |
0.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11900.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 11596.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.456 |
0.000 |
| C2 |
1.161 |
-0.355 |
0.000 |
| N3 |
-1.286 |
0.031 |
0.000 |
| H4 |
0.125 |
1.541 |
0.000 |
| H5 |
2.155 |
0.089 |
0.000 |
| H6 |
1.087 |
-1.443 |
0.000 |
| H7 |
-1.329 |
-1.009 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4161 | 1.3545 | 1.0918 | 2.1856 | 2.1885 | 1.9780 |
C2 | 1.4161 | | 2.4773 | 2.1601 | 1.0883 | 1.0911 | 2.5746 | N3 | 1.3545 | 2.4773 | | 2.0663 | 3.4412 | 2.7939 | 1.0407 | H4 | 1.0918 | 2.1601 | 2.0663 | | 2.4954 | 3.1353 | 2.9349 | H5 | 2.1856 | 1.0883 | 3.4412 | 2.4954 | | 1.8676 | 3.6527 | H6 | 2.1885 | 1.0911 | 2.7939 | 3.1353 | 1.8676 | | 2.4551 | H7 | 1.9780 | 2.5746 | 1.0407 | 2.9349 | 3.6527 | 2.4551 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
120.997 |
|
C1 |
C2 |
H6 |
121.050 |
| C1 |
N3 |
H7 |
110.662 |
|
C2 |
C1 |
N3 |
126.783 |
| C2 |
C1 |
H4 |
118.366 |
|
N3 |
C1 |
H4 |
114.850 |
| H5 |
C2 |
H6 |
117.953 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.176 |
|
|
|
| 2 |
C |
-0.270 |
|
|
|
| 3 |
N |
-0.173 |
|
|
|
| 4 |
H |
0.141 |
|
|
|
| 5 |
H |
0.140 |
|
|
|
| 6 |
H |
0.140 |
|
|
|
| 7 |
H |
0.198 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.356 |
-1.217 |
0.000 |
2.652 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.649 |
2.457 |
0.000 |
| y |
2.457 |
-16.293 |
0.000 |
| z |
0.000 |
0.000 |
-20.566 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.220 |
2.457 |
0.000 |
| y |
2.457 |
4.814 |
0.000 |
| z |
0.000 |
0.000 |
-1.595 |
|
| Polar |
|---|
| 3z2-r2 | -3.190 |
|---|
| x2-y2 | -5.356 |
|---|
| xy | 2.457 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.635 |
-0.120 |
0.000 |
| y |
-0.120 |
4.567 |
0.000 |
| z |
0.000 |
0.000 |
2.413 |
<r2> (average value of r
2) Å
2
| <r2> |
40.272 |
| (<r2>)1/2 |
6.346 |
