Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -27.715947 |
| Energy at 298.15K | |
| HF Energy | -27.715947 |
| Nuclear repulsion energy | 25.781341 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3429 |
3341 |
91.03 |
51.90 |
0.22 |
0.36 |
| 2 |
Σ |
1942 |
1893 |
97.72 |
1.82 |
0.42 |
0.59 |
| 3 |
Σ |
1205 |
1175 |
28.49 |
42.09 |
0.23 |
0.37 |
| 4 |
Π |
508 |
495 |
7.38 |
0.66 |
0.75 |
0.86 |
| 4 |
Π |
496 |
483 |
42.11 |
2.58 |
0.75 |
0.86 |
| 5 |
Π |
459 |
447 |
0.17 |
1.21 |
0.75 |
0.86 |
| 5 |
Π |
55 |
54 |
174.86 |
14.81 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4046.8 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 3943.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.004 |
| C2 |
0.000 |
0.000 |
-1.268 |
| O3 |
0.000 |
0.000 |
1.239 |
| H4 |
0.000 |
0.000 |
-2.335 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2722 | 1.2350 | 2.3393 |
C2 | 1.2722 | | 2.5072 | 1.0671 | O3 | 1.2350 | 2.5072 | | 3.5743 | H4 | 2.3393 | 1.0671 | 3.5743 | |
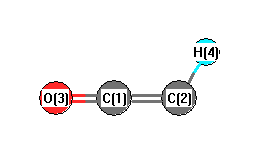 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.048 |
|
|
|
| 2 |
C |
-0.227 |
|
|
|
| 3 |
O |
-0.014 |
|
|
|
| 4 |
H |
0.289 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.510 |
2.510 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.031 |
0.000 |
0.000 |
| y |
0.000 |
-16.283 |
0.000 |
| z |
0.000 |
0.000 |
-15.239 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.270 |
0.000 |
0.000 |
| y |
0.000 |
0.352 |
0.000 |
| z |
0.000 |
0.000 |
1.918 |
|
| Polar |
|---|
| 3z2-r2 | 3.836 |
|---|
| x2-y2 | -1.748 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.628 |
0.000 |
0.000 |
| y |
0.000 |
1.527 |
0.000 |
| z |
0.000 |
0.000 |
6.128 |
<r2> (average value of r
2) Å
2
| <r2> |
31.414 |
| (<r2>)1/2 |
5.605 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -27.715948 |
| Energy at 298.15K | |
| HF Energy | -27.715948 |
| Nuclear repulsion energy | 25.777518 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3411 |
3324 |
78.65 |
51.99 |
0.35 |
0.52 |
| 2 |
A' |
1935 |
1886 |
110.63 |
5.90 |
0.24 |
0.38 |
| 3 |
A' |
1205 |
1174 |
26.54 |
39.33 |
0.35 |
0.52 |
| 4 |
A' |
507 |
494 |
10.85 |
34.01 |
0.35 |
0.51 |
| 5 |
A' |
94 |
91 |
182.97 |
11.73 |
0.60 |
0.75 |
| 6 |
A" |
461 |
449 |
0.46 |
15.29 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3805.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 3708.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.025 |
0.000 |
| C2 |
1.230 |
-0.320 |
0.000 |
| O3 |
-1.210 |
0.260 |
0.000 |
| H4 |
2.299 |
-0.312 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2775 | 1.2328 | 2.3236 |
C2 | 1.2775 | | 2.5082 | 1.0689 | O3 | 1.2328 | 2.5082 | | 3.5555 | H4 | 2.3236 | 1.0689 | 3.5555 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
163.953 |
|
C2 |
C1 |
O3 |
175.383 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.036 |
|
|
|
| 2 |
C |
-0.260 |
|
|
|
| 3 |
O |
-0.016 |
|
|
|
| 4 |
H |
0.241 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.465 |
-0.008 |
0.000 |
2.465 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.179 |
0.420 |
0.000 |
| y |
0.420 |
-18.323 |
0.000 |
| z |
0.000 |
0.000 |
-16.296 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.131 |
0.420 |
0.000 |
| y |
0.420 |
-2.586 |
0.000 |
| z |
0.000 |
0.000 |
0.455 |
|
| Polar |
|---|
| 3z2-r2 | 0.910 |
|---|
| x2-y2 | 3.144 |
|---|
| xy | 0.420 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.872 |
-1.136 |
0.000 |
| y |
-1.136 |
1.991 |
0.000 |
| z |
0.000 |
0.000 |
2.300 |
<r2> (average value of r
2) Å
2
| <r2> |
31.407 |
| (<r2>)1/2 |
5.604 |
