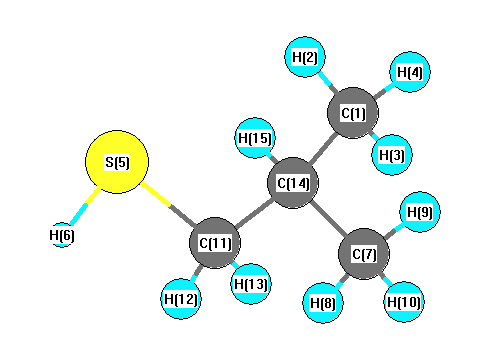
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3096 |
3017 |
46.60 |
|
|
|
| 2 |
A |
3084 |
3005 |
41.48 |
|
|
|
| 3 |
A |
3070 |
2992 |
85.20 |
|
|
|
| 4 |
A |
3064 |
2986 |
51.14 |
|
|
|
| 5 |
A |
3056 |
2978 |
9.79 |
|
|
|
| 6 |
A |
3019 |
2942 |
37.50 |
|
|
|
| 7 |
A |
2990 |
2914 |
8.64 |
|
|
|
| 8 |
A |
2981 |
2905 |
33.21 |
|
|
|
| 9 |
A |
2974 |
2898 |
38.52 |
|
|
|
| 10 |
A |
2477 |
2414 |
40.04 |
|
|
|
| 11 |
A |
1529 |
1490 |
14.61 |
|
|
|
| 12 |
A |
1525 |
1486 |
7.88 |
|
|
|
| 13 |
A |
1515 |
1476 |
5.33 |
|
|
|
| 14 |
A |
1511 |
1472 |
0.27 |
|
|
|
| 15 |
A |
1490 |
1452 |
3.98 |
|
|
|
| 16 |
A |
1438 |
1401 |
11.01 |
|
|
|
| 17 |
A |
1420 |
1384 |
10.58 |
|
|
|
| 18 |
A |
1378 |
1343 |
3.10 |
|
|
|
| 19 |
A |
1357 |
1323 |
1.72 |
|
|
|
| 20 |
A |
1284 |
1252 |
35.82 |
|
|
|
| 21 |
A |
1239 |
1208 |
3.09 |
|
|
|
| 22 |
A |
1200 |
1170 |
7.76 |
|
|
|
| 23 |
A |
1143 |
1114 |
4.64 |
|
|
|
| 24 |
A |
1082 |
1054 |
3.44 |
|
|
|
| 25 |
A |
974 |
949 |
0.13 |
|
|
|
| 26 |
A |
967 |
942 |
3.27 |
|
|
|
| 27 |
A |
941 |
917 |
2.77 |
|
|
|
| 28 |
A |
880 |
857 |
3.95 |
|
|
|
| 29 |
A |
868 |
846 |
1.55 |
|
|
|
| 30 |
A |
770 |
750 |
5.57 |
|
|
|
| 31 |
A |
683 |
666 |
5.95 |
|
|
|
| 32 |
A |
419 |
408 |
0.30 |
|
|
|
| 33 |
A |
383 |
374 |
1.74 |
|
|
|
| 34 |
A |
327 |
318 |
0.41 |
|
|
|
| 35 |
A |
235 |
229 |
0.36 |
|
|
|
| 36 |
A |
208 |
202 |
0.28 |
|
|
|
| 37 |
A |
194 |
190 |
2.30 |
|
|
|
| 38 |
A |
137 |
134 |
13.56 |
|
|
|
| 39 |
A |
71 |
69 |
15.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28489.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 27762.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.479 |
|
|
|
| 2 |
H |
0.163 |
|
|
|
| 3 |
H |
0.136 |
|
|
|
| 4 |
H |
0.144 |
|
|
|
| 5 |
S |
0.049 |
|
|
|
| 6 |
H |
0.038 |
|
|
|
| 7 |
C |
-0.495 |
|
|
|
| 8 |
H |
0.146 |
|
|
|
| 9 |
H |
0.153 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
C |
-0.577 |
|
|
|
| 12 |
H |
0.181 |
|
|
|
| 13 |
H |
0.173 |
|
|
|
| 14 |
C |
0.088 |
|
|
|
| 15 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.131 |
-1.387 |
0.753 |
1.942 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.573 |
3.509 |
-2.316 |
| y |
3.509 |
-40.390 |
-0.828 |
| z |
-2.316 |
-0.828 |
-41.657 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.550 |
3.509 |
-2.316 |
| y |
3.509 |
1.225 |
-0.828 |
| z |
-2.316 |
-0.828 |
-0.675 |
|
| Polar |
|---|
| 3z2-r2 | -1.350 |
|---|
| x2-y2 | -1.183 |
|---|
| xy | 3.509 |
|---|
| xz | -2.316 |
|---|
| yz | -0.828 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.116 |
-0.220 |
0.146 |
| y |
-0.220 |
9.262 |
-0.380 |
| z |
0.146 |
-0.380 |
8.101 |
<r2> (average value of r
2) Å
2
| <r2> |
145.990 |
| (<r2>)1/2 |
12.083 |