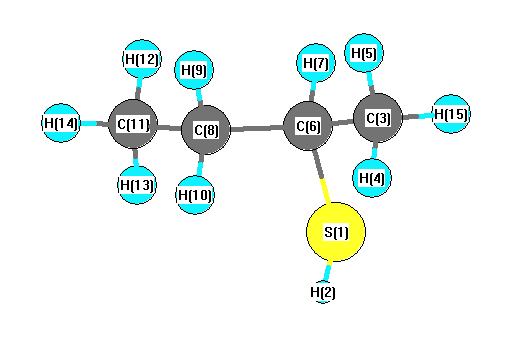
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3095 |
3017 |
35.90 |
|
|
|
| 2 |
A |
3092 |
3013 |
39.07 |
|
|
|
| 3 |
A |
3067 |
2988 |
85.41 |
|
|
|
| 4 |
A |
3060 |
2982 |
39.42 |
|
|
|
| 5 |
A |
3031 |
2954 |
9.16 |
|
|
|
| 6 |
A |
3009 |
2932 |
12.67 |
|
|
|
| 7 |
A |
2986 |
2910 |
30.09 |
|
|
|
| 8 |
A |
2979 |
2903 |
64.68 |
|
|
|
| 9 |
A |
2964 |
2888 |
31.39 |
|
|
|
| 10 |
A |
2467 |
2404 |
32.61 |
|
|
|
| 11 |
A |
1531 |
1492 |
7.96 |
|
|
|
| 12 |
A |
1518 |
1479 |
9.06 |
|
|
|
| 13 |
A |
1518 |
1479 |
3.22 |
|
|
|
| 14 |
A |
1512 |
1473 |
12.86 |
|
|
|
| 15 |
A |
1504 |
1466 |
1.54 |
|
|
|
| 16 |
A |
1431 |
1395 |
16.48 |
|
|
|
| 17 |
A |
1429 |
1393 |
1.80 |
|
|
|
| 18 |
A |
1379 |
1344 |
2.05 |
|
|
|
| 19 |
A |
1324 |
1290 |
12.42 |
|
|
|
| 20 |
A |
1312 |
1279 |
4.47 |
|
|
|
| 21 |
A |
1267 |
1235 |
5.86 |
|
|
|
| 22 |
A |
1184 |
1154 |
2.06 |
|
|
|
| 23 |
A |
1124 |
1095 |
2.89 |
|
|
|
| 24 |
A |
1118 |
1089 |
11.22 |
|
|
|
| 25 |
A |
1033 |
1006 |
0.40 |
|
|
|
| 26 |
A |
1015 |
989 |
9.34 |
|
|
|
| 27 |
A |
978 |
953 |
5.86 |
|
|
|
| 28 |
A |
852 |
830 |
1.83 |
|
|
|
| 29 |
A |
830 |
808 |
9.55 |
|
|
|
| 30 |
A |
789 |
768 |
9.58 |
|
|
|
| 31 |
A |
560 |
546 |
10.60 |
|
|
|
| 32 |
A |
447 |
435 |
0.50 |
|
|
|
| 33 |
A |
364 |
354 |
0.84 |
|
|
|
| 34 |
A |
317 |
309 |
1.11 |
|
|
|
| 35 |
A |
242 |
235 |
0.24 |
|
|
|
| 36 |
A |
225 |
220 |
0.31 |
|
|
|
| 37 |
A |
204 |
199 |
1.53 |
|
|
|
| 38 |
A |
174 |
169 |
20.82 |
|
|
|
| 39 |
A |
108 |
105 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28516.3 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 27789.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.021 |
|
|
|
| 2 |
H |
0.036 |
|
|
|
| 3 |
C |
-0.439 |
|
|
|
| 4 |
H |
0.143 |
|
|
|
| 5 |
H |
0.145 |
|
|
|
| 6 |
C |
-0.331 |
|
|
|
| 7 |
H |
0.173 |
|
|
|
| 8 |
C |
-0.167 |
|
|
|
| 9 |
H |
0.131 |
|
|
|
| 10 |
H |
0.129 |
|
|
|
| 11 |
C |
-0.459 |
|
|
|
| 12 |
H |
0.137 |
|
|
|
| 13 |
H |
0.162 |
|
|
|
| 14 |
H |
0.146 |
|
|
|
| 15 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.154 |
-1.992 |
0.639 |
2.098 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.708 |
0.967 |
-1.220 |
| y |
0.967 |
-44.446 |
2.263 |
| z |
-1.220 |
2.263 |
-38.719 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.125 |
0.967 |
-1.220 |
| y |
0.967 |
-4.233 |
2.263 |
| z |
-1.220 |
2.263 |
4.358 |
|
| Polar |
|---|
| 3z2-r2 | 8.716 |
|---|
| x2-y2 | 2.738 |
|---|
| xy | 0.967 |
|---|
| xz | -1.220 |
|---|
| yz | 2.263 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.885 |
-0.421 |
-0.214 |
| y |
-0.421 |
11.042 |
0.215 |
| z |
-0.214 |
0.215 |
8.773 |
<r2> (average value of r
2) Å
2
| <r2> |
143.916 |
| (<r2>)1/2 |
11.996 |