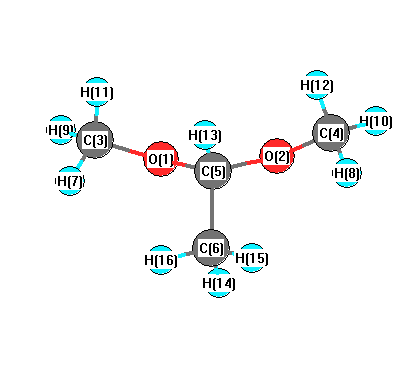
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3122 |
3042 |
28.14 |
|
|
|
| 2 |
A |
3110 |
3031 |
3.46 |
|
|
|
| 3 |
A |
3053 |
2976 |
136.98 |
|
|
|
| 4 |
A |
3020 |
2943 |
18.29 |
|
|
|
| 5 |
A |
2961 |
2885 |
136.98 |
|
|
|
| 6 |
A |
2914 |
2839 |
65.55 |
|
|
|
| 7 |
A |
1525 |
1486 |
12.87 |
|
|
|
| 8 |
A |
1504 |
1466 |
3.33 |
|
|
|
| 9 |
A |
1499 |
1461 |
7.18 |
|
|
|
| 10 |
A |
1477 |
1439 |
5.10 |
|
|
|
| 11 |
A |
1435 |
1398 |
27.60 |
|
|
|
| 12 |
A |
1360 |
1325 |
35.92 |
|
|
|
| 13 |
A |
1218 |
1187 |
14.07 |
|
|
|
| 14 |
A |
1154 |
1125 |
61.24 |
|
|
|
| 15 |
A |
1141 |
1112 |
3.12 |
|
|
|
| 16 |
A |
1087 |
1059 |
91.46 |
|
|
|
| 17 |
A |
1016 |
990 |
33.95 |
|
|
|
| 18 |
A |
804 |
784 |
17.65 |
|
|
|
| 19 |
A |
536 |
522 |
2.94 |
|
|
|
| 20 |
A |
497 |
484 |
9.19 |
|
|
|
| 21 |
A |
247 |
240 |
7.63 |
|
|
|
| 22 |
A |
213 |
208 |
3.46 |
|
|
|
| 23 |
A |
114 |
111 |
2.75 |
|
|
|
| 24 |
A |
3118 |
3039 |
33.50 |
|
|
|
| 25 |
A |
3107 |
3028 |
80.65 |
|
|
|
| 26 |
A |
3042 |
2964 |
3.98 |
|
|
|
| 27 |
A |
2952 |
2877 |
6.35 |
|
|
|
| 28 |
A |
1512 |
1473 |
7.20 |
|
|
|
| 29 |
A |
1510 |
1471 |
1.05 |
|
|
|
| 30 |
A |
1492 |
1454 |
6.28 |
|
|
|
| 31 |
A |
1472 |
1435 |
3.84 |
|
|
|
| 32 |
A |
1379 |
1344 |
26.31 |
|
|
|
| 33 |
A |
1184 |
1154 |
40.48 |
|
|
|
| 34 |
A |
1145 |
1116 |
12.92 |
|
|
|
| 35 |
A |
1112 |
1083 |
55.04 |
|
|
|
| 36 |
A |
1038 |
1012 |
138.42 |
|
|
|
| 37 |
A |
825 |
804 |
86.41 |
|
|
|
| 38 |
A |
370 |
361 |
0.74 |
|
|
|
| 39 |
A |
322 |
313 |
2.36 |
|
|
|
| 40 |
A |
190 |
185 |
0.47 |
|
|
|
| 41 |
A |
126 |
122 |
1.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30450.5 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29674.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.243 |
|
|
|
| 2 |
O |
-0.243 |
|
|
|
| 3 |
C |
-0.287 |
|
|
|
| 4 |
C |
-0.287 |
|
|
|
| 5 |
C |
0.058 |
|
|
|
| 6 |
C |
-0.447 |
|
|
|
| 7 |
H |
0.127 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.172 |
|
|
|
| 10 |
H |
0.172 |
|
|
|
| 11 |
H |
0.145 |
|
|
|
| 12 |
H |
0.145 |
|
|
|
| 13 |
H |
0.093 |
|
|
|
| 14 |
H |
0.172 |
|
|
|
| 15 |
H |
0.148 |
|
|
|
| 16 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.490 |
-2.472 |
0.000 |
2.887 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.963 |
-2.035 |
0.000 |
| y |
-2.035 |
-35.127 |
0.000 |
| z |
0.000 |
0.000 |
-38.552 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.124 |
-2.035 |
0.000 |
| y |
-2.035 |
3.631 |
0.000 |
| z |
0.000 |
0.000 |
-1.507 |
|
| Polar |
|---|
| 3z2-r2 | -3.014 |
|---|
| x2-y2 | -3.836 |
|---|
| xy | -2.035 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.379 |
-0.543 |
0.000 |
| y |
-0.543 |
8.898 |
0.000 |
| z |
0.000 |
0.000 |
8.911 |
<r2> (average value of r
2) Å
2
| <r2> |
161.959 |
| (<r2>)1/2 |
12.726 |