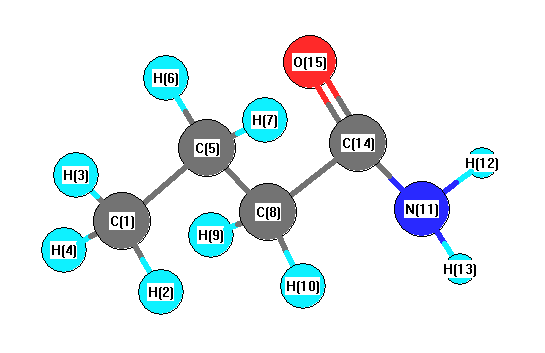
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3718 |
3623 |
15.56 |
|
|
|
| 2 |
A |
3558 |
3467 |
19.59 |
|
|
|
| 3 |
A |
3080 |
3002 |
83.24 |
|
|
|
| 4 |
A |
3069 |
2991 |
63.60 |
|
|
|
| 5 |
A |
3047 |
2970 |
7.36 |
|
|
|
| 6 |
A |
3035 |
2958 |
23.59 |
|
|
|
| 7 |
A |
2998 |
2922 |
39.16 |
|
|
|
| 8 |
A |
2988 |
2912 |
26.45 |
|
|
|
| 9 |
A |
2976 |
2900 |
39.51 |
|
|
|
| 10 |
A |
1668 |
1626 |
269.43 |
|
|
|
| 11 |
A |
1633 |
1591 |
95.06 |
|
|
|
| 12 |
A |
1533 |
1494 |
12.43 |
|
|
|
| 13 |
A |
1523 |
1484 |
8.57 |
|
|
|
| 14 |
A |
1518 |
1480 |
2.43 |
|
|
|
| 15 |
A |
1496 |
1458 |
8.59 |
|
|
|
| 16 |
A |
1433 |
1396 |
2.93 |
|
|
|
| 17 |
A |
1399 |
1363 |
75.46 |
|
|
|
| 18 |
A |
1365 |
1330 |
47.75 |
|
|
|
| 19 |
A |
1319 |
1286 |
22.43 |
|
|
|
| 20 |
A |
1293 |
1260 |
48.26 |
|
|
|
| 21 |
A |
1254 |
1222 |
15.42 |
|
|
|
| 22 |
A |
1138 |
1109 |
4.04 |
|
|
|
| 23 |
A |
1121 |
1092 |
7.11 |
|
|
|
| 24 |
A |
1069 |
1042 |
2.86 |
|
|
|
| 25 |
A |
1052 |
1025 |
2.42 |
|
|
|
| 26 |
A |
919 |
896 |
2.66 |
|
|
|
| 27 |
A |
890 |
868 |
1.82 |
|
|
|
| 28 |
A |
853 |
831 |
1.82 |
|
|
|
| 29 |
A |
758 |
739 |
8.22 |
|
|
|
| 30 |
A |
686 |
668 |
55.23 |
|
|
|
| 31 |
A |
602 |
587 |
65.02 |
|
|
|
| 32 |
A |
532 |
518 |
77.12 |
|
|
|
| 33 |
A |
493 |
481 |
124.12 |
|
|
|
| 34 |
A |
417 |
407 |
3.40 |
|
|
|
| 35 |
A |
332 |
323 |
1.81 |
|
|
|
| 36 |
A |
236 |
230 |
0.15 |
|
|
|
| 37 |
A |
185 |
180 |
6.60 |
|
|
|
| 38 |
A |
91 |
88 |
2.61 |
|
|
|
| 39 |
A |
43 |
42 |
5.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28660.6 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 27929.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.478 |
|
|
|
| 2 |
H |
0.143 |
|
|
|
| 3 |
H |
0.153 |
|
|
|
| 4 |
H |
0.136 |
|
|
|
| 5 |
C |
-0.214 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.172 |
|
|
|
| 8 |
C |
-0.403 |
|
|
|
| 9 |
H |
0.118 |
|
|
|
| 10 |
H |
0.163 |
|
|
|
| 11 |
N |
-0.498 |
|
|
|
| 12 |
H |
0.325 |
|
|
|
| 13 |
H |
0.315 |
|
|
|
| 14 |
C |
0.139 |
|
|
|
| 15 |
O |
-0.202 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.127 |
-4.086 |
-0.304 |
4.100 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.753 |
-7.522 |
-1.447 |
| y |
-7.522 |
-40.294 |
0.064 |
| z |
-1.447 |
0.064 |
-38.388 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.588 |
-7.522 |
-1.447 |
| y |
-7.522 |
-4.224 |
0.064 |
| z |
-1.447 |
0.064 |
-1.364 |
|
| Polar |
|---|
| 3z2-r2 | -2.729 |
|---|
| x2-y2 | 6.541 |
|---|
| xy | -7.522 |
|---|
| xz | -1.447 |
|---|
| yz | 0.064 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.815 |
-0.345 |
-0.285 |
| y |
-0.345 |
8.728 |
0.124 |
| z |
-0.285 |
0.124 |
6.651 |
<r2> (average value of r
2) Å
2
| <r2> |
181.389 |
| (<r2>)1/2 |
13.468 |