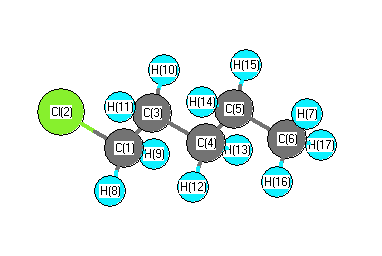
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3068 |
2989 |
65.65 |
|
|
|
| 2 |
A' |
3059 |
2981 |
31.85 |
|
|
|
| 3 |
A' |
3008 |
2931 |
32.51 |
|
|
|
| 4 |
A' |
2979 |
2903 |
20.80 |
|
|
|
| 5 |
A' |
2976 |
2900 |
89.37 |
|
|
|
| 6 |
A' |
2965 |
2889 |
10.66 |
|
|
|
| 7 |
A' |
1537 |
1498 |
12.30 |
|
|
|
| 8 |
A' |
1526 |
1487 |
0.64 |
|
|
|
| 9 |
A' |
1516 |
1477 |
1.04 |
|
|
|
| 10 |
A' |
1513 |
1475 |
0.98 |
|
|
|
| 11 |
A' |
1496 |
1458 |
1.36 |
|
|
|
| 12 |
A' |
1430 |
1394 |
4.87 |
|
|
|
| 13 |
A' |
1392 |
1356 |
0.04 |
|
|
|
| 14 |
A' |
1373 |
1338 |
17.73 |
|
|
|
| 15 |
A' |
1320 |
1286 |
16.54 |
|
|
|
| 16 |
A' |
1263 |
1231 |
12.37 |
|
|
|
| 17 |
A' |
1129 |
1100 |
3.92 |
|
|
|
| 18 |
A' |
1067 |
1040 |
0.27 |
|
|
|
| 19 |
A' |
1054 |
1027 |
11.98 |
|
|
|
| 20 |
A' |
1004 |
979 |
0.39 |
|
|
|
| 21 |
A' |
899 |
876 |
1.50 |
|
|
|
| 22 |
A' |
673 |
656 |
57.14 |
|
|
|
| 23 |
A' |
436 |
425 |
6.02 |
|
|
|
| 24 |
A' |
332 |
323 |
1.44 |
|
|
|
| 25 |
A' |
246 |
240 |
3.44 |
|
|
|
| 26 |
A' |
112 |
109 |
1.89 |
|
|
|
| 27 |
A" |
3144 |
3064 |
30.38 |
|
|
|
| 28 |
A" |
3067 |
2989 |
142.07 |
|
|
|
| 29 |
A" |
3060 |
2982 |
1.36 |
|
|
|
| 30 |
A" |
3025 |
2947 |
20.67 |
|
|
|
| 31 |
A" |
3000 |
2923 |
1.93 |
|
|
|
| 32 |
A" |
1522 |
1483 |
8.91 |
|
|
|
| 33 |
A" |
1329 |
1295 |
0.47 |
|
|
|
| 34 |
A" |
1326 |
1292 |
0.71 |
|
|
|
| 35 |
A" |
1287 |
1254 |
0.31 |
|
|
|
| 36 |
A" |
1226 |
1195 |
0.19 |
|
|
|
| 37 |
A" |
1095 |
1067 |
1.08 |
|
|
|
| 38 |
A" |
975 |
950 |
0.02 |
|
|
|
| 39 |
A" |
850 |
828 |
1.09 |
|
|
|
| 40 |
A" |
771 |
751 |
0.18 |
|
|
|
| 41 |
A" |
743 |
724 |
5.45 |
|
|
|
| 42 |
A" |
233 |
227 |
0.00 |
|
|
|
| 43 |
A" |
136 |
133 |
0.59 |
|
|
|
| 44 |
A" |
93 |
91 |
0.36 |
|
|
|
| 45 |
A" |
60 |
58 |
1.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33155.7 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 32310.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.510 |
|
|
|
| 2 |
Cl |
-0.042 |
|
|
|
| 3 |
C |
-0.156 |
|
|
|
| 4 |
C |
-0.243 |
|
|
|
| 5 |
C |
-0.196 |
|
|
|
| 6 |
C |
-0.474 |
|
|
|
| 7 |
H |
0.151 |
|
|
|
| 8 |
H |
0.195 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.148 |
|
|
|
| 11 |
H |
0.148 |
|
|
|
| 12 |
H |
0.122 |
|
|
|
| 13 |
H |
0.122 |
|
|
|
| 14 |
H |
0.129 |
|
|
|
| 15 |
H |
0.129 |
|
|
|
| 16 |
H |
0.142 |
|
|
|
| 17 |
H |
0.142 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.422 |
-2.936 |
0.000 |
2.966 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.309 |
3.637 |
0.000 |
| y |
3.637 |
-57.947 |
0.000 |
| z |
0.000 |
0.000 |
-45.471 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.400 |
3.637 |
0.000 |
| y |
3.637 |
-12.057 |
0.000 |
| z |
0.000 |
0.000 |
6.657 |
|
| Polar |
|---|
| 3z2-r2 | 13.313 |
|---|
| x2-y2 | 11.638 |
|---|
| xy | 3.637 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.523 |
-1.482 |
0.000 |
| y |
-1.482 |
13.524 |
0.000 |
| z |
0.000 |
0.000 |
8.392 |
<r2> (average value of r
2) Å
2
| <r2> |
272.795 |
| (<r2>)1/2 |
16.517 |