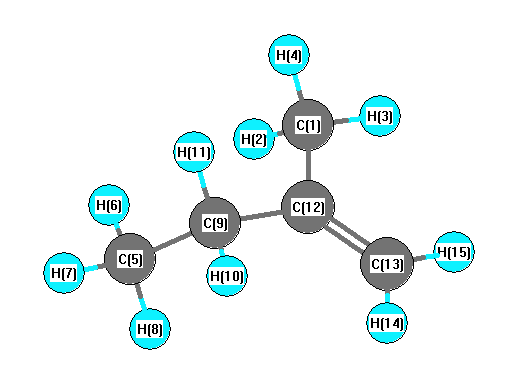
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3192 |
3110 |
39.46 |
|
|
|
| 2 |
A |
3095 |
3016 |
18.46 |
|
|
|
| 3 |
A |
3084 |
3005 |
39.63 |
|
|
|
| 4 |
A |
3078 |
3000 |
54.56 |
|
|
|
| 5 |
A |
3071 |
2993 |
73.09 |
|
|
|
| 6 |
A |
3040 |
2962 |
41.73 |
|
|
|
| 7 |
A |
3032 |
2955 |
16.64 |
|
|
|
| 8 |
A |
2987 |
2911 |
42.83 |
|
|
|
| 9 |
A |
2975 |
2899 |
40.45 |
|
|
|
| 10 |
A |
2968 |
2892 |
46.91 |
|
|
|
| 11 |
A |
1680 |
1638 |
21.91 |
|
|
|
| 12 |
A |
1531 |
1492 |
5.10 |
|
|
|
| 13 |
A |
1523 |
1484 |
7.78 |
|
|
|
| 14 |
A |
1518 |
1479 |
12.91 |
|
|
|
| 15 |
A |
1512 |
1473 |
13.56 |
|
|
|
| 16 |
A |
1501 |
1463 |
4.63 |
|
|
|
| 17 |
A |
1461 |
1424 |
0.25 |
|
|
|
| 18 |
A |
1432 |
1396 |
7.91 |
|
|
|
| 19 |
A |
1426 |
1389 |
5.78 |
|
|
|
| 20 |
A |
1353 |
1319 |
2.11 |
|
|
|
| 21 |
A |
1314 |
1281 |
2.11 |
|
|
|
| 22 |
A |
1276 |
1243 |
1.28 |
|
|
|
| 23 |
A |
1123 |
1094 |
0.24 |
|
|
|
| 24 |
A |
1099 |
1071 |
5.98 |
|
|
|
| 25 |
A |
1058 |
1031 |
2.00 |
|
|
|
| 26 |
A |
1023 |
997 |
0.92 |
|
|
|
| 27 |
A |
1008 |
983 |
1.02 |
|
|
|
| 28 |
A |
953 |
928 |
61.89 |
|
|
|
| 29 |
A |
939 |
915 |
1.51 |
|
|
|
| 30 |
A |
806 |
786 |
1.99 |
|
|
|
| 31 |
A |
764 |
745 |
0.96 |
|
|
|
| 32 |
A |
721 |
703 |
0.08 |
|
|
|
| 33 |
A |
531 |
517 |
8.53 |
|
|
|
| 34 |
A |
426 |
415 |
1.49 |
|
|
|
| 35 |
A |
392 |
382 |
1.00 |
|
|
|
| 36 |
A |
257 |
250 |
0.33 |
|
|
|
| 37 |
A |
215 |
209 |
0.60 |
|
|
|
| 38 |
A |
173 |
168 |
0.33 |
|
|
|
| 39 |
A |
67 |
65 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29801.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29041.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.666 |
|
|
|
| 2 |
H |
0.161 |
|
|
|
| 3 |
H |
0.160 |
|
|
|
| 4 |
H |
0.151 |
|
|
|
| 5 |
C |
-0.489 |
|
|
|
| 6 |
H |
0.146 |
|
|
|
| 7 |
H |
0.148 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
| 9 |
C |
-0.344 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
| 11 |
H |
0.131 |
|
|
|
| 12 |
C |
0.616 |
|
|
|
| 13 |
C |
-0.569 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.415 |
0.445 |
0.130 |
0.622 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.477 |
0.743 |
-0.877 |
| y |
0.743 |
-32.864 |
0.150 |
| z |
-0.877 |
0.150 |
-35.062 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.486 |
0.743 |
-0.877 |
| y |
0.743 |
1.405 |
0.150 |
| z |
-0.877 |
0.150 |
-1.891 |
|
| Polar |
|---|
| 3z2-r2 | -3.783 |
|---|
| x2-y2 | -0.612 |
|---|
| xy | 0.743 |
|---|
| xz | -0.877 |
|---|
| yz | 0.150 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.989 |
-1.057 |
-0.692 |
| y |
-1.057 |
9.600 |
0.408 |
| z |
-0.692 |
0.408 |
6.961 |
<r2> (average value of r
2) Å
2
| <r2> |
123.678 |
| (<r2>)1/2 |
11.121 |