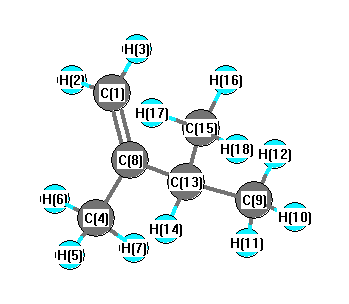
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3188 |
3107 |
39.78 |
|
|
|
| 2 |
A' |
3092 |
3014 |
22.14 |
|
|
|
| 3 |
A' |
3083 |
3005 |
37.33 |
|
|
|
| 4 |
A' |
3074 |
2996 |
45.40 |
|
|
|
| 5 |
A' |
3068 |
2990 |
131.25 |
|
|
|
| 6 |
A' |
2990 |
2914 |
5.53 |
|
|
|
| 7 |
A' |
2984 |
2908 |
62.31 |
|
|
|
| 8 |
A' |
2979 |
2903 |
17.42 |
|
|
|
| 9 |
A' |
1675 |
1632 |
22.92 |
|
|
|
| 10 |
A' |
1532 |
1493 |
12.61 |
|
|
|
| 11 |
A' |
1527 |
1488 |
6.03 |
|
|
|
| 12 |
A' |
1514 |
1476 |
13.64 |
|
|
|
| 13 |
A' |
1464 |
1427 |
0.51 |
|
|
|
| 14 |
A' |
1436 |
1400 |
6.46 |
|
|
|
| 15 |
A' |
1429 |
1393 |
11.07 |
|
|
|
| 16 |
A' |
1346 |
1312 |
5.08 |
|
|
|
| 17 |
A' |
1308 |
1275 |
0.60 |
|
|
|
| 18 |
A' |
1192 |
1161 |
3.60 |
|
|
|
| 19 |
A' |
1115 |
1086 |
13.19 |
|
|
|
| 20 |
A' |
1015 |
989 |
1.38 |
|
|
|
| 21 |
A' |
963 |
939 |
0.63 |
|
|
|
| 22 |
A' |
892 |
869 |
1.20 |
|
|
|
| 23 |
A' |
716 |
697 |
0.97 |
|
|
|
| 24 |
A' |
519 |
506 |
1.17 |
|
|
|
| 25 |
A' |
436 |
425 |
0.74 |
|
|
|
| 26 |
A' |
327 |
319 |
0.31 |
|
|
|
| 27 |
A' |
271 |
264 |
0.50 |
|
|
|
| 28 |
A' |
245 |
239 |
0.06 |
|
|
|
| 29 |
A" |
3071 |
2993 |
29.94 |
|
|
|
| 30 |
A" |
3060 |
2982 |
5.02 |
|
|
|
| 31 |
A" |
3045 |
2967 |
40.26 |
|
|
|
| 32 |
A" |
2977 |
2901 |
39.64 |
|
|
|
| 33 |
A" |
1520 |
1481 |
0.28 |
|
|
|
| 34 |
A" |
1512 |
1474 |
0.80 |
|
|
|
| 35 |
A" |
1502 |
1463 |
12.31 |
|
|
|
| 36 |
A" |
1414 |
1378 |
7.99 |
|
|
|
| 37 |
A" |
1347 |
1312 |
3.00 |
|
|
|
| 38 |
A" |
1124 |
1095 |
3.63 |
|
|
|
| 39 |
A" |
1074 |
1047 |
1.67 |
|
|
|
| 40 |
A" |
968 |
943 |
0.19 |
|
|
|
| 41 |
A" |
955 |
930 |
58.23 |
|
|
|
| 42 |
A" |
936 |
912 |
0.70 |
|
|
|
| 43 |
A" |
741 |
722 |
0.57 |
|
|
|
| 44 |
A" |
550 |
536 |
9.87 |
|
|
|
| 45 |
A" |
247 |
241 |
0.19 |
|
|
|
| 46 |
A" |
209 |
203 |
0.60 |
|
|
|
| 47 |
A" |
158 |
154 |
0.24 |
|
|
|
| 48 |
A" |
52 |
51 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35919.9 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 35003.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.594 |
|
|
|
| 2 |
C |
0.635 |
|
|
|
| 3 |
C |
-0.059 |
|
|
|
| 4 |
C |
-0.701 |
|
|
|
| 5 |
C |
-0.521 |
|
|
|
| 6 |
C |
-0.521 |
|
|
|
| 7 |
H |
0.129 |
|
|
|
| 8 |
H |
0.133 |
|
|
|
| 9 |
H |
0.134 |
|
|
|
| 10 |
H |
0.159 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
| 12 |
H |
0.162 |
|
|
|
| 13 |
H |
0.145 |
|
|
|
| 14 |
H |
0.154 |
|
|
|
| 15 |
H |
0.142 |
|
|
|
| 16 |
H |
0.145 |
|
|
|
| 17 |
H |
0.154 |
|
|
|
| 18 |
H |
0.142 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.350 |
0.458 |
0.000 |
0.576 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.419 |
-0.896 |
0.000 |
| y |
-0.896 |
-40.331 |
0.000 |
| z |
0.000 |
0.000 |
-42.216 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.854 |
-0.896 |
0.000 |
| y |
-0.896 |
0.487 |
0.000 |
| z |
0.000 |
0.000 |
-2.341 |
|
| Polar |
|---|
| 3z2-r2 | -4.683 |
|---|
| x2-y2 | 0.911 |
|---|
| xy | -0.896 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.723 |
0.986 |
0.000 |
| y |
0.986 |
12.139 |
0.000 |
| z |
0.000 |
0.000 |
8.803 |
<r2> (average value of r
2) Å
2
| <r2> |
163.534 |
| (<r2>)1/2 |
12.788 |