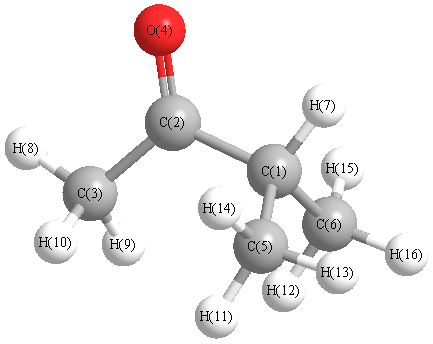
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3117 |
3038 |
33.16 |
|
|
|
| 2 |
A |
3086 |
3007 |
15.34 |
|
|
|
| 3 |
A |
3080 |
3002 |
67.73 |
|
|
|
| 4 |
A |
3077 |
2999 |
79.88 |
|
|
|
| 5 |
A |
3076 |
2997 |
26.78 |
|
|
|
| 6 |
A |
3068 |
2990 |
0.62 |
|
|
|
| 7 |
A |
3027 |
2950 |
2.39 |
|
|
|
| 8 |
A |
3005 |
2928 |
6.73 |
|
|
|
| 9 |
A |
2988 |
2912 |
24.23 |
|
|
|
| 10 |
A |
2984 |
2908 |
39.39 |
|
|
|
| 11 |
A |
1646 |
1604 |
141.45 |
|
|
|
| 12 |
A |
1531 |
1492 |
23.80 |
|
|
|
| 13 |
A |
1526 |
1487 |
10.88 |
|
|
|
| 14 |
A |
1516 |
1478 |
7.98 |
|
|
|
| 15 |
A |
1511 |
1473 |
0.20 |
|
|
|
| 16 |
A |
1498 |
1460 |
10.93 |
|
|
|
| 17 |
A |
1494 |
1456 |
5.18 |
|
|
|
| 18 |
A |
1439 |
1402 |
3.80 |
|
|
|
| 19 |
A |
1418 |
1382 |
8.03 |
|
|
|
| 20 |
A |
1414 |
1378 |
38.51 |
|
|
|
| 21 |
A |
1354 |
1320 |
1.18 |
|
|
|
| 22 |
A |
1342 |
1308 |
5.96 |
|
|
|
| 23 |
A |
1269 |
1236 |
46.75 |
|
|
|
| 24 |
A |
1190 |
1159 |
1.49 |
|
|
|
| 25 |
A |
1125 |
1096 |
0.22 |
|
|
|
| 26 |
A |
1088 |
1060 |
26.82 |
|
|
|
| 27 |
A |
1022 |
996 |
3.57 |
|
|
|
| 28 |
A |
975 |
950 |
0.77 |
|
|
|
| 29 |
A |
973 |
948 |
25.76 |
|
|
|
| 30 |
A |
942 |
918 |
1.23 |
|
|
|
| 31 |
A |
893 |
871 |
3.82 |
|
|
|
| 32 |
A |
705 |
687 |
3.02 |
|
|
|
| 33 |
A |
610 |
595 |
0.00 |
|
|
|
| 34 |
A |
518 |
505 |
3.76 |
|
|
|
| 35 |
A |
486 |
474 |
9.91 |
|
|
|
| 36 |
A |
327 |
319 |
2.37 |
|
|
|
| 37 |
A |
269 |
262 |
1.61 |
|
|
|
| 38 |
A |
238 |
232 |
0.43 |
|
|
|
| 39 |
A |
221 |
215 |
0.08 |
|
|
|
| 40 |
A |
190 |
185 |
0.24 |
|
|
|
| 41 |
A |
69 |
67 |
3.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30653.0 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 29871.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.151 |
|
|
|
| 2 |
C |
0.116 |
|
|
|
| 3 |
H |
0.184 |
|
|
|
| 4 |
H |
0.184 |
|
|
|
| 5 |
H |
0.157 |
|
|
|
| 6 |
C |
-0.568 |
|
|
|
| 7 |
H |
0.157 |
|
|
|
| 8 |
C |
0.006 |
|
|
|
| 9 |
H |
0.165 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.141 |
|
|
|
| 12 |
H |
0.142 |
|
|
|
| 13 |
H |
0.157 |
|
|
|
| 14 |
H |
0.157 |
|
|
|
| 15 |
C |
-0.508 |
|
|
|
| 16 |
C |
-0.506 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.213 |
0.428 |
2.489 |
3.358 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.711 |
0.959 |
5.602 |
| y |
0.959 |
-37.661 |
-0.636 |
| z |
5.602 |
-0.636 |
-41.385 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.188 |
0.959 |
5.602 |
| y |
0.959 |
3.887 |
-0.636 |
| z |
5.602 |
-0.636 |
-1.698 |
|
| Polar |
|---|
| 3z2-r2 | -3.397 |
|---|
| x2-y2 | -4.050 |
|---|
| xy | 0.959 |
|---|
| xz | 5.602 |
|---|
| yz | -0.636 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.747 |
-0.037 |
-0.203 |
| y |
-0.037 |
8.218 |
0.133 |
| z |
-0.203 |
0.133 |
8.878 |
<r2> (average value of r
2) Å
2
| <r2> |
153.860 |
| (<r2>)1/2 |
12.404 |