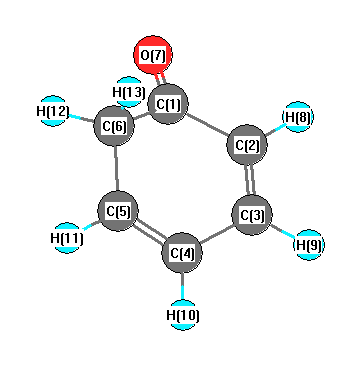
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3179 |
3098 |
0.10 |
|
|
|
| 2 |
A1 |
3133 |
3053 |
12.21 |
|
|
|
| 3 |
A1 |
2960 |
2884 |
18.24 |
|
|
|
| 4 |
A1 |
1685 |
1642 |
108.32 |
|
|
|
| 5 |
A1 |
1594 |
1553 |
158.60 |
|
|
|
| 6 |
A1 |
1468 |
1431 |
25.54 |
|
|
|
| 7 |
A1 |
1406 |
1370 |
0.18 |
|
|
|
| 8 |
A1 |
1199 |
1168 |
10.57 |
|
|
|
| 9 |
A1 |
958 |
934 |
5.48 |
|
|
|
| 10 |
A1 |
873 |
851 |
10.35 |
|
|
|
| 11 |
A1 |
764 |
744 |
0.84 |
|
|
|
| 12 |
A1 |
500 |
488 |
3.84 |
|
|
|
| 13 |
A2 |
1202 |
1171 |
0.00 |
|
|
|
| 14 |
A2 |
1019 |
993 |
0.00 |
|
|
|
| 15 |
A2 |
763 |
744 |
0.00 |
|
|
|
| 16 |
A2 |
365 |
355 |
0.00 |
|
|
|
| 17 |
B1 |
2981 |
2905 |
16.34 |
|
|
|
| 18 |
B1 |
1037 |
1010 |
0.21 |
|
|
|
| 19 |
B1 |
963 |
938 |
26.58 |
|
|
|
| 20 |
B1 |
849 |
827 |
48.72 |
|
|
|
| 21 |
B1 |
584 |
570 |
28.63 |
|
|
|
| 22 |
B1 |
313 |
305 |
3.91 |
|
|
|
| 23 |
B1 |
132 |
129 |
0.95 |
|
|
|
| 24 |
B2 |
3175 |
3094 |
37.07 |
|
|
|
| 25 |
B2 |
3131 |
3052 |
24.47 |
|
|
|
| 26 |
B2 |
1631 |
1589 |
5.02 |
|
|
|
| 27 |
B2 |
1413 |
1377 |
31.81 |
|
|
|
| 28 |
B2 |
1379 |
1343 |
0.04 |
|
|
|
| 29 |
B2 |
1289 |
1256 |
25.72 |
|
|
|
| 30 |
B2 |
1150 |
1120 |
10.92 |
|
|
|
| 31 |
B2 |
995 |
969 |
9.81 |
|
|
|
| 32 |
B2 |
579 |
564 |
1.27 |
|
|
|
| 33 |
B2 |
438 |
427 |
14.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22552.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 21977.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.039 |
|
|
|
| 2 |
C |
-0.528 |
|
|
|
| 3 |
C |
-0.176 |
|
|
|
| 4 |
C |
-0.176 |
|
|
|
| 5 |
C |
0.029 |
|
|
|
| 6 |
C |
0.029 |
|
|
|
| 7 |
O |
-0.140 |
|
|
|
| 8 |
H |
0.165 |
|
|
|
| 9 |
H |
0.165 |
|
|
|
| 10 |
H |
0.158 |
|
|
|
| 11 |
H |
0.158 |
|
|
|
| 12 |
H |
0.178 |
|
|
|
| 13 |
H |
0.178 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-5.249 |
5.249 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.729 |
0.000 |
0.000 |
| y |
0.000 |
-36.253 |
0.000 |
| z |
0.000 |
0.000 |
-47.217 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.995 |
0.000 |
0.000 |
| y |
0.000 |
8.720 |
0.000 |
| z |
0.000 |
0.000 |
-7.725 |
|
| Polar |
|---|
| 3z2-r2 | -15.451 |
|---|
| x2-y2 | -6.477 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.284 |
0.000 |
0.000 |
| y |
0.000 |
9.608 |
0.000 |
| z |
0.000 |
0.000 |
15.056 |
<r2> (average value of r
2) Å
2
| <r2> |
154.653 |
| (<r2>)1/2 |
12.436 |