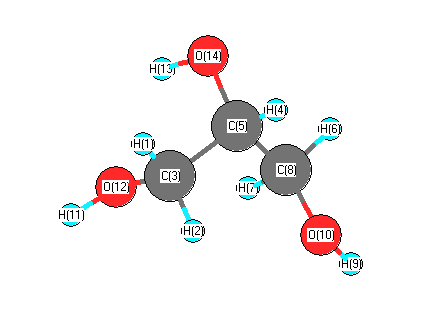
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3701 |
3607 |
10.29 |
|
|
|
| 2 |
A |
3679 |
3585 |
5.11 |
|
|
|
| 3 |
A |
3614 |
3522 |
25.68 |
|
|
|
| 4 |
A |
3084 |
3005 |
60.70 |
|
|
|
| 5 |
A |
3080 |
3002 |
34.34 |
|
|
|
| 6 |
A |
3059 |
2981 |
25.22 |
|
|
|
| 7 |
A |
3002 |
2925 |
29.23 |
|
|
|
| 8 |
A |
2997 |
2921 |
65.12 |
|
|
|
| 9 |
A |
1525 |
1487 |
7.80 |
|
|
|
| 10 |
A |
1514 |
1475 |
10.37 |
|
|
|
| 11 |
A |
1443 |
1406 |
3.75 |
|
|
|
| 12 |
A |
1419 |
1383 |
11.17 |
|
|
|
| 13 |
A |
1406 |
1370 |
17.88 |
|
|
|
| 14 |
A |
1368 |
1333 |
18.63 |
|
|
|
| 15 |
A |
1296 |
1263 |
56.88 |
|
|
|
| 16 |
A |
1268 |
1236 |
14.19 |
|
|
|
| 17 |
A |
1211 |
1180 |
6.52 |
|
|
|
| 18 |
A |
1202 |
1171 |
1.23 |
|
|
|
| 19 |
A |
1150 |
1121 |
26.48 |
|
|
|
| 20 |
A |
1125 |
1096 |
9.30 |
|
|
|
| 21 |
A |
1073 |
1046 |
72.60 |
|
|
|
| 22 |
A |
1017 |
991 |
107.18 |
|
|
|
| 23 |
A |
997 |
971 |
82.23 |
|
|
|
| 24 |
A |
960 |
935 |
26.95 |
|
|
|
| 25 |
A |
928 |
904 |
18.77 |
|
|
|
| 26 |
A |
816 |
795 |
24.07 |
|
|
|
| 27 |
A |
630 |
614 |
37.10 |
|
|
|
| 28 |
A |
491 |
478 |
177.28 |
|
|
|
| 29 |
A |
463 |
451 |
8.59 |
|
|
|
| 30 |
A |
387 |
377 |
8.05 |
|
|
|
| 31 |
A |
265 |
259 |
20.50 |
|
|
|
| 32 |
A |
253 |
246 |
150.72 |
|
|
|
| 33 |
A |
240 |
234 |
53.28 |
|
|
|
| 34 |
A |
230 |
224 |
97.87 |
|
|
|
| 35 |
A |
139 |
136 |
34.42 |
|
|
|
| 36 |
A |
89 |
87 |
2.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25560.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 24908.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.148 |
|
|
|
| 2 |
H |
0.167 |
|
|
|
| 3 |
C |
-0.227 |
|
|
|
| 4 |
H |
0.150 |
|
|
|
| 5 |
C |
0.130 |
|
|
|
| 6 |
H |
0.149 |
|
|
|
| 7 |
H |
0.139 |
|
|
|
| 8 |
C |
-0.203 |
|
|
|
| 9 |
H |
0.342 |
|
|
|
| 10 |
O |
-0.493 |
|
|
|
| 11 |
H |
0.352 |
|
|
|
| 12 |
O |
-0.522 |
|
|
|
| 13 |
H |
0.353 |
|
|
|
| 14 |
O |
-0.486 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.083 |
-2.529 |
-0.750 |
2.639 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.510 |
0.057 |
0.100 |
| y |
0.057 |
6.785 |
-0.187 |
| z |
0.100 |
-0.187 |
6.497 |
<r2> (average value of r
2) Å
2
| <r2> |
148.598 |
| (<r2>)1/2 |
12.190 |